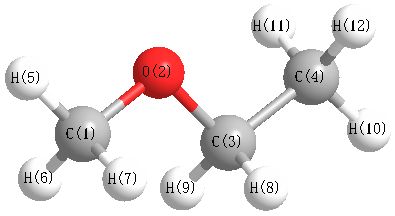
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3129 |
3012 |
36.03 |
|
|
|
| 2 |
A' |
3124 |
3008 |
31.96 |
|
|
|
| 3 |
A' |
3049 |
2935 |
20.43 |
|
|
|
| 4 |
A' |
2976 |
2865 |
100.67 |
|
|
|
| 5 |
A' |
2958 |
2848 |
39.60 |
|
|
|
| 6 |
A' |
1546 |
1488 |
2.79 |
|
|
|
| 7 |
A' |
1521 |
1464 |
4.92 |
|
|
|
| 8 |
A' |
1513 |
1456 |
8.25 |
|
|
|
| 9 |
A' |
1489 |
1433 |
0.36 |
|
|
|
| 10 |
A' |
1439 |
1385 |
38.47 |
|
|
|
| 11 |
A' |
1403 |
1350 |
0.79 |
|
|
|
| 12 |
A' |
1248 |
1201 |
46.83 |
|
|
|
| 13 |
A' |
1179 |
1135 |
145.67 |
|
|
|
| 14 |
A' |
1127 |
1085 |
5.95 |
|
|
|
| 15 |
A' |
1047 |
1008 |
19.14 |
|
|
|
| 16 |
A' |
879 |
847 |
11.46 |
|
|
|
| 17 |
A' |
469 |
451 |
0.55 |
|
|
|
| 18 |
A' |
288 |
277 |
2.75 |
|
|
|
| 19 |
A" |
3128 |
3011 |
35.53 |
|
|
|
| 20 |
A" |
3019 |
2906 |
76.11 |
|
|
|
| 21 |
A" |
2985 |
2874 |
83.70 |
|
|
|
| 22 |
A" |
1498 |
1442 |
11.17 |
|
|
|
| 23 |
A" |
1495 |
1439 |
5.04 |
|
|
|
| 24 |
A" |
1312 |
1263 |
1.97 |
|
|
|
| 25 |
A" |
1206 |
1161 |
10.11 |
|
|
|
| 26 |
A" |
1175 |
1131 |
0.31 |
|
|
|
| 27 |
A" |
828 |
797 |
0.36 |
|
|
|
| 28 |
A" |
269 |
259 |
2.27 |
|
|
|
| 29 |
A" |
214 |
206 |
1.33 |
|
|
|
| 30 |
A" |
117 |
113 |
2.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23813.3 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 22925.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.450 |
|
|
|
| 2 |
O |
-0.328 |
|
|
|
| 3 |
C |
-0.215 |
|
|
|
| 4 |
C |
-0.668 |
|
|
|
| 5 |
H |
0.223 |
|
|
|
| 6 |
H |
0.195 |
|
|
|
| 7 |
H |
0.195 |
|
|
|
| 8 |
H |
0.189 |
|
|
|
| 9 |
H |
0.189 |
|
|
|
| 10 |
H |
0.215 |
|
|
|
| 11 |
H |
0.228 |
|
|
|
| 12 |
H |
0.228 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.976 |
-0.684 |
0.000 |
1.192 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.669 |
1.753 |
0.000 |
| y |
1.753 |
-25.543 |
0.000 |
| z |
0.000 |
0.000 |
-26.552 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.379 |
1.753 |
0.000 |
| y |
1.753 |
0.567 |
0.000 |
| z |
0.000 |
0.000 |
-0.946 |
|
| Polar |
|---|
| 3z2-r2 | -1.892 |
|---|
| x2-y2 | -0.126 |
|---|
| xy | 1.753 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.187 |
0.728 |
0.000 |
| y |
0.728 |
6.169 |
0.000 |
| z |
0.000 |
0.000 |
5.343 |
<r2> (average value of r
2) Å
2
| <r2> |
102.902 |
| (<r2>)1/2 |
10.144 |