return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (1-Butanethiol, 3-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-594.935572
Energy at 298.15K-594.948596
HF Energy-593.892609
Nuclear repulsion energy302.665984
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3164 2996 60.90      
2 A' 3160 2993 45.50      
3 A' 3114 2949 22.34      
4 A' 3077 2914 21.68      
5 A' 3072 2910 18.40      
6 A' 3064 2902 5.89      
7 A' 2682 2540 12.78      
8 A' 1553 1471 17.88      
9 A' 1548 1466 10.28      
10 A' 1534 1453 0.17      
11 A' 1521 1441 8.71      
12 A' 1459 1381 11.00      
13 A' 1422 1346 3.95      
14 A' 1382 1309 19.96      
15 A' 1307 1238 9.71      
16 A' 1239 1173 1.84      
17 A' 1172 1110 2.65      
18 A' 1050 995 0.62      
19 A' 1016 962 2.51      
20 A' 901 854 1.03      
21 A' 808 765 2.08      
22 A' 795 753 2.63      
23 A' 539 510 1.35      
24 A' 390 370 0.38      
25 A' 274 260 0.46      
26 A' 251 238 0.53      
27 A' 194 183 1.94      
28 A" 3182 3013 9.44      
29 A" 3159 2992 20.55      
30 A" 3157 2990 3.27      
31 A" 3122 2957 11.97      
32 A" 3070 2908 28.26      
33 A" 1538 1457 1.93      
34 A" 1523 1442 0.85      
35 A" 1440 1364 16.25      
36 A" 1404 1330 0.16      
37 A" 1345 1273 0.11      
38 A" 1248 1182 4.90      
39 A" 1142 1081 1.24      
40 A" 1014 961 0.59      
41 A" 981 929 1.30      
42 A" 943 893 0.13      
43 A" 789 747 4.57      
44 A" 383 363 0.03      
45 A" 252 239 0.06      
46 A" 169 160 21.35      
47 A" 97 92 0.14      
48 A" 49 47 9.04      

Unscaled Zero Point Vibrational Energy (zpe) 35845.6 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 33949.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.19870 0.04048 0.03777

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.531 0.656 0.000
H2 1.902 1.689 0.000
S3 -2.519 -0.251 0.000
H4 -2.917 -1.535 0.000
C5 -0.727 -0.581 0.000
C6 0.000 0.759 0.000
C7 2.067 -0.025 1.259
C8 2.067 -0.025 -1.259
H9 -0.468 -1.158 0.889
H10 -0.468 -1.158 -0.889
H11 -0.316 1.332 -0.880
H12 -0.316 1.332 0.880
H13 3.157 0.051 -1.304
H14 3.157 0.051 1.304
H15 1.814 -1.088 1.278
H16 1.814 -1.088 -1.278
H17 1.661 0.437 2.164
H18 1.661 0.437 -2.164

Atom - Atom Distances (Å)
  C1 H2 S3 H4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.09804.15054.95842.57451.53451.52791.52792.84162.84162.15502.15502.17022.17022.18062.18062.17872.1787
H21.09804.82845.79853.47362.11762.13282.13283.80963.80962.41312.41312.44112.44113.05863.05862.51182.5118
S34.15054.82841.34431.82222.71414.76114.76112.41242.41242.85192.85195.83175.83174.59444.59444.75704.7570
H44.95845.79851.34432.38903.71115.35785.35782.63272.63273.96993.96996.41166.41164.92084.92085.43405.4340
C52.57453.47361.82222.38901.52423.11453.11451.09101.09102.14522.14524.14534.14532.88912.88913.37933.3793
C61.53452.11762.71413.71111.52422.54382.54382.16402.16401.09661.09663.48833.48832.88712.88712.74682.7468
C71.52792.13284.76115.35783.11452.54382.51742.80113.51023.47762.76862.78581.09351.09342.76231.09383.4771
C81.52792.13284.76115.35783.11452.54382.51743.51022.80112.76863.47761.09352.78582.76231.09343.47711.0938
H92.84163.80962.41242.63271.09102.16402.80113.51021.77793.05792.49464.40553.84342.31563.14752.94944.0488
H102.84163.80962.41242.63271.09102.16403.51022.80111.77792.49463.05793.84344.40553.14752.31564.04882.9494
H112.15502.41312.85193.96992.14521.09663.47762.76863.05792.49461.75923.72644.29813.87963.24883.73822.5223
H122.15502.41312.85193.96992.14521.09662.76863.47762.49463.05791.75924.29813.72643.24883.87962.52233.7382
H132.17022.44115.83176.41164.14533.48832.78581.09354.40553.84343.72644.29812.60793.12541.76123.79621.7679
H142.17022.44115.83176.41164.14533.48831.09352.78583.84344.40554.29813.72642.60791.76123.12541.76793.7962
H152.18063.05864.59444.92082.88912.88711.09342.76232.31563.14753.87963.24883.12541.76122.55621.76983.7675
H162.18063.05864.59444.92082.88912.88712.76231.09343.14752.31563.24883.87961.76123.12542.55623.76751.7698
H172.17872.51184.75705.43403.37932.74681.09383.47712.94944.04883.73822.52233.79621.76791.76983.76754.3275
H182.17872.51184.75705.43403.37932.74683.47711.09384.04882.94942.52233.73821.76793.79623.76751.76984.3275

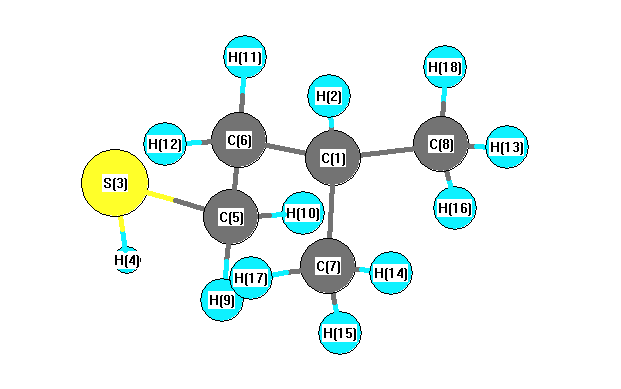
picture of 1-Butanethiol, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C6 C5 114.635 C1 C6 H11 108.841
C1 C6 H12 108.841 C1 C7 H14 110.673
C1 C7 H15 111.504 C1 C7 H17 111.328
C1 C8 H13 110.673 C1 C8 H16 111.504
C1 C8 H18 111.328 H2 C1 C6 105.908
H2 C1 C7 107.489 H2 C1 C8 107.489
S3 C5 C6 108.066 S3 C5 H9 109.224
S3 C5 H10 109.224 H4 S3 C5 96.794
C5 C6 H11 108.782 C5 C6 H12 108.782
C6 C1 C7 112.328 C6 C1 C8 112.328
C6 C5 H9 110.581 C6 C5 H10 110.581
C7 C1 C8 110.931 H9 C5 H10 109.134
H11 C6 H12 106.667 H13 C8 H16 107.285
H13 C8 H18 107.855 H14 C7 H15 107.285
H14 C7 H17 107.855 H15 C7 H17 108.021
H16 C8 H18 108.021
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability