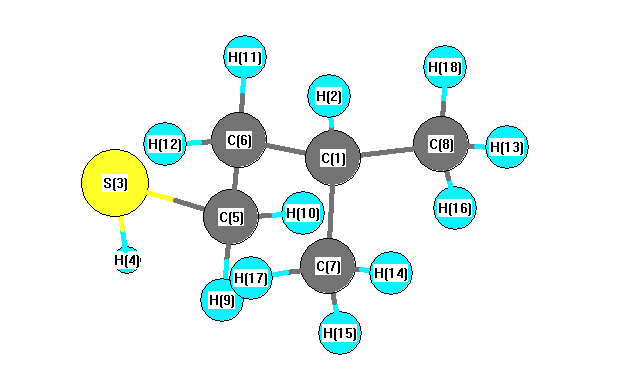
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3125 |
2982 |
67.78 |
|
|
|
| 2 |
A' |
3123 |
2981 |
78.12 |
|
|
|
| 3 |
A' |
3090 |
2949 |
28.24 |
|
|
|
| 4 |
A' |
3055 |
2916 |
7.38 |
|
|
|
| 5 |
A' |
3051 |
2912 |
36.34 |
|
|
|
| 6 |
A' |
3035 |
2897 |
9.54 |
|
|
|
| 7 |
A' |
2660 |
2539 |
15.13 |
|
|
|
| 8 |
A' |
1533 |
1463 |
15.23 |
|
|
|
| 9 |
A' |
1529 |
1459 |
10.64 |
|
|
|
| 10 |
A' |
1512 |
1443 |
0.97 |
|
|
|
| 11 |
A' |
1502 |
1434 |
7.61 |
|
|
|
| 12 |
A' |
1439 |
1374 |
9.12 |
|
|
|
| 13 |
A' |
1406 |
1342 |
3.15 |
|
|
|
| 14 |
A' |
1362 |
1300 |
23.50 |
|
|
|
| 15 |
A' |
1285 |
1227 |
12.98 |
|
|
|
| 16 |
A' |
1220 |
1164 |
2.65 |
|
|
|
| 17 |
A' |
1157 |
1104 |
2.56 |
|
|
|
| 18 |
A' |
1036 |
989 |
0.61 |
|
|
|
| 19 |
A' |
998 |
953 |
2.37 |
|
|
|
| 20 |
A' |
885 |
845 |
0.85 |
|
|
|
| 21 |
A' |
794 |
757 |
1.64 |
|
|
|
| 22 |
A' |
771 |
736 |
5.85 |
|
|
|
| 23 |
A' |
528 |
504 |
1.11 |
|
|
|
| 24 |
A' |
380 |
363 |
0.48 |
|
|
|
| 25 |
A' |
257 |
245 |
0.60 |
|
|
|
| 26 |
A' |
245 |
234 |
0.55 |
|
|
|
| 27 |
A' |
183 |
175 |
1.84 |
|
|
|
| 28 |
A" |
3148 |
3004 |
18.77 |
|
|
|
| 29 |
A" |
3123 |
2980 |
18.39 |
|
|
|
| 30 |
A" |
3115 |
2973 |
11.93 |
|
|
|
| 31 |
A" |
3088 |
2947 |
13.84 |
|
|
|
| 32 |
A" |
3047 |
2908 |
34.92 |
|
|
|
| 33 |
A" |
1519 |
1450 |
1.55 |
|
|
|
| 34 |
A" |
1504 |
1436 |
0.64 |
|
|
|
| 35 |
A" |
1420 |
1356 |
14.18 |
|
|
|
| 36 |
A" |
1386 |
1323 |
0.29 |
|
|
|
| 37 |
A" |
1334 |
1274 |
0.08 |
|
|
|
| 38 |
A" |
1233 |
1177 |
4.85 |
|
|
|
| 39 |
A" |
1120 |
1069 |
0.84 |
|
|
|
| 40 |
A" |
994 |
948 |
0.56 |
|
|
|
| 41 |
A" |
965 |
921 |
1.60 |
|
|
|
| 42 |
A" |
931 |
888 |
0.10 |
|
|
|
| 43 |
A" |
778 |
742 |
4.93 |
|
|
|
| 44 |
A" |
371 |
354 |
0.01 |
|
|
|
| 45 |
A" |
244 |
233 |
0.09 |
|
|
|
| 46 |
A" |
179 |
171 |
19.17 |
|
|
|
| 47 |
A" |
97 |
93 |
1.05 |
|
|
|
| 48 |
A" |
52 |
50 |
8.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35404.5 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 33790.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.259 |
|
|
|
| 2 |
H |
0.224 |
|
|
|
| 3 |
S |
-0.112 |
|
|
|
| 4 |
H |
0.152 |
|
|
|
| 5 |
C |
-0.600 |
|
|
|
| 6 |
C |
-0.459 |
|
|
|
| 7 |
C |
-0.649 |
|
|
|
| 8 |
C |
-0.649 |
|
|
|
| 9 |
H |
0.262 |
|
|
|
| 10 |
H |
0.262 |
|
|
|
| 11 |
H |
0.241 |
|
|
|
| 12 |
H |
0.241 |
|
|
|
| 13 |
H |
0.227 |
|
|
|
| 14 |
H |
0.227 |
|
|
|
| 15 |
H |
0.219 |
|
|
|
| 16 |
H |
0.219 |
|
|
|
| 17 |
H |
0.226 |
|
|
|
| 18 |
H |
0.226 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.665 |
-0.914 |
0.000 |
1.900 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-51.493 |
3.587 |
0.000 |
| y |
3.587 |
-45.217 |
0.000 |
| z |
0.000 |
0.000 |
-49.595 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.087 |
3.587 |
0.000 |
| y |
3.587 |
5.327 |
0.000 |
| z |
0.000 |
0.000 |
-1.240 |
|
| Polar |
|---|
| 3z2-r2 | -2.480 |
|---|
| x2-y2 | -6.276 |
|---|
| xy | 3.587 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
308.855 |
| (<r2>)1/2 |
17.574 |