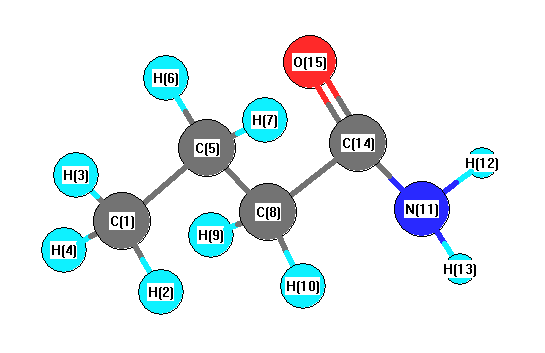
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3979 |
3598 |
34.76 |
|
|
|
| 2 |
A |
3848 |
3480 |
40.91 |
|
|
|
| 3 |
A |
3257 |
2945 |
82.90 |
|
|
|
| 4 |
A |
3237 |
2928 |
75.78 |
|
|
|
| 5 |
A |
3215 |
2908 |
31.08 |
|
|
|
| 6 |
A |
3208 |
2901 |
8.57 |
|
|
|
| 7 |
A |
3200 |
2894 |
23.31 |
|
|
|
| 8 |
A |
3174 |
2870 |
44.77 |
|
|
|
| 9 |
A |
3168 |
2865 |
22.26 |
|
|
|
| 10 |
A |
1959 |
1772 |
407.79 |
|
|
|
| 11 |
A |
1810 |
1637 |
123.74 |
|
|
|
| 12 |
A |
1641 |
1484 |
6.25 |
|
|
|
| 13 |
A |
1633 |
1477 |
7.02 |
|
|
|
| 14 |
A |
1623 |
1468 |
2.53 |
|
|
|
| 15 |
A |
1605 |
1451 |
7.19 |
|
|
|
| 16 |
A |
1571 |
1420 |
63.17 |
|
|
|
| 17 |
A |
1551 |
1403 |
7.75 |
|
|
|
| 18 |
A |
1476 |
1335 |
56.23 |
|
|
|
| 19 |
A |
1450 |
1312 |
5.84 |
|
|
|
| 20 |
A |
1374 |
1243 |
107.15 |
|
|
|
| 21 |
A |
1371 |
1240 |
5.97 |
|
|
|
| 22 |
A |
1237 |
1118 |
2.75 |
|
|
|
| 23 |
A |
1227 |
1109 |
1.81 |
|
|
|
| 24 |
A |
1175 |
1063 |
6.47 |
|
|
|
| 25 |
A |
1116 |
1010 |
2.27 |
|
|
|
| 26 |
A |
994 |
899 |
2.94 |
|
|
|
| 27 |
A |
966 |
874 |
2.84 |
|
|
|
| 28 |
A |
913 |
826 |
1.14 |
|
|
|
| 29 |
A |
810 |
732 |
8.05 |
|
|
|
| 30 |
A |
682 |
617 |
11.09 |
|
|
|
| 31 |
A |
670 |
606 |
26.53 |
|
|
|
| 32 |
A |
540 |
489 |
6.33 |
|
|
|
| 33 |
A |
464 |
420 |
2.48 |
|
|
|
| 34 |
A |
367 |
332 |
4.09 |
|
|
|
| 35 |
A |
263 |
238 |
0.33 |
|
|
|
| 36 |
A |
240 |
217 |
283.84 |
|
|
|
| 37 |
A |
195 |
176 |
9.72 |
|
|
|
| 38 |
A |
107 |
97 |
0.92 |
|
|
|
| 39 |
A |
31 |
28 |
1.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30671.6 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 27739.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.615 |
-0.204 |
|
-0.335 |
| 2 |
H |
0.202 |
0.039 |
|
0.075 |
| 3 |
H |
0.216 |
0.041 |
|
0.073 |
| 4 |
H |
0.201 |
0.031 |
|
0.067 |
| 5 |
C |
-0.396 |
0.310 |
|
0.347 |
| 6 |
H |
0.217 |
-0.055 |
|
-0.046 |
| 7 |
H |
0.227 |
-0.054 |
|
-0.045 |
| 8 |
C |
-0.546 |
-0.262 |
|
-0.377 |
| 9 |
H |
0.225 |
0.065 |
|
0.102 |
| 10 |
H |
0.229 |
0.046 |
|
0.081 |
| 11 |
N |
-0.902 |
-1.102 |
|
-1.116 |
| 12 |
H |
0.378 |
0.454 |
|
0.461 |
| 13 |
H |
0.374 |
0.421 |
|
0.428 |
| 14 |
C |
0.667 |
0.929 |
|
0.938 |
| 15 |
O |
-0.477 |
-0.660 |
|
-0.653 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.095 |
-3.731 |
0.364 |
3.750 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.074 |
-3.752 |
0.396 |
3.773 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.795 |
-6.415 |
0.945 |
| y |
-6.415 |
-39.730 |
-0.596 |
| z |
0.945 |
-0.596 |
-38.077 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.108 |
-6.415 |
0.945 |
| y |
-6.415 |
-3.793 |
-0.596 |
| z |
0.945 |
-0.596 |
-1.315 |
|
| Polar |
|---|
| 3z2-r2 | -2.630 |
|---|
| x2-y2 | 5.934 |
|---|
| xy | -6.415 |
|---|
| xz | 0.945 |
|---|
| yz | -0.596 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.434 |
-0.209 |
-0.031 |
| y |
-0.209 |
7.760 |
0.006 |
| z |
-0.031 |
0.006 |
5.896 |
<r2> (average value of r
2) Å
2
| <r2> |
216.838 |
| (<r2>)1/2 |
14.725 |