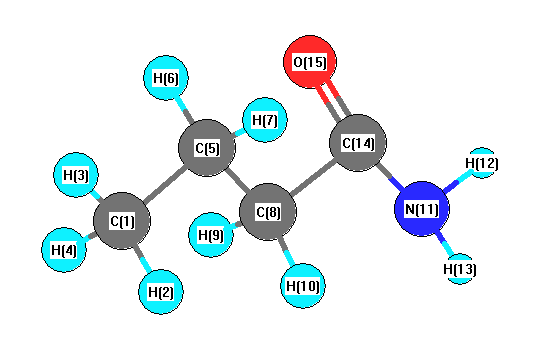
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3751 |
3611 |
24.47 |
|
|
|
| 2 |
A |
3616 |
3481 |
25.10 |
|
|
|
| 3 |
A |
3113 |
2997 |
53.70 |
|
|
|
| 4 |
A |
3107 |
2991 |
45.05 |
|
|
|
| 5 |
A |
3085 |
2970 |
9.80 |
|
|
|
| 6 |
A |
3061 |
2947 |
21.83 |
|
|
|
| 7 |
A |
3049 |
2935 |
27.68 |
|
|
|
| 8 |
A |
3030 |
2917 |
33.66 |
|
|
|
| 9 |
A |
3021 |
2908 |
14.20 |
|
|
|
| 10 |
A |
1803 |
1735 |
307.77 |
|
|
|
| 11 |
A |
1654 |
1593 |
116.91 |
|
|
|
| 12 |
A |
1519 |
1463 |
9.76 |
|
|
|
| 13 |
A |
1513 |
1456 |
9.93 |
|
|
|
| 14 |
A |
1503 |
1447 |
3.48 |
|
|
|
| 15 |
A |
1475 |
1420 |
9.98 |
|
|
|
| 16 |
A |
1424 |
1371 |
69.76 |
|
|
|
| 17 |
A |
1422 |
1369 |
10.01 |
|
|
|
| 18 |
A |
1368 |
1317 |
33.51 |
|
|
|
| 19 |
A |
1332 |
1282 |
34.66 |
|
|
|
| 20 |
A |
1275 |
1227 |
37.82 |
|
|
|
| 21 |
A |
1259 |
1212 |
6.60 |
|
|
|
| 22 |
A |
1145 |
1102 |
0.94 |
|
|
|
| 23 |
A |
1124 |
1083 |
2.33 |
|
|
|
| 24 |
A |
1087 |
1046 |
1.01 |
|
|
|
| 25 |
A |
1063 |
1023 |
2.56 |
|
|
|
| 26 |
A |
937 |
902 |
2.90 |
|
|
|
| 27 |
A |
892 |
858 |
1.08 |
|
|
|
| 28 |
A |
851 |
820 |
3.72 |
|
|
|
| 29 |
A |
752 |
724 |
7.63 |
|
|
|
| 30 |
A |
676 |
651 |
6.46 |
|
|
|
| 31 |
A |
618 |
595 |
14.45 |
|
|
|
| 32 |
A |
522 |
502 |
6.82 |
|
|
|
| 33 |
A |
426 |
410 |
2.26 |
|
|
|
| 34 |
A |
342 |
330 |
2.61 |
|
|
|
| 35 |
A |
246 |
237 |
1.67 |
|
|
|
| 36 |
A |
218 |
210 |
231.74 |
|
|
|
| 37 |
A |
185 |
178 |
7.97 |
|
|
|
| 38 |
A |
89 |
86 |
1.28 |
|
|
|
| 39 |
A |
36 |
35 |
3.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28793.6 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 27719.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.653 |
|
|
|
| 2 |
H |
0.217 |
|
|
|
| 3 |
H |
0.227 |
|
|
|
| 4 |
H |
0.213 |
|
|
|
| 5 |
C |
-0.438 |
|
|
|
| 6 |
H |
0.223 |
|
|
|
| 7 |
H |
0.245 |
|
|
|
| 8 |
C |
-0.561 |
|
|
|
| 9 |
H |
0.220 |
|
|
|
| 10 |
H |
0.245 |
|
|
|
| 11 |
N |
-0.810 |
|
|
|
| 12 |
H |
0.360 |
|
|
|
| 13 |
H |
0.359 |
|
|
|
| 14 |
C |
0.532 |
|
|
|
| 15 |
O |
-0.378 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.372 |
-3.583 |
-0.039 |
3.603 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.706 |
-6.371 |
-0.301 |
| y |
-6.371 |
-38.833 |
-0.160 |
| z |
-0.301 |
-0.160 |
-37.935 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.678 |
-6.371 |
-0.301 |
| y |
-6.371 |
-3.513 |
-0.160 |
| z |
-0.301 |
-0.160 |
-2.165 |
|
| Polar |
|---|
| 3z2-r2 | -4.331 |
|---|
| x2-y2 | 6.127 |
|---|
| xy | -6.371 |
|---|
| xz | -0.301 |
|---|
| yz | -0.160 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
216.955 |
| (<r2>)1/2 |
14.729 |