return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

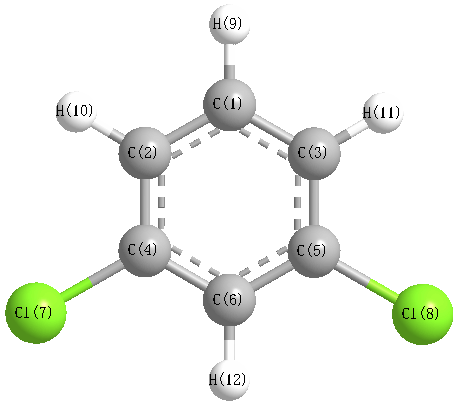
All results from a given calculation for C6H4Cl2 (1,3-dichlorobenzene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-1149.874184
Energy at 298.15K-1149.878066
HF Energy-1148.587482
Nuclear repulsion energy452.643786
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3246 3074 3.23      
2 A1 3245 3073 0.00      
3 A1 3221 3050 3.73      
4 A1 1630 1543 25.17      
5 A1 1453 1377 15.98      
6 A1 1167 1105 20.73      
7 A1 1109 1051 12.03      
8 A1 1007 953 5.43      
9 A1 678 642 1.34      
10 A1 409 388 1.92      
11 A1 199 188 0.18      
12 A2 832 788 0.00      
13 A2 502 476 0.00      
14 A2 199 188 0.00      
15 B1 874 827 0.26      
16 B1 806 763 2.94      
17 B1 740 701 83.86      
18 B1 423 400 5.62      
19 B1 401 380 0.20      
20 B1 166 157 0.39      
21 B2 3240 3069 0.64      
22 B2 1633 1547 76.26      
23 B2 1506 1426 68.08      
24 B2 1457 1380 2.17      
25 B2 1310 1240 2.48      
26 B2 1204 1140 0.77      
27 B2 1125 1066 23.84      
28 B2 814 771 72.49      
29 B2 437 414 1.91      
30 B2 370 350 0.51      

Unscaled Zero Point Vibrational Energy (zpe) 17699.6 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 16763.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.09418 0.02869 0.02199

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 2.077
C2 0.000 1.215 1.389
C3 0.000 -1.215 1.389
C4 0.000 1.199 -0.006
C5 0.000 -1.199 -0.006
C6 0.000 0.000 -0.720
Cl7 0.000 2.696 -0.881
Cl8 0.000 -2.696 -0.881
H9 0.000 0.000 3.163
H10 0.000 2.161 1.921
H11 0.000 -2.161 1.921
H12 0.000 0.000 -1.804

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C11.39611.39612.40362.40362.79654.00194.00191.08622.16642.16643.8811
C21.39612.42971.39552.78802.43362.71024.52172.15021.08493.41723.4165
C31.39612.42972.78801.39552.43364.52172.71022.15023.41721.08493.4165
C42.40361.39552.78802.39761.39491.73373.99173.38862.15363.87292.1609
C52.40362.78801.39552.39761.39493.99171.73373.38863.87292.15362.1609
C62.79652.43362.43361.39491.39492.70072.70073.88273.41153.41151.0846
Cl74.00192.71024.52171.73373.99172.70075.39194.86002.85175.60652.8497
Cl84.00194.52172.71023.99171.73372.70075.39194.86005.60652.85172.8497
H91.08622.15022.15023.38863.38863.88274.86004.86002.49262.49264.9673
H102.16641.08493.41722.15363.87293.41152.85175.60652.49264.32144.3060
H112.16643.41721.08493.87292.15363.41155.60652.85172.49264.32144.3060
H123.88113.41653.41652.16092.16091.08462.84972.84974.96734.30604.3060

picture of 1,3-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 118.860 C1 C2 H10 121.152
C1 C3 C5 118.860 C1 C3 H11 121.152
C2 C1 C3 120.962 C2 C1 H9 119.519
C2 C4 C6 121.410 C2 C4 Cl7 119.623
C3 C1 H9 119.519 C3 C5 C6 121.410
C3 C5 Cl8 119.623 C4 C2 H10 119.988
C4 C6 C5 118.498 C4 C6 H12 120.751
C5 C3 H11 119.988 C5 C6 H12 120.751
C6 C4 Cl7 118.966 C6 C5 Cl8 118.966
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability