Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -322.907432 |
| Energy at 298.15K | |
| HF Energy | -321.809116 |
| Nuclear repulsion energy | 234.565842 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3178 |
3010 |
24.02 |
|
|
|
| 2 |
A' |
3104 |
2939 |
28.33 |
|
|
|
| 3 |
A' |
3086 |
2923 |
35.18 |
|
|
|
| 4 |
A' |
3082 |
2919 |
8.27 |
|
|
|
| 5 |
A' |
1657 |
1570 |
124.19 |
|
|
|
| 6 |
A' |
1560 |
1477 |
18.85 |
|
|
|
| 7 |
A' |
1546 |
1464 |
3.76 |
|
|
|
| 8 |
A' |
1533 |
1452 |
2.27 |
|
|
|
| 9 |
A' |
1454 |
1377 |
12.39 |
|
|
|
| 10 |
A' |
1452 |
1375 |
18.44 |
|
|
|
| 11 |
A' |
1358 |
1286 |
3.90 |
|
|
|
| 12 |
A' |
1187 |
1124 |
5.58 |
|
|
|
| 13 |
A' |
1110 |
1052 |
34.12 |
|
|
|
| 14 |
A' |
1080 |
1023 |
8.88 |
|
|
|
| 15 |
A' |
949 |
899 |
97.73 |
|
|
|
| 16 |
A' |
855 |
810 |
495.92 |
|
|
|
| 17 |
A' |
672 |
636 |
76.40 |
|
|
|
| 18 |
A' |
398 |
377 |
0.11 |
|
|
|
| 19 |
A' |
358 |
340 |
3.36 |
|
|
|
| 20 |
A' |
159 |
150 |
0.38 |
|
|
|
| 21 |
A" |
3176 |
3008 |
60.34 |
|
|
|
| 22 |
A" |
3157 |
2990 |
3.20 |
|
|
|
| 23 |
A" |
3140 |
2974 |
6.06 |
|
|
|
| 24 |
A" |
1545 |
1463 |
10.85 |
|
|
|
| 25 |
A" |
1354 |
1282 |
0.03 |
|
|
|
| 26 |
A" |
1304 |
1235 |
0.36 |
|
|
|
| 27 |
A" |
1216 |
1152 |
0.93 |
|
|
|
| 28 |
A" |
917 |
869 |
1.53 |
|
|
|
| 29 |
A" |
776 |
735 |
2.24 |
|
|
|
| 30 |
A" |
244 |
231 |
0.03 |
|
|
|
| 31 |
A" |
214 |
203 |
1.02 |
|
|
|
| 32 |
A" |
103 |
98 |
1.36 |
|
|
|
| 33 |
A" |
47i |
45i |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23437.9 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 22198.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.914 |
2.201 |
0.000 |
| C2 |
-1.506 |
0.730 |
0.000 |
| C3 |
0.000 |
0.576 |
0.000 |
| O4 |
0.282 |
-0.835 |
0.000 |
| N5 |
1.674 |
-1.026 |
0.000 |
| O6 |
1.943 |
-2.179 |
0.000 |
| H7 |
-3.000 |
2.305 |
0.000 |
| H8 |
-1.531 |
2.718 |
0.884 |
| H9 |
-1.531 |
2.718 |
-0.884 |
| H10 |
-1.910 |
0.222 |
-0.880 |
| H11 |
-1.910 |
0.222 |
0.880 |
| H12 |
0.442 |
1.034 |
0.890 |
| H13 |
0.442 |
1.034 |
-0.890 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5264 | 2.5109 | 3.7467 | 4.8249 | 5.8362 | 1.0915 | 1.0929 | 1.0929 | 2.1661 | 2.1661 | 2.7761 | 2.7761 |
C2 | 1.5264 | | 1.5137 | 2.3760 | 3.6319 | 4.5120 | 2.1708 | 2.1754 | 2.1754 | 1.0936 | 1.0936 | 2.1633 | 2.1633 | C3 | 2.5109 | 1.5137 | | 1.4382 | 2.3162 | 3.3711 | 3.4629 | 2.7774 | 2.7774 | 2.1325 | 2.1325 | 1.0944 | 1.0944 | O4 | 3.7467 | 2.3760 | 1.4382 | | 1.4044 | 2.1368 | 4.5420 | 4.0852 | 4.0852 | 2.5875 | 2.5875 | 2.0756 | 2.0756 | N5 | 4.8249 | 3.6319 | 2.3162 | 1.4044 | | 1.1848 | 5.7389 | 5.0063 | 5.0063 | 3.8950 | 3.8950 | 2.5589 | 2.5589 | O6 | 5.8362 | 4.5120 | 3.3711 | 2.1368 | 1.1848 | | 6.6739 | 6.0688 | 6.0688 | 4.6241 | 4.6241 | 3.6560 | 3.6560 | H7 | 1.0915 | 2.1708 | 3.4629 | 4.5420 | 5.7389 | 6.6739 | | 1.7637 | 1.7637 | 2.5103 | 2.5103 | 3.7763 | 3.7763 | H8 | 1.0929 | 2.1754 | 2.7774 | 4.0852 | 5.0063 | 6.0688 | 1.7637 | | 1.7673 | 3.0797 | 2.5250 | 2.5945 | 3.1428 | H9 | 1.0929 | 2.1754 | 2.7774 | 4.0852 | 5.0063 | 6.0688 | 1.7637 | 1.7673 | | 2.5250 | 3.0797 | 3.1428 | 2.5945 | H10 | 2.1661 | 1.0936 | 2.1325 | 2.5875 | 3.8950 | 4.6241 | 2.5103 | 3.0797 | 2.5250 | | 1.7591 | 3.0537 | 2.4888 | H11 | 2.1661 | 1.0936 | 2.1325 | 2.5875 | 3.8950 | 4.6241 | 2.5103 | 2.5250 | 3.0797 | 1.7591 | | 2.4888 | 3.0537 | H12 | 2.7761 | 2.1633 | 1.0944 | 2.0756 | 2.5589 | 3.6560 | 3.7763 | 2.5945 | 3.1428 | 3.0537 | 2.4888 | | 1.7801 | H13 | 2.7761 | 2.1633 | 1.0944 | 2.0756 | 2.5589 | 3.6560 | 3.7763 | 3.1428 | 2.5945 | 2.4888 | 3.0537 | 1.7801 | |
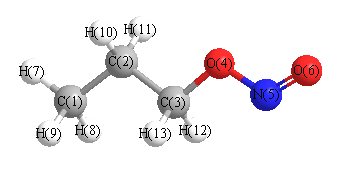 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.366 |
|
C1 |
C2 |
H10 |
110.453 |
| C1 |
C2 |
H11 |
110.453 |
|
C2 |
C1 |
H7 |
110.951 |
| C2 |
C1 |
H8 |
111.227 |
|
C2 |
C1 |
H9 |
111.227 |
| C2 |
C3 |
O4 |
107.178 |
|
C2 |
C3 |
H12 |
111.066 |
| C2 |
C3 |
H13 |
111.066 |
|
C3 |
C2 |
H10 |
108.682 |
| C3 |
C2 |
H11 |
108.682 |
|
C3 |
O4 |
N5 |
109.133 |
| O4 |
C3 |
H12 |
109.328 |
|
O4 |
C3 |
H13 |
109.328 |
| O4 |
N5 |
O6 |
110.954 |
|
H7 |
C1 |
H8 |
107.685 |
| H7 |
C1 |
H9 |
107.685 |
|
H8 |
C1 |
H9 |
107.902 |
| H10 |
C2 |
H11 |
107.082 |
|
H12 |
C3 |
H13 |
108.839 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -322.909763 |
| Energy at 298.15K | -322.918868 |
| HF Energy | -321.808278 |
| Nuclear repulsion energy | 242.126017 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3185 |
3017 |
23.70 |
|
|
|
| 2 |
A |
3172 |
3004 |
32.65 |
|
|
|
| 3 |
A |
3161 |
2994 |
20.29 |
|
|
|
| 4 |
A |
3139 |
2973 |
11.82 |
|
|
|
| 5 |
A |
3093 |
2930 |
35.78 |
|
|
|
| 6 |
A |
3086 |
2923 |
14.86 |
|
|
|
| 7 |
A |
3086 |
2922 |
17.63 |
|
|
|
| 8 |
A |
1666 |
1578 |
147.69 |
|
|
|
| 9 |
A |
1550 |
1468 |
13.82 |
|
|
|
| 10 |
A |
1542 |
1461 |
9.59 |
|
|
|
| 11 |
A |
1530 |
1449 |
3.23 |
|
|
|
| 12 |
A |
1521 |
1440 |
3.30 |
|
|
|
| 13 |
A |
1455 |
1378 |
8.27 |
|
|
|
| 14 |
A |
1432 |
1356 |
12.31 |
|
|
|
| 15 |
A |
1408 |
1334 |
0.89 |
|
|
|
| 16 |
A |
1333 |
1262 |
4.92 |
|
|
|
| 17 |
A |
1321 |
1251 |
9.52 |
|
|
|
| 18 |
A |
1214 |
1150 |
11.90 |
|
|
|
| 19 |
A |
1143 |
1083 |
8.73 |
|
|
|
| 20 |
A |
1121 |
1062 |
33.38 |
|
|
|
| 21 |
A |
1029 |
975 |
11.81 |
|
|
|
| 22 |
A |
949 |
899 |
68.90 |
|
|
|
| 23 |
A |
905 |
857 |
1.81 |
|
|
|
| 24 |
A |
840 |
796 |
358.56 |
|
|
|
| 25 |
A |
792 |
750 |
92.89 |
|
|
|
| 26 |
A |
615 |
583 |
70.78 |
|
|
|
| 27 |
A |
463 |
439 |
17.33 |
|
|
|
| 28 |
A |
396 |
375 |
3.41 |
|
|
|
| 29 |
A |
285 |
270 |
1.83 |
|
|
|
| 30 |
A |
259 |
246 |
1.11 |
|
|
|
| 31 |
A |
186 |
176 |
0.36 |
|
|
|
| 32 |
A |
114 |
108 |
0.96 |
|
|
|
| 33 |
A |
57 |
54 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23524.3 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 22279.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.162 |
-0.928 |
0.182 |
| C2 |
-1.757 |
0.458 |
-0.308 |
| C3 |
-0.409 |
0.898 |
0.221 |
| O4 |
0.561 |
-0.030 |
-0.297 |
| N5 |
1.780 |
0.211 |
0.313 |
| O6 |
2.570 |
-0.553 |
-0.139 |
| H7 |
-3.143 |
-1.208 |
-0.206 |
| H8 |
-2.212 |
-0.956 |
1.274 |
| H9 |
-1.440 |
-1.679 |
-0.142 |
| H10 |
-1.728 |
0.482 |
-1.402 |
| H11 |
-2.493 |
1.205 |
0.008 |
| H12 |
-0.149 |
1.906 |
-0.116 |
| H13 |
-0.379 |
0.866 |
1.314 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5255 | 2.5317 | 2.9067 | 4.1045 | 4.7577 | 1.0920 | 1.0933 | 1.0904 | 2.1646 | 2.1661 | 3.4886 | 2.7709 |
C2 | 1.5255 | | 1.5135 | 2.3685 | 3.5990 | 4.4471 | 2.1703 | 2.1703 | 2.1668 | 1.0941 | 1.0954 | 2.1718 | 2.1672 | C3 | 2.5317 | 1.5135 | | 1.4388 | 2.2955 | 3.3335 | 3.4780 | 2.7926 | 2.7994 | 2.1322 | 2.1175 | 1.0935 | 1.0941 | O4 | 2.9067 | 2.3685 | 1.4388 | | 1.3841 | 2.0827 | 3.8879 | 3.3192 | 2.5975 | 2.5923 | 3.3083 | 2.0694 | 2.0695 | N5 | 4.1045 | 3.5990 | 2.2955 | 1.3841 | | 1.1891 | 5.1493 | 4.2685 | 3.7610 | 3.9135 | 4.3973 | 2.6028 | 2.4680 | O6 | 4.7577 | 4.4471 | 3.3335 | 2.0827 | 1.1891 | | 5.7513 | 5.0034 | 4.1655 | 4.5980 | 5.3621 | 3.6662 | 3.5813 | H7 | 1.0920 | 2.1703 | 3.4780 | 3.8879 | 5.1493 | 5.7513 | | 1.7664 | 1.7677 | 2.5080 | 2.5088 | 4.3209 | 3.7754 | H8 | 1.0933 | 2.1703 | 2.7926 | 3.3192 | 4.2685 | 5.0034 | 1.7664 | | 1.7679 | 3.0758 | 2.5201 | 3.7916 | 2.5848 | H9 | 1.0904 | 2.1668 | 2.7994 | 2.5975 | 3.7610 | 4.1655 | 1.7677 | 1.7679 | | 2.5175 | 3.0739 | 3.8099 | 3.1183 | H10 | 2.1646 | 1.0941 | 2.1322 | 2.5923 | 3.9135 | 4.5980 | 2.5080 | 3.0758 | 2.5175 | | 1.7594 | 2.4846 | 3.0566 | H11 | 2.1661 | 1.0954 | 2.1175 | 3.3083 | 4.3973 | 5.3621 | 2.5088 | 2.5201 | 3.0739 | 1.7594 | | 2.4496 | 2.5080 | H12 | 3.4886 | 2.1718 | 1.0935 | 2.0694 | 2.6028 | 3.6662 | 4.3209 | 3.7916 | 3.8099 | 2.4846 | 2.4496 | | 1.7829 | H13 | 2.7709 | 2.1672 | 1.0941 | 2.0695 | 2.4680 | 3.5813 | 3.7754 | 2.5848 | 3.1183 | 3.0566 | 2.5080 | 1.7829 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.832 |
|
C1 |
C2 |
H10 |
110.364 |
| C1 |
C2 |
H11 |
110.403 |
|
C2 |
C1 |
H7 |
110.941 |
| C2 |
C1 |
H8 |
110.863 |
|
C2 |
C1 |
H9 |
110.757 |
| C2 |
C3 |
O4 |
106.666 |
|
C2 |
C3 |
H12 |
111.819 |
| C2 |
C3 |
H13 |
111.412 |
|
C3 |
C2 |
H10 |
108.645 |
| C3 |
C2 |
H11 |
107.427 |
|
C3 |
O4 |
N5 |
108.802 |
| O4 |
C3 |
H12 |
108.847 |
|
O4 |
C3 |
H13 |
108.825 |
| O4 |
N5 |
O6 |
107.832 |
|
H7 |
C1 |
H8 |
107.862 |
| H7 |
C1 |
H9 |
108.190 |
|
H8 |
C1 |
H9 |
108.110 |
| H10 |
C2 |
H11 |
106.951 |
|
H12 |
C3 |
H13 |
109.174 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
