Jump to
S1C2
Vibrational Frequencies calculated at PBE1PBE/6-311G*
Geometric Data calculated at PBE1PBE/6-311G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -323.350314 |
| Energy at 298.15K | -323.359370 |
| HF Energy | -323.350314 |
| Nuclear repulsion energy | 242.608318 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3140 |
3010 |
31.94 |
|
|
|
| 2 |
A |
3131 |
3001 |
46.60 |
|
|
|
| 3 |
A |
3122 |
2993 |
24.29 |
|
|
|
| 4 |
A |
3095 |
2967 |
15.69 |
|
|
|
| 5 |
A |
3064 |
2937 |
29.41 |
|
|
|
| 6 |
A |
3056 |
2930 |
31.65 |
|
|
|
| 7 |
A |
3056 |
2929 |
10.59 |
|
|
|
| 8 |
A |
1816 |
1741 |
261.59 |
|
|
|
| 9 |
A |
1522 |
1458 |
11.56 |
|
|
|
| 10 |
A |
1515 |
1452 |
11.87 |
|
|
|
| 11 |
A |
1499 |
1437 |
3.02 |
|
|
|
| 12 |
A |
1489 |
1427 |
3.68 |
|
|
|
| 13 |
A |
1430 |
1371 |
8.95 |
|
|
|
| 14 |
A |
1414 |
1356 |
9.59 |
|
|
|
| 15 |
A |
1388 |
1331 |
1.04 |
|
|
|
| 16 |
A |
1320 |
1265 |
6.07 |
|
|
|
| 17 |
A |
1307 |
1253 |
9.06 |
|
|
|
| 18 |
A |
1197 |
1148 |
9.37 |
|
|
|
| 19 |
A |
1130 |
1084 |
13.21 |
|
|
|
| 20 |
A |
1110 |
1064 |
26.70 |
|
|
|
| 21 |
A |
1025 |
982 |
7.30 |
|
|
|
| 22 |
A |
941 |
902 |
121.28 |
|
|
|
| 23 |
A |
893 |
856 |
1.85 |
|
|
|
| 24 |
A |
863 |
827 |
347.13 |
|
|
|
| 25 |
A |
787 |
754 |
21.28 |
|
|
|
| 26 |
A |
632 |
606 |
31.52 |
|
|
|
| 27 |
A |
460 |
441 |
6.42 |
|
|
|
| 28 |
A |
388 |
372 |
2.71 |
|
|
|
| 29 |
A |
284 |
272 |
0.84 |
|
|
|
| 30 |
A |
246 |
236 |
0.81 |
|
|
|
| 31 |
A |
184 |
177 |
0.51 |
|
|
|
| 32 |
A |
109 |
104 |
0.98 |
|
|
|
| 33 |
A |
53 |
51 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23334.3 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 22366.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.179 |
-0.918 |
0.181 |
| C2 |
-1.751 |
0.459 |
-0.311 |
| C3 |
-0.403 |
0.892 |
0.222 |
| O4 |
0.573 |
-0.028 |
-0.288 |
| N5 |
1.788 |
0.204 |
0.310 |
| O6 |
2.561 |
-0.555 |
-0.142 |
| H7 |
-3.160 |
-1.188 |
-0.218 |
| H8 |
-2.245 |
-0.944 |
1.273 |
| H9 |
-1.466 |
-1.683 |
-0.132 |
| H10 |
-1.718 |
0.478 |
-1.405 |
| H11 |
-2.482 |
1.216 |
-0.006 |
| H12 |
-0.139 |
1.898 |
-0.112 |
| H13 |
-0.373 |
0.862 |
1.316 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5231 | 2.5356 | 2.9303 | 4.1253 | 4.7652 | 1.0926 | 1.0944 | 1.0916 | 2.1624 | 2.1630 | 3.4900 | 2.7781 |
C2 | 1.5231 | | 1.5125 | 2.3747 | 3.6023 | 4.4328 | 2.1692 | 2.1726 | 2.1678 | 1.0949 | 1.0959 | 2.1704 | 2.1695 | C3 | 2.5356 | 1.5125 | | 1.4354 | 2.2985 | 3.3185 | 3.4810 | 2.8044 | 2.8075 | 2.1321 | 2.1164 | 1.0935 | 1.0943 | O4 | 2.9303 | 2.3747 | 1.4354 | | 1.3744 | 2.0616 | 3.9099 | 3.3489 | 2.6305 | 2.5984 | 3.3111 | 2.0616 | 2.0639 | N5 | 4.1253 | 3.6023 | 2.2985 | 1.3744 | | 1.1740 | 5.1677 | 4.3024 | 3.7879 | 3.9126 | 4.4000 | 2.6003 | 2.4725 | O6 | 4.7652 | 4.4328 | 3.3185 | 2.0616 | 1.1740 | | 5.7568 | 5.0248 | 4.1822 | 4.5792 | 5.3469 | 3.6480 | 3.5692 | H7 | 1.0926 | 2.1692 | 3.4810 | 3.9099 | 5.1677 | 5.7568 | | 1.7661 | 1.7672 | 2.5032 | 2.5063 | 4.3202 | 3.7843 | H8 | 1.0944 | 2.1726 | 2.8044 | 3.3489 | 4.3024 | 5.0248 | 1.7661 | | 1.7676 | 3.0776 | 2.5213 | 3.7993 | 2.6013 | H9 | 1.0916 | 2.1678 | 2.8075 | 2.6305 | 3.7879 | 4.1822 | 1.7672 | 1.7676 | | 2.5208 | 3.0741 | 3.8192 | 3.1248 | H10 | 2.1624 | 1.0949 | 2.1321 | 2.5984 | 3.9126 | 4.5792 | 2.5032 | 3.0776 | 2.5208 | | 1.7569 | 2.4859 | 3.0589 | H11 | 2.1630 | 1.0959 | 2.1164 | 3.3111 | 4.4000 | 5.3469 | 2.5063 | 2.5213 | 3.0741 | 1.7569 | | 2.4432 | 2.5141 | H12 | 3.4900 | 2.1704 | 1.0935 | 2.0616 | 2.6003 | 3.6480 | 4.3202 | 3.7993 | 3.8192 | 2.4859 | 2.4432 | | 1.7803 | H13 | 2.7781 | 2.1695 | 1.0943 | 2.0639 | 2.4725 | 3.5692 | 3.7843 | 2.6013 | 3.1248 | 3.0589 | 2.5141 | 1.7803 | |
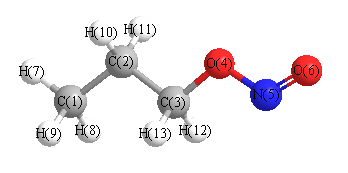 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.297 |
|
C1 |
C2 |
H10 |
110.310 |
| C1 |
C2 |
H11 |
110.293 |
|
C2 |
C1 |
H7 |
110.988 |
| C2 |
C1 |
H8 |
111.147 |
|
C2 |
C1 |
H9 |
110.933 |
| C2 |
C3 |
O4 |
107.303 |
|
C2 |
C3 |
H12 |
111.784 |
| C2 |
C3 |
H13 |
111.661 |
|
C3 |
C2 |
H10 |
108.660 |
| C3 |
C2 |
H11 |
107.384 |
|
C3 |
O4 |
N5 |
109.753 |
| O4 |
C3 |
H12 |
108.461 |
|
O4 |
C3 |
H13 |
108.595 |
| O4 |
N5 |
O6 |
107.734 |
|
H7 |
C1 |
H8 |
107.711 |
| H7 |
C1 |
H9 |
108.004 |
|
H8 |
C1 |
H9 |
107.914 |
| H10 |
C2 |
H11 |
106.636 |
|
H12 |
C3 |
H13 |
108.930 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.674 |
|
|
|
| 2 |
C |
-0.455 |
|
|
|
| 3 |
C |
-0.254 |
|
|
|
| 4 |
O |
-0.258 |
|
|
|
| 5 |
N |
0.205 |
|
|
|
| 6 |
O |
-0.184 |
|
|
|
| 7 |
H |
0.234 |
|
|
|
| 8 |
H |
0.225 |
|
|
|
| 9 |
H |
0.236 |
|
|
|
| 10 |
H |
0.236 |
|
|
|
| 11 |
H |
0.226 |
|
|
|
| 12 |
H |
0.238 |
|
|
|
| 13 |
H |
0.225 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.360 |
-0.899 |
0.751 |
2.634 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.675 |
0.492 |
-0.648 |
| y |
0.492 |
-34.912 |
-0.809 |
| z |
-0.648 |
-0.809 |
-37.073 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.683 |
0.492 |
-0.648 |
| y |
0.492 |
2.962 |
-0.809 |
| z |
-0.648 |
-0.809 |
-0.279 |
|
| Polar |
|---|
| 3z2-r2 | -0.559 |
|---|
| x2-y2 | -3.763 |
|---|
| xy | 0.492 |
|---|
| xz | -0.648 |
|---|
| yz | -0.809 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.148 |
-1.098 |
-0.007 |
| y |
-1.098 |
6.859 |
-0.038 |
| z |
-0.007 |
-0.038 |
5.988 |
<r2> (average value of r
2) Å
2
| <r2> |
193.269 |
| (<r2>)1/2 |
13.902 |
