return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6S (Thiophene, 2-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-591.406527
Energy at 298.15K-591.412547
HF Energy-590.381789
Nuclear repulsion energy272.955190
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3277 3103 1.47      
2 A 3247 3075 4.67      
3 A 3230 3059 6.57      
4 A 3175 3007 10.67      
5 A 3078 2915 26.20      
6 A 1579 1495 1.49      
7 A 1531 1450 7.55      
8 A 1486 1407 8.12      
9 A 1452 1375 0.87      
10 A 1403 1329 0.54      
11 A 1275 1207 15.22      
12 A 1219 1155 12.14      
13 A 1111 1053 3.95      
14 A 1085 1028 4.31      
15 A 1008 955 1.84      
16 A 887 840 14.93      
17 A 771 730 0.10      
18 A 695 658 1.39      
19 A 560 531 1.00      
20 A 307 290 0.79      
21 A 3155 2989 12.57      
22 A 1516 1436 10.79      
23 A 1069 1013 0.24      
24 A 813 770 0.06      
25 A 774 733 21.78      
26 A 665 630 81.01      
27 A 526 498 2.88      
28 A 455 431 6.32      
29 A 221 209 3.60      
30 A 68 65 0.19      

Unscaled Zero Point Vibrational Energy (zpe) 20818.7 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 19717.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.17609 0.10403 0.06620

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -2.373 -1.581 0.000
C2 -1.431 -1.045 0.000
H3 -2.151 1.056 0.000
C4 -1.310 0.370 0.000
C5 0.000 0.807 0.000
H6 0.006 -2.733 0.000
C7 -0.205 -1.672 0.000
S8 1.081 -0.536 0.000
H9 -0.345 2.906 0.000
H10 1.109 2.433 0.884
H11 1.109 2.433 -0.884
C12 0.503 2.218 0.000

Atom - Atom Distances (Å)
  H1 C2 H3 C4 C5 H6 C7 S8 H9 H10 H11 C12
H11.08442.64672.22233.36642.64322.16993.60834.92425.38745.38744.7649
C21.08442.22051.41972.33942.21721.37712.56244.09684.39634.39633.7923
H32.64672.22051.08552.16544.36013.35093.60292.58523.64803.64802.8970
C42.22231.41971.08551.38043.37062.32172.55662.71303.29983.29982.5883
C53.36642.33942.16541.38043.53972.48691.72402.12752.15822.15821.4983
H62.64322.21724.36013.37063.53971.08202.44545.64995.35635.35634.9759
C72.16991.37713.35092.32172.48691.08201.71524.57994.40014.40013.9537
S83.60832.56243.60292.55661.72402.44541.71523.72613.09863.09862.8145
H94.92424.09682.58522.71302.12755.64994.57993.72611.76621.76621.0918
H105.38744.39633.64803.29982.15825.35634.40013.09861.76621.76711.0932
H115.38744.39633.64803.29982.15825.35634.40013.09861.76621.76711.0932
C124.76493.79232.89702.58831.49834.97593.95372.81451.09181.09321.0932

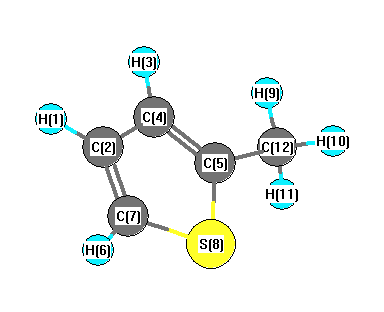
picture of Thiophene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 124.571 H1 C2 C7 123.217
C2 C4 H3 124.292 C2 C4 C5 113.326
C2 C7 H6 128.352 C2 C7 S8 111.448
H3 C4 C5 122.382 C4 C2 C7 112.212
C4 C5 S8 110.394 C4 C5 C12 128.041
C5 S8 C7 92.621 C5 C12 H9 109.448
C5 C12 H10 111.826 C5 C12 H11 111.826
H6 C7 S8 120.200 S8 C5 C12 121.565
H9 C12 H10 107.867 H9 C12 H11 107.867
H10 C12 H11 107.847
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability