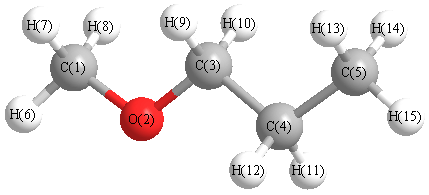
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3129 |
3012 |
37.20 |
|
|
|
| 2 |
A' |
3112 |
2996 |
39.59 |
|
|
|
| 3 |
A' |
3047 |
2933 |
32.68 |
|
|
|
| 4 |
A' |
3035 |
2922 |
30.67 |
|
|
|
| 5 |
A' |
2976 |
2865 |
90.04 |
|
|
|
| 6 |
A' |
2949 |
2839 |
44.27 |
|
|
|
| 7 |
A' |
1541 |
1484 |
3.85 |
|
|
|
| 8 |
A' |
1521 |
1464 |
13.03 |
|
|
|
| 9 |
A' |
1516 |
1459 |
5.25 |
|
|
|
| 10 |
A' |
1502 |
1446 |
1.96 |
|
|
|
| 11 |
A' |
1490 |
1435 |
0.02 |
|
|
|
| 12 |
A' |
1438 |
1384 |
20.71 |
|
|
|
| 13 |
A' |
1424 |
1370 |
3.34 |
|
|
|
| 14 |
A' |
1340 |
1290 |
1.98 |
|
|
|
| 15 |
A' |
1242 |
1195 |
40.24 |
|
|
|
| 16 |
A' |
1181 |
1137 |
144.41 |
|
|
|
| 17 |
A' |
1137 |
1095 |
15.11 |
|
|
|
| 18 |
A' |
1069 |
1029 |
3.36 |
|
|
|
| 19 |
A' |
997 |
960 |
26.18 |
|
|
|
| 20 |
A' |
916 |
881 |
7.00 |
|
|
|
| 21 |
A' |
440 |
423 |
0.57 |
|
|
|
| 22 |
A' |
409 |
394 |
3.04 |
|
|
|
| 23 |
A' |
193 |
186 |
1.44 |
|
|
|
| 24 |
A" |
3106 |
2990 |
75.96 |
|
|
|
| 25 |
A" |
3079 |
2964 |
2.83 |
|
|
|
| 26 |
A" |
3020 |
2907 |
76.95 |
|
|
|
| 27 |
A" |
2976 |
2865 |
73.54 |
|
|
|
| 28 |
A" |
1516 |
1460 |
9.70 |
|
|
|
| 29 |
A" |
1499 |
1443 |
8.53 |
|
|
|
| 30 |
A" |
1327 |
1278 |
0.36 |
|
|
|
| 31 |
A" |
1280 |
1232 |
2.31 |
|
|
|
| 32 |
A" |
1205 |
1160 |
5.32 |
|
|
|
| 33 |
A" |
1178 |
1134 |
0.06 |
|
|
|
| 34 |
A" |
906 |
872 |
2.32 |
|
|
|
| 35 |
A" |
767 |
739 |
1.96 |
|
|
|
| 36 |
A" |
245 |
235 |
1.08 |
|
|
|
| 37 |
A" |
236 |
227 |
2.78 |
|
|
|
| 38 |
A" |
112 |
108 |
2.70 |
|
|
|
| 39 |
A" |
104 |
101 |
0.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30079.1 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 28957.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.452 |
|
|
|
| 2 |
O |
-0.330 |
|
|
|
| 3 |
C |
-0.191 |
|
|
|
| 4 |
C |
-0.462 |
|
|
|
| 5 |
C |
-0.662 |
|
|
|
| 6 |
H |
0.222 |
|
|
|
| 7 |
H |
0.195 |
|
|
|
| 8 |
H |
0.195 |
|
|
|
| 9 |
H |
0.187 |
|
|
|
| 10 |
H |
0.187 |
|
|
|
| 11 |
H |
0.224 |
|
|
|
| 12 |
H |
0.224 |
|
|
|
| 13 |
H |
0.218 |
|
|
|
| 14 |
H |
0.218 |
|
|
|
| 15 |
H |
0.228 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.200 |
1.086 |
0.000 |
1.105 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.244 |
-2.189 |
0.000 |
| y |
-2.189 |
-33.587 |
0.000 |
| z |
0.000 |
0.000 |
-33.057 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.078 |
-2.189 |
0.000 |
| y |
-2.189 |
-1.436 |
0.000 |
| z |
0.000 |
0.000 |
-0.641 |
|
| Polar |
|---|
| 3z2-r2 | -1.283 |
|---|
| x2-y2 | 2.343 |
|---|
| xy | -2.189 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.960 |
-0.405 |
0.000 |
| y |
-0.405 |
7.040 |
0.000 |
| z |
0.000 |
0.000 |
6.743 |
<r2> (average value of r
2) Å
2
| <r2> |
180.567 |
| (<r2>)1/2 |
13.438 |