return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (2,3-dimethylbut-1-ene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-235.159032
Energy at 298.15K 
HF Energy-234.217380
Nuclear repulsion energy244.073355
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3267 3094 17.78      
2 A' 3174 3006 37.44      
3 A' 3172 3004 2.60      
4 A' 3168 3000 20.04      
5 A' 3161 2994 62.00      
6 A' 3078 2915 1.79      
7 A' 3076 2914 34.61      
8 A' 3072 2909 13.95      
9 A' 1710 1620 8.92      
10 A' 1549 1468 9.38      
11 A' 1547 1465 12.15      
12 A' 1531 1450 13.86      
13 A' 1474 1396 2.07      
14 A' 1452 1375 9.01      
15 A' 1442 1366 9.73      
16 A' 1364 1291 1.81      
17 A' 1335 1264 0.17      
18 A' 1217 1152 1.92      
19 A' 1127 1068 11.02      
20 A' 1023 969 1.61      
21 A' 993 940 0.44      
22 A' 929 880 1.63      
23 A' 751 711 1.11      
24 A' 529 501 1.16      
25 A' 447 423 0.64      
26 A' 337 319 0.18      
27 A' 289 274 0.22      
28 A' 268 254 0.25      
29 A" 3167 3000 8.76      
30 A" 3159 2992 8.03      
31 A" 3149 2983 18.42      
32 A" 3071 2909 23.80      
33 A" 1538 1457 0.23      
34 A" 1524 1444 3.60      
35 A" 1515 1435 11.51      
36 A" 1429 1354 11.41      
37 A" 1367 1294 1.02      
38 A" 1156 1095 2.93      
39 A" 1078 1021 0.00      
40 A" 989 936 0.18      
41 A" 944 894 0.92      
42 A" 870 824 47.21      
43 A" 746 706 0.73      
44 A" 551 522 8.93      
45 A" 278 263 0.05      
46 A" 208 197 0.55      
47 A" 164 156 0.22      
48 A" 23i 22i 0.14      

Unscaled Zero Point Vibrational Energy (zpe) 36679.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 34739.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.14797 0.08376 0.08274

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.756 -1.062 0.000
C2 -0.475 -0.663 0.000
C3 -0.121 0.808 0.000
C4 0.655 -1.659 0.000
C5 0.655 1.204 1.259
C6 0.655 1.204 -1.259
H7 -2.027 -2.114 0.000
H8 -2.572 -0.345 0.000
H9 -1.066 1.365 0.000
H10 0.272 -2.682 0.000
H11 1.294 -1.540 -0.880
H12 1.294 -1.540 0.880
H13 0.796 2.288 1.296
H14 0.120 0.899 2.162
H15 1.647 0.744 1.276
H16 0.796 2.288 -1.296
H17 0.120 0.899 -2.162
H18 1.647 0.744 -1.276

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.34252.48412.48453.54073.54071.08601.08592.52302.59583.21033.21034.40643.47014.05874.40643.47014.0587
C21.34251.51271.50622.51962.51962.12492.12122.11252.15232.16092.16093.46462.73292.84773.46462.73292.8477
C32.48411.51272.58621.53141.53143.48842.70831.09733.51152.87852.87852.17072.17762.18122.17072.17762.1812
C42.48451.50622.58623.12813.12812.72073.48443.47971.09191.09381.09384.15733.39192.89634.15733.39192.8963
C53.54072.51961.53143.12812.51874.44843.79442.13894.10273.53732.84241.09391.09301.09322.77943.47662.7611
C63.54072.51961.53143.12812.51874.44843.79442.13894.10272.84243.53732.77943.47662.76111.09391.09301.0932
H71.08602.12493.48842.72074.44844.44841.85083.60882.36843.48333.48335.38754.28504.82655.38754.28504.8265
H81.08592.12122.70833.48443.79443.79441.85082.27803.68084.14054.14054.46673.66994.53994.46673.66994.5399
H92.52302.11251.09733.47972.13892.13893.60882.27804.26213.84443.84442.44932.51003.06182.44932.51003.0618
H102.59582.15233.51151.09194.10274.10272.36843.68084.26211.76711.76715.16294.18553.90585.16294.18553.9058
H113.21032.16092.87851.09383.53732.84243.48334.14053.84441.76711.76004.43134.07163.16063.88262.99452.3446
H123.21032.16092.87851.09382.84243.53733.48334.14053.84441.76711.76003.88262.99452.34464.43134.07163.1606
H134.40643.46462.17074.15731.09392.77945.38754.46672.44935.16294.43133.88261.77131.76332.59213.78773.1186
H143.47012.73292.17763.39191.09303.47664.28503.66992.51004.18554.07162.99451.77131.77193.78774.32463.7653
H154.05872.84772.18122.89631.09322.76114.82654.53993.06183.90583.16062.34461.76331.77193.11863.76532.5524
H164.40643.46462.17074.15732.77941.09395.38754.46672.44935.16293.88264.43132.59213.78773.11861.77131.7633
H173.47012.73292.17763.39193.47661.09304.28503.66992.51004.18552.99454.07163.78774.32463.76531.77131.7719
H184.05872.84772.18122.89632.76111.09324.82654.53993.06183.90582.34463.16063.11863.76532.55241.76331.7719

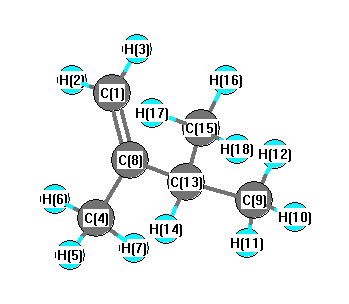
picture of 2,3-dimethylbut-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.804 C1 C2 C4 121.310
C2 C1 H7 121.726 C2 C1 H8 121.378
C2 C3 C5 111.722 C2 C3 C6 111.722
C2 C3 H9 106.991 C2 C4 H10 110.862
C2 C4 H11 111.438 C2 C4 H12 111.438
C3 C2 C4 117.886 C3 C5 H13 110.436
C3 C5 H14 111.040 C3 C5 H15 111.313
C3 C6 H16 110.436 C3 C6 H17 111.040
C3 C6 H18 111.313 C5 C3 C6 110.638
C5 C3 H9 107.769 C6 C3 H9 107.769
H7 C1 H8 116.896 H10 C4 H11 107.898
H10 C4 H12 107.898 H11 C4 H12 107.125
H13 C5 H14 108.177 H13 C5 H15 107.458
H14 C5 H15 108.284 H16 C6 H17 108.177
H16 C6 H18 107.458 H17 C6 H18 108.284
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability