return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (2,3-dimethylbut-1-ene)

using model chemistry: B2PLYP=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at B2PLYP=FULL/6-311G*
 hartrees
Energy at 0K-235.627906
Energy at 298.15K-235.640230
HF Energy-235.320137
Nuclear repulsion energy243.652335
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3234 3234 25.00      
2 A' 3152 3152 17.69      
3 A' 3139 3139 30.14      
4 A' 3131 3131 34.43      
5 A' 3125 3125 86.60      
6 A' 3059 3059 26.89      
7 A' 3053 3053 7.46      
8 A' 3051 3051 25.34      
9 A' 1711 1711 14.76      
10 A' 1546 1546 11.55      
11 A' 1541 1541 6.60      
12 A' 1528 1528 13.31      
13 A' 1474 1474 1.44      
14 A' 1448 1448 5.29      
15 A' 1439 1439 8.19      
16 A' 1361 1361 3.32      
17 A' 1324 1324 0.17      
18 A' 1208 1208 2.31      
19 A' 1127 1127 9.92      
20 A' 1024 1024 1.29      
21 A' 979 979 0.59      
22 A' 913 913 1.37      
23 A' 739 739 0.99      
24 A' 527 527 0.97      
25 A' 447 447 0.88      
26 A' 336 336 0.17      
27 A' 281 281 0.40      
28 A' 264 264 0.09      
29 A" 3130 3130 16.86      
30 A" 3121 3121 6.26      
31 A" 3107 3107 28.12      
32 A" 3052 3052 27.62      
33 A" 1533 1533 0.14      
34 A" 1522 1522 1.52      
35 A" 1512 1512 11.39      
36 A" 1425 1425 7.58      
37 A" 1364 1364 1.71      
38 A" 1140 1140 3.20      
39 A" 1079 1079 0.13      
40 A" 979 979 0.02      
41 A" 943 943 1.12      
42 A" 902 902 45.58      
43 A" 747 747 0.75      
44 A" 556 556 8.80      
45 A" 266 266 0.06      
46 A" 214 214 0.52      
47 A" 165 165 0.16      
48 A" 37 37 0.15      

Unscaled Zero Point Vibrational Energy (zpe) 36476.0 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 36476.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULL/6-311G*
ABC
0.14765 0.08314 0.08228

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.953 -1.817 0.000
C2 -0.064 -0.818 0.000
C3 -0.518 0.630 0.000
C4 1.419 -1.094 0.000
C5 -0.064 1.376 1.263
C6 -0.064 1.376 -1.263
H7 -0.645 -2.857 0.000
H8 -2.021 -1.626 0.000
H9 -1.614 0.619 0.000
H10 1.618 -2.167 0.000
H11 1.909 -0.665 -0.878
H12 1.909 -0.665 0.878
H13 -0.494 2.380 1.290
H14 -0.377 0.850 2.167
H15 1.023 1.488 1.295
H16 -0.494 2.380 -1.290
H17 -0.377 0.850 -2.167
H18 1.023 1.488 -1.295

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.33712.48522.47983.54703.54701.08481.08482.52402.59473.20773.20774.41463.48434.06204.41463.48434.0620
C21.33711.51831.50822.53222.53222.11992.11772.11422.15522.16452.16453.47582.75302.85923.47582.75302.8592
C32.48521.51832.59381.53571.53573.48942.71111.09553.51932.88762.88762.17422.18272.18802.17422.18272.1880
C42.47981.50822.59383.14623.14622.71453.48113.48351.09051.09331.09334.17133.42132.91584.17133.42132.9158
C53.54702.53221.53573.14622.52584.45573.80032.13784.12013.55532.86441.09281.09201.09252.77673.48422.7817
C63.54702.53221.53573.14622.52584.45573.80032.13784.12012.86443.55532.77673.48422.78171.09281.09201.0925
H71.08482.11993.48942.71454.45574.45571.84593.60862.36603.47863.47865.39584.30254.83055.39584.30254.8305
H81.08482.11772.71113.48113.80033.80031.84592.28233.67884.14014.14014.47733.67854.54294.47733.67854.5429
H92.52402.11421.09553.48352.13782.13783.60862.28234.26673.85073.85072.45302.50563.06302.45302.50563.0630
H102.59472.15523.51931.09054.12014.12012.36603.67884.26671.76401.76405.17694.21653.92245.17694.21653.9224
H113.20772.16452.88761.09333.55532.86443.47864.14013.85071.76401.75594.44384.09803.18443.90103.03042.3646
H123.20772.16452.88761.09332.86443.55533.47864.14013.85071.76401.75593.90103.03042.36464.44384.09803.1844
H134.41463.47582.17424.17131.09282.77675.39584.47732.45305.17694.44383.90101.76751.76042.57973.78213.1277
H143.48432.75302.18273.42131.09203.48424.30253.67852.50564.21654.09803.03041.76751.76793.78214.33413.7887
H154.06202.85922.18802.91581.09252.78174.83054.54293.06303.92243.18442.36461.76041.76793.12773.78872.5908
H164.41463.47582.17424.17132.77671.09285.39584.47732.45305.17693.90104.44382.57973.78213.12771.76751.7604
H173.48432.75302.18273.42133.48421.09204.30253.67852.50564.21653.03044.09803.78214.33413.78871.76751.7679
H184.06202.85922.18802.91582.78171.09254.83054.54293.06303.92242.36463.18443.12773.78872.59081.76041.7679

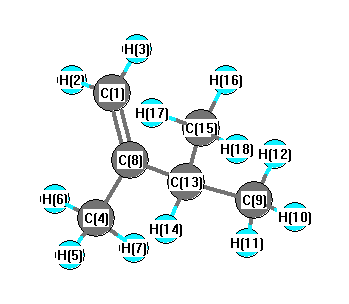
picture of 2,3-dimethylbut-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.870 C1 C2 C4 121.159
C2 C1 H7 121.809 C2 C1 H8 121.595
C2 C3 C5 112.020 C2 C3 C6 112.020
C2 C3 H9 106.857 C2 C4 H10 111.041
C2 C4 H11 111.617 C2 C4 H12 111.617
C3 C2 C4 117.970 C3 C5 H13 110.483
C3 C5 H14 111.214 C3 C5 H15 111.607
C3 C6 H16 110.483 C3 C6 H17 111.214
C3 C6 H18 111.607 C5 C3 C6 110.649
C5 C3 H9 107.506 C6 C3 H9 107.506
H7 C1 H8 116.596 H10 C4 H11 107.754
H10 C4 H12 107.754 H11 C4 H12 106.840
H13 C5 H14 107.992 H13 C5 H15 107.326
H14 C5 H15 108.056 H16 C6 H17 107.992
H16 C6 H18 107.326 H17 C6 H18 108.056
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability