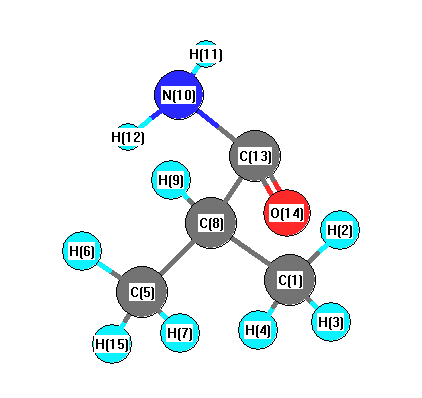
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3996 |
3614 |
39.56 |
|
|
|
| 2 |
A |
3857 |
3488 |
45.52 |
|
|
|
| 3 |
A |
3274 |
2961 |
62.69 |
|
|
|
| 4 |
A |
3267 |
2954 |
3.28 |
|
|
|
| 5 |
A |
3239 |
2930 |
80.35 |
|
|
|
| 6 |
A |
3235 |
2926 |
27.56 |
|
|
|
| 7 |
A |
3190 |
2885 |
25.40 |
|
|
|
| 8 |
A |
3181 |
2877 |
56.74 |
|
|
|
| 9 |
A |
3175 |
2872 |
36.20 |
|
|
|
| 10 |
A |
1946 |
1760 |
397.47 |
|
|
|
| 11 |
A |
1808 |
1635 |
126.33 |
|
|
|
| 12 |
A |
1650 |
1492 |
18.22 |
|
|
|
| 13 |
A |
1636 |
1479 |
10.29 |
|
|
|
| 14 |
A |
1625 |
1470 |
1.94 |
|
|
|
| 15 |
A |
1621 |
1466 |
0.79 |
|
|
|
| 16 |
A |
1559 |
1410 |
68.30 |
|
|
|
| 17 |
A |
1555 |
1406 |
5.98 |
|
|
|
| 18 |
A |
1537 |
1390 |
2.22 |
|
|
|
| 19 |
A |
1462 |
1322 |
0.54 |
|
|
|
| 20 |
A |
1394 |
1261 |
153.46 |
|
|
|
| 21 |
A |
1302 |
1178 |
7.22 |
|
|
|
| 22 |
A |
1242 |
1124 |
9.84 |
|
|
|
| 23 |
A |
1216 |
1100 |
0.01 |
|
|
|
| 24 |
A |
1165 |
1054 |
11.21 |
|
|
|
| 25 |
A |
1049 |
949 |
0.28 |
|
|
|
| 26 |
A |
1012 |
916 |
1.37 |
|
|
|
| 27 |
A |
982 |
888 |
0.86 |
|
|
|
| 28 |
A |
857 |
775 |
14.00 |
|
|
|
| 29 |
A |
818 |
740 |
1.28 |
|
|
|
| 30 |
A |
669 |
605 |
11.77 |
|
|
|
| 31 |
A |
633 |
573 |
16.38 |
|
|
|
| 32 |
A |
513 |
464 |
4.53 |
|
|
|
| 33 |
A |
346 |
313 |
0.96 |
|
|
|
| 34 |
A |
312 |
282 |
6.43 |
|
|
|
| 35 |
A |
264 |
239 |
4.22 |
|
|
|
| 36 |
A |
262 |
237 |
0.03 |
|
|
|
| 37 |
A |
235 |
213 |
0.23 |
|
|
|
| 38 |
A |
122 |
110 |
262.16 |
|
|
|
| 39 |
A |
12 |
11 |
10.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30608.8 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 27682.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.607 |
-0.263 |
|
-0.472 |
| 2 |
H |
0.213 |
0.041 |
|
0.101 |
| 3 |
H |
0.237 |
0.071 |
|
0.133 |
| 4 |
H |
0.210 |
0.071 |
|
0.128 |
| 5 |
C |
-0.607 |
-0.263 |
|
-0.472 |
| 6 |
H |
0.213 |
0.041 |
|
0.101 |
| 7 |
H |
0.237 |
0.071 |
|
0.133 |
| 8 |
C |
-0.338 |
0.202 |
|
0.227 |
| 9 |
H |
0.198 |
-0.039 |
|
-0.001 |
| 10 |
N |
-0.906 |
-1.184 |
|
-1.165 |
| 11 |
H |
0.383 |
0.484 |
|
0.483 |
| 12 |
H |
0.379 |
0.447 |
|
0.443 |
| 13 |
C |
0.677 |
0.925 |
|
0.882 |
| 14 |
O |
-0.501 |
-0.675 |
|
-0.648 |
| 15 |
H |
0.210 |
0.071 |
|
0.128 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.058 |
0.002 |
-3.861 |
3.862 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.036 |
0.002 |
-3.859 |
3.859 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.173 |
0.002 |
-4.889 |
| y |
0.002 |
-38.585 |
0.003 |
| z |
-4.889 |
0.003 |
-39.686 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.963 |
0.002 |
-4.889 |
| y |
0.002 |
-2.155 |
0.003 |
| z |
-4.889 |
0.003 |
-3.807 |
|
| Polar |
|---|
| 3z2-r2 | -7.615 |
|---|
| x2-y2 | 5.412 |
|---|
| xy | 0.002 |
|---|
| xz | -4.889 |
|---|
| yz | 0.003 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.936 |
0.000 |
-0.217 |
| y |
0.000 |
6.626 |
-0.001 |
| z |
-0.217 |
-0.001 |
7.443 |
<r2> (average value of r
2) Å
2
| <r2> |
171.794 |
| (<r2>)1/2 |
13.107 |