return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9NO (Propanamide, 2-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-287.135401
Energy at 298.15K-287.145948
HF Energy-286.106756
Nuclear repulsion energy248.281132
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3786 3586 32.13      
2 A 3644 3451 38.97      
3 A 3186 3017 22.56      
4 A 3181 3013 5.70      
5 A 3168 3000 29.05      
6 A 3164 2997 21.61      
7 A 3085 2922 25.42      
8 A 3080 2917 21.67      
9 A 3078 2915 25.51      
10 A 1808 1713 248.83      
11 A 1686 1597 97.43      
12 A 1554 1472 22.94      
13 A 1542 1460 14.59      
14 A 1528 1447 2.64      
15 A 1524 1444 1.79      
16 A 1455 1378 71.76      
17 A 1446 1369 14.34      
18 A 1430 1355 9.79      
19 A 1361 1289 1.25      
20 A 1310 1241 87.51      
21 A 1227 1162 3.52      
22 A 1161 1099 3.94      
23 A 1158 1097 3.71      
24 A 1088 1031 7.32      
25 A 1002 949 0.32      
26 A 953 903 2.88      
27 A 949 899 2.09      
28 A 788 746 7.28      
29 A 782 741 3.69      
30 A 627 594 12.60      
31 A 596 565 28.26      
32 A 487 462 2.93      
33 A 334 316 11.86      
34 A 304 288 196.61      
35 A 299 283 26.45      
36 A 259 245 5.60      
37 A 248 235 8.45      
38 A 217 206 5.49      
39 A 24 23 7.67      

Unscaled Zero Point Vibrational Energy (zpe) 29258.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 27710.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.16438 0.08822 0.08501

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.324 1.267 0.182
H2 0.793 2.205 -0.003
H3 1.378 1.114 1.262
H4 2.339 1.367 -0.209
C5 1.367 -1.223 -0.226
H6 0.866 -2.067 -0.706
H7 1.418 -1.423 0.846
C8 0.610 0.082 -0.469
H9 0.537 0.254 -1.550
N10 -1.820 0.112 -0.748
H11 -2.747 0.166 -0.358
H12 -1.671 0.507 -1.659
C13 -0.783 -0.039 0.132
O14 -0.957 -0.284 1.315
H15 2.384 -1.157 -0.621

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 N10 H11 H12 C13 O14 H15
C11.09341.09171.09312.52283.48032.77161.52882.15463.47594.25093.59632.47892.98172.7646
H21.09341.76901.77083.48234.33023.77742.18152.50263.43004.10043.42022.74523.31553.7703
H31.09171.76901.77512.76993.77532.57082.15643.05763.90774.53134.26532.69732.72203.1167
H41.09311.77081.77512.76603.76963.12122.17032.50704.37815.22814.35033.44133.98952.5575
C52.52283.48232.76992.76601.09301.09141.52762.15033.49494.34413.77842.48062.94281.0931
H63.48034.33023.77533.76961.09301.76872.17712.49223.45974.26203.73822.74543.25451.7720
H72.77163.77742.57083.12121.09141.76872.15533.05433.92264.61774.42072.69682.67631.7760
C81.52882.18152.15642.17031.52762.17712.15531.09692.44643.35972.60761.52182.40272.1692
H92.15462.50263.05762.50702.15032.49223.05431.09692.49333.49412.22472.15743.27542.5038
N103.47593.43003.90774.37813.49493.45973.92262.44642.49331.00701.00371.36842.27144.3937
H114.25094.10044.53135.22814.34414.26204.61773.35973.49411.00701.72242.03412.49075.3055
H123.59633.42024.26534.35033.77843.73824.42072.60762.22471.00371.72242.07183.15914.5047
C132.47892.74522.69733.44132.48062.74542.69681.52182.15741.36842.03412.07181.22113.4423
O142.98173.31552.72203.98952.94283.25452.67632.40273.27542.27142.49073.15911.22113.9595
H152.76463.77033.11672.55751.09311.77201.77602.16922.50384.39375.30554.50473.44233.9595

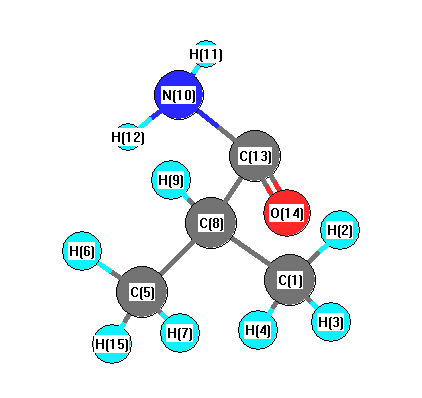
picture of Propanamide, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C8 C5 111.265 C1 C8 H9 109.183
C1 C8 C13 108.702 H2 C1 H3 108.112
H2 C1 H4 108.172 H2 C1 C8 111.516
H3 C1 H4 108.686 H3 C1 C8 109.625
H4 C1 C8 110.647 C5 C8 H9 108.931
C5 C8 C13 108.869 H6 C5 H7 108.130
H6 C5 C8 111.275 H6 C5 H15 108.309
H7 C5 C8 109.635 H7 C5 H15 108.782
C8 C5 H15 110.636 C8 C13 N10 115.547
C8 C13 O14 121.934 H9 C8 C13 109.879
N10 C13 O14 122.500 H11 N10 H12 117.878
H11 N10 C13 116.991 H12 N10 C13 120.940
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability