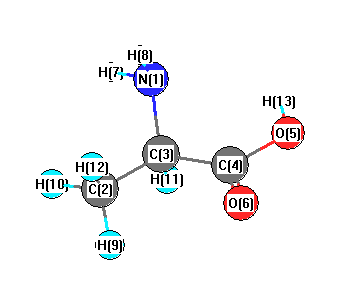
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3615 |
3480 |
4.98 |
|
|
|
| 2 |
A |
3526 |
3395 |
0.17 |
|
|
|
| 3 |
A |
3423 |
3296 |
233.17 |
|
|
|
| 4 |
A |
3149 |
3031 |
8.91 |
|
|
|
| 5 |
A |
3113 |
2997 |
32.27 |
|
|
|
| 6 |
A |
3051 |
2937 |
19.43 |
|
|
|
| 7 |
A |
3043 |
2929 |
12.38 |
|
|
|
| 8 |
A |
1874 |
1805 |
303.72 |
|
|
|
| 9 |
A |
1709 |
1645 |
45.47 |
|
|
|
| 10 |
A |
1511 |
1455 |
16.59 |
|
|
|
| 11 |
A |
1501 |
1445 |
5.39 |
|
|
|
| 12 |
A |
1459 |
1404 |
441.15 |
|
|
|
| 13 |
A |
1415 |
1362 |
14.72 |
|
|
|
| 14 |
A |
1401 |
1349 |
2.28 |
|
|
|
| 15 |
A |
1322 |
1272 |
8.73 |
|
|
|
| 16 |
A |
1245 |
1198 |
9.13 |
|
|
|
| 17 |
A |
1225 |
1180 |
14.12 |
|
|
|
| 18 |
A |
1161 |
1118 |
10.45 |
|
|
|
| 19 |
A |
1105 |
1063 |
50.41 |
|
|
|
| 20 |
A |
1024 |
986 |
1.38 |
|
|
|
| 21 |
A |
947 |
912 |
64.91 |
|
|
|
| 22 |
A |
916 |
882 |
68.21 |
|
|
|
| 23 |
A |
879 |
847 |
111.72 |
|
|
|
| 24 |
A |
811 |
781 |
8.87 |
|
|
|
| 25 |
A |
744 |
716 |
8.65 |
|
|
|
| 26 |
A |
568 |
547 |
5.12 |
|
|
|
| 27 |
A |
535 |
515 |
0.93 |
|
|
|
| 28 |
A |
409 |
394 |
5.21 |
|
|
|
| 29 |
A |
373 |
359 |
16.45 |
|
|
|
| 30 |
A |
316 |
305 |
13.46 |
|
|
|
| 31 |
A |
275 |
265 |
12.21 |
|
|
|
| 32 |
A |
228 |
219 |
0.34 |
|
|
|
| 33 |
A |
76 |
73 |
1.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23972.9 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 23078.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.824 |
|
|
|
| 2 |
C |
-0.646 |
|
|
|
| 3 |
C |
-0.257 |
|
|
|
| 4 |
C |
0.496 |
|
|
|
| 5 |
O |
-0.500 |
|
|
|
| 6 |
O |
-0.346 |
|
|
|
| 7 |
H |
0.346 |
|
|
|
| 8 |
H |
0.354 |
|
|
|
| 9 |
H |
0.259 |
|
|
|
| 10 |
H |
0.222 |
|
|
|
| 11 |
H |
0.258 |
|
|
|
| 12 |
H |
0.219 |
|
|
|
| 13 |
H |
0.420 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
5.024 |
2.220 |
0.333 |
5.502 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.279 |
0.040 |
-0.337 |
| y |
0.040 |
-37.372 |
-0.083 |
| z |
-0.337 |
-0.083 |
-33.443 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.872 |
0.040 |
-0.337 |
| y |
0.040 |
-1.011 |
-0.083 |
| z |
-0.337 |
-0.083 |
4.883 |
|
| Polar |
|---|
| 3z2-r2 | 9.766 |
|---|
| x2-y2 | -1.908 |
|---|
| xy | 0.040 |
|---|
| xz | -0.337 |
|---|
| yz | -0.083 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.139 |
0.178 |
0.080 |
| y |
0.178 |
7.629 |
-0.048 |
| z |
0.080 |
-0.048 |
5.427 |
<r2> (average value of r
2) Å
2
| <r2> |
162.292 |
| (<r2>)1/2 |
12.739 |