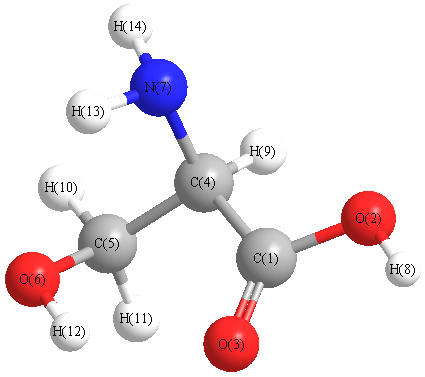
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3792 |
3635 |
48.31 |
|
|
|
| 2 |
A |
3779 |
3622 |
62.40 |
|
|
|
| 3 |
A |
3625 |
3475 |
2.37 |
|
|
|
| 4 |
A |
3539 |
3393 |
0.53 |
|
|
|
| 5 |
A |
3144 |
3013 |
23.09 |
|
|
|
| 6 |
A |
3066 |
2939 |
15.82 |
|
|
|
| 7 |
A |
2996 |
2872 |
64.33 |
|
|
|
| 8 |
A |
1832 |
1756 |
278.14 |
|
|
|
| 9 |
A |
1724 |
1653 |
43.80 |
|
|
|
| 10 |
A |
1516 |
1453 |
2.97 |
|
|
|
| 11 |
A |
1461 |
1400 |
51.71 |
|
|
|
| 12 |
A |
1413 |
1355 |
4.99 |
|
|
|
| 13 |
A |
1408 |
1350 |
50.65 |
|
|
|
| 14 |
A |
1395 |
1337 |
28.68 |
|
|
|
| 15 |
A |
1337 |
1281 |
8.20 |
|
|
|
| 16 |
A |
1259 |
1206 |
19.07 |
|
|
|
| 17 |
A |
1217 |
1167 |
69.14 |
|
|
|
| 18 |
A |
1199 |
1149 |
170.25 |
|
|
|
| 19 |
A |
1163 |
1115 |
22.52 |
|
|
|
| 20 |
A |
1128 |
1082 |
115.31 |
|
|
|
| 21 |
A |
1047 |
1003 |
36.68 |
|
|
|
| 22 |
A |
1010 |
968 |
0.56 |
|
|
|
| 23 |
A |
900 |
863 |
198.35 |
|
|
|
| 24 |
A |
829 |
795 |
33.95 |
|
|
|
| 25 |
A |
758 |
727 |
39.46 |
|
|
|
| 26 |
A |
663 |
635 |
103.57 |
|
|
|
| 27 |
A |
584 |
560 |
42.41 |
|
|
|
| 28 |
A |
573 |
549 |
200.64 |
|
|
|
| 29 |
A |
529 |
507 |
16.32 |
|
|
|
| 30 |
A |
439 |
421 |
8.17 |
|
|
|
| 31 |
A |
309 |
296 |
6.01 |
|
|
|
| 32 |
A |
293 |
281 |
10.29 |
|
|
|
| 33 |
A |
257 |
247 |
33.38 |
|
|
|
| 34 |
A |
224 |
215 |
8.50 |
|
|
|
| 35 |
A |
170 |
163 |
2.55 |
|
|
|
| 36 |
A |
31 |
29 |
0.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25304.6 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 24254.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.468 |
|
|
|
| 2 |
O |
-0.515 |
|
|
|
| 3 |
O |
-0.368 |
|
|
|
| 4 |
C |
-0.255 |
|
|
|
| 5 |
C |
-0.206 |
|
|
|
| 6 |
O |
-0.562 |
|
|
|
| 7 |
N |
-0.745 |
|
|
|
| 8 |
H |
0.410 |
|
|
|
| 9 |
H |
0.265 |
|
|
|
| 10 |
H |
0.190 |
|
|
|
| 11 |
H |
0.242 |
|
|
|
| 12 |
H |
0.406 |
|
|
|
| 13 |
H |
0.344 |
|
|
|
| 14 |
H |
0.326 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.527 |
0.237 |
0.775 |
3.619 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.632 |
-2.220 |
-1.141 |
| y |
-2.220 |
-44.242 |
1.423 |
| z |
-1.141 |
1.423 |
-39.722 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.350 |
-2.220 |
-1.141 |
| y |
-2.220 |
-3.565 |
1.423 |
| z |
-1.141 |
1.423 |
3.215 |
|
| Polar |
|---|
| 3z2-r2 | 6.430 |
|---|
| x2-y2 | 2.610 |
|---|
| xy | -2.220 |
|---|
| xz | -1.141 |
|---|
| yz | 1.423 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.961 |
0.144 |
-0.173 |
| y |
0.144 |
7.734 |
-0.126 |
| z |
-0.173 |
-0.126 |
5.823 |
<r2> (average value of r
2) Å
2
| <r2> |
214.551 |
| (<r2>)1/2 |
14.648 |