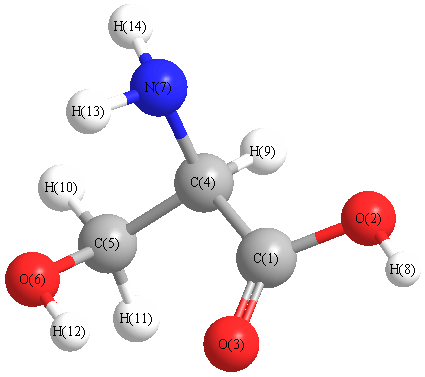
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3783 |
3630 |
48.11 |
|
|
|
| 2 |
A |
3770 |
3618 |
62.32 |
|
|
|
| 3 |
A |
3621 |
3475 |
2.23 |
|
|
|
| 4 |
A |
3535 |
3393 |
0.51 |
|
|
|
| 5 |
A |
3141 |
3015 |
23.80 |
|
|
|
| 6 |
A |
3064 |
2940 |
16.52 |
|
|
|
| 7 |
A |
2993 |
2872 |
65.29 |
|
|
|
| 8 |
A |
1828 |
1754 |
278.11 |
|
|
|
| 9 |
A |
1724 |
1654 |
43.43 |
|
|
|
| 10 |
A |
1516 |
1455 |
2.89 |
|
|
|
| 11 |
A |
1461 |
1402 |
53.46 |
|
|
|
| 12 |
A |
1413 |
1355 |
6.04 |
|
|
|
| 13 |
A |
1407 |
1351 |
43.04 |
|
|
|
| 14 |
A |
1394 |
1338 |
31.62 |
|
|
|
| 15 |
A |
1337 |
1283 |
7.86 |
|
|
|
| 16 |
A |
1258 |
1208 |
18.44 |
|
|
|
| 17 |
A |
1216 |
1167 |
69.01 |
|
|
|
| 18 |
A |
1197 |
1148 |
174.52 |
|
|
|
| 19 |
A |
1161 |
1114 |
22.17 |
|
|
|
| 20 |
A |
1125 |
1079 |
114.91 |
|
|
|
| 21 |
A |
1046 |
1004 |
36.77 |
|
|
|
| 22 |
A |
1010 |
969 |
0.38 |
|
|
|
| 23 |
A |
899 |
863 |
198.55 |
|
|
|
| 24 |
A |
829 |
795 |
33.74 |
|
|
|
| 25 |
A |
757 |
727 |
38.88 |
|
|
|
| 26 |
A |
662 |
635 |
103.48 |
|
|
|
| 27 |
A |
585 |
561 |
61.71 |
|
|
|
| 28 |
A |
575 |
552 |
179.44 |
|
|
|
| 29 |
A |
529 |
508 |
17.77 |
|
|
|
| 30 |
A |
439 |
421 |
8.06 |
|
|
|
| 31 |
A |
309 |
296 |
5.90 |
|
|
|
| 32 |
A |
293 |
281 |
9.94 |
|
|
|
| 33 |
A |
258 |
247 |
33.39 |
|
|
|
| 34 |
A |
226 |
217 |
8.95 |
|
|
|
| 35 |
A |
171 |
164 |
2.47 |
|
|
|
| 36 |
A |
31 |
30 |
0.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25280.8 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 24259.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.462 |
|
|
|
| 2 |
O |
-0.513 |
|
|
|
| 3 |
O |
-0.365 |
|
|
|
| 4 |
C |
-0.251 |
|
|
|
| 5 |
C |
-0.206 |
|
|
|
| 6 |
O |
-0.561 |
|
|
|
| 7 |
N |
-0.742 |
|
|
|
| 8 |
H |
0.409 |
|
|
|
| 9 |
H |
0.264 |
|
|
|
| 10 |
H |
0.190 |
|
|
|
| 11 |
H |
0.241 |
|
|
|
| 12 |
H |
0.405 |
|
|
|
| 13 |
H |
0.343 |
|
|
|
| 14 |
H |
0.325 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.529 |
0.249 |
0.773 |
3.621 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.659 |
-2.255 |
-1.135 |
| y |
-2.255 |
-44.246 |
1.425 |
| z |
-1.135 |
1.425 |
-39.753 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.341 |
-2.255 |
-1.135 |
| y |
-2.255 |
-3.540 |
1.425 |
| z |
-1.135 |
1.425 |
3.199 |
|
| Polar |
|---|
| 3z2-r2 | 6.398 |
|---|
| x2-y2 | 2.587 |
|---|
| xy | -2.255 |
|---|
| xz | -1.135 |
|---|
| yz | 1.425 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.980 |
0.140 |
-0.175 |
| y |
0.140 |
7.749 |
-0.126 |
| z |
-0.175 |
-0.126 |
5.835 |
<r2> (average value of r
2) Å
2
| <r2> |
214.506 |
| (<r2>)1/2 |
14.646 |