return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HO2CCHNH2CH2OH (Serine)

using model chemistry: B2PLYP=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP=FULL/6-311G*
 hartrees
Energy at 0K-398.721282
Energy at 298.15K-398.731377
HF Energy-398.299679
Nuclear repulsion energy325.799903
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3794 3794 44.63      
2 A 3749 3749 51.13      
3 A 3614 3614 1.92      
4 A 3526 3526 0.58      
5 A 3156 3156 23.30      
6 A 3086 3086 15.38      
7 A 3019 3019 58.31      
8 A 1793 1793 248.81      
9 A 1735 1735 39.23      
10 A 1535 1535 2.44      
11 A 1462 1462 50.61      
12 A 1425 1425 2.43      
13 A 1417 1417 22.41      
14 A 1398 1398 34.01      
15 A 1343 1343 6.48      
16 A 1267 1267 15.63      
17 A 1211 1211 25.00      
18 A 1186 1186 217.50      
19 A 1153 1153 43.12      
20 A 1103 1103 100.82      
21 A 1047 1047 39.62      
22 A 1013 1013 0.22      
23 A 913 913 201.38      
24 A 826 826 23.18      
25 A 750 750 34.47      
26 A 654 654 107.18      
27 A 585 585 6.74      
28 A 561 561 238.53      
29 A 522 522 14.36      
30 A 446 446 7.48      
31 A 306 306 6.18      
32 A 292 292 16.49      
33 A 268 268 30.33      
34 A 221 221 3.68      
35 A 172 172 3.28      
36 A 35 35 0.95      

Unscaled Zero Point Vibrational Energy (zpe) 25292.2 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 25292.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULL/6-311G*
ABC
0.11887 0.07853 0.05154

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.775 -0.547 0.001
O2 2.057 -0.351 -0.357
O3 0.393 -1.560 0.543
C4 -0.085 0.677 -0.310
C5 -1.512 0.450 0.188
O6 -2.107 -0.692 -0.380
N7 0.446 1.925 0.227
H8 2.553 -1.147 -0.124
H9 -0.124 0.765 -1.400
H10 -1.503 0.397 1.286
H11 -2.108 1.315 -0.099
H12 -1.659 -1.458 -0.010
H13 1.345 2.143 -0.182
H14 0.570 1.862 1.230

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 O6 N7 H8 H9 H10 H11 H12 H13 H14
C11.34591.21041.52792.50192.91052.50441.88092.11922.78083.43342.59882.75612.7129
O21.34592.24532.37693.69884.17842.84930.96632.66303.99224.49373.89342.59993.1034
O31.21042.24532.44102.79182.80263.49962.29813.07352.82413.86392.12773.89133.4948
C41.52792.37692.44101.52852.44281.45923.21281.09392.15342.13162.66932.05222.0511
C52.50193.69882.79181.52851.40802.45224.37892.13281.09871.08891.92403.34162.7230
O62.91054.17842.80262.44281.40803.70674.68922.66382.08012.02660.96134.47154.0357
N72.50442.84933.49961.45922.45223.70673.74192.07782.69422.64663.99191.01091.0127
H81.88090.96632.29813.21284.37894.68923.74193.52864.56365.27154.22523.50573.8502
H92.11922.66303.07351.09392.13282.66382.07783.52863.04162.43533.03822.35372.9329
H102.78083.99222.82412.15341.09872.08012.69424.56363.04161.76842.26803.64922.5397
H113.43344.49373.86392.13161.08892.02662.64665.27152.43531.76842.81033.55203.0397
H122.59883.89342.12772.66931.92400.96133.99194.22523.03822.26802.81034.69284.1871
H132.75612.59993.89132.05223.34164.47151.01093.50572.35373.64923.55204.69281.6349
H142.71293.10343.49482.05112.72304.03571.01273.85022.93292.53973.03974.18711.6349

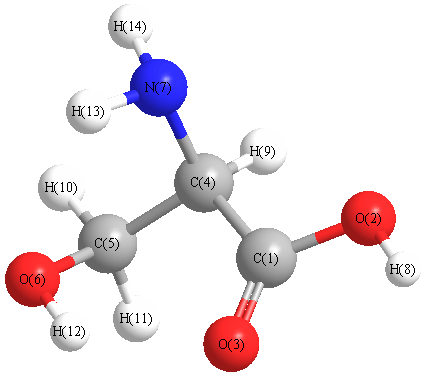
picture of Serine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 H8 107.740 C1 C4 C5 109.891
C1 C4 N7 113.924 C1 C4 H9 106.691
O2 C1 O3 122.794 O2 C1 C4 111.442
O3 C1 C4 125.713 C4 C5 O6 112.521
C4 C5 H10 109.001 C4 C5 H11 107.879
C4 N7 H13 111.057 C4 N7 H14 110.855
C5 C4 N7 110.298 C5 C4 H9 107.687
C5 O6 H12 107.082 O6 C5 H10 111.561
O6 C5 H11 107.832 N7 C4 H9 108.090
H10 C5 H11 107.868 H13 N7 H14 107.783
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability