return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6 (Dewar Benzene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-231.520329
Energy at 298.15K 
HF Energy-230.603208
Nuclear repulsion energy208.641527
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3244 3072 8.53 276.87 0.04 0.07
2 A1 3151 2985 29.19 148.08 0.14 0.24
3 A1 1589 1505 0.23 15.22 0.02 0.05
4 A1 1184 1121 5.35 20.13 0.14 0.25
5 A1 1045 989 1.90 9.26 0.74 0.85
6 A1 973 922 4.32 9.26 0.27 0.42
7 A1 886 839 0.79 5.21 0.73 0.84
8 A1 816 772 84.40 4.89 0.08 0.16
9 A1 391 370 5.52 9.71 0.64 0.78
10 A2 3211 3041 0.00 163.91 0.75 0.86
11 A2 1302 1233 0.00 6.08 0.75 0.86
12 A2 1215 1151 0.00 8.74 0.75 0.86
13 A2 981 929 0.00 0.28 0.75 0.86
14 A2 885 839 0.00 0.41 0.75 0.86
15 A2 777 736 0.00 2.07 0.75 0.86
16 A2 321 304 0.00 2.01 0.75 0.86
17 B1 3241 3070 41.33 69.28 0.75 0.86
18 B1 1565 1482 5.58 1.48 0.75 0.86
19 B1 1230 1165 1.60 1.67 0.75 0.86
20 B1 1117 1058 0.77 0.25 0.75 0.86
21 B1 1029 974 0.00 6.98 0.75 0.86
22 B1 718 680 54.93 3.96 0.75 0.86
23 B2 3212 3042 25.29 46.44 0.75 0.86
24 B2 3145 2978 12.01 115.84 0.75 0.86
25 B2 1303 1234 38.04 0.02 0.75 0.86
26 B2 1168 1106 7.33 0.70 0.75 0.86
27 B2 932 883 9.14 0.02 0.75 0.86
28 B2 915 866 4.54 0.89 0.75 0.86
29 B2 833 789 8.58 0.73 0.75 0.86
30 B2 480 454 4.84 5.13 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 21427.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 20294.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.25902 0.15015 0.11677

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
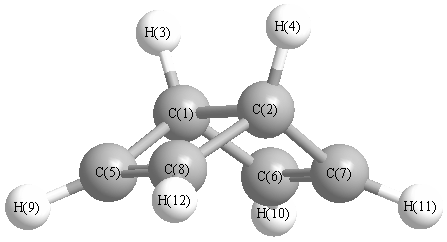
C1 0.000 0.788 0.527
C2 0.000 -0.788 0.527
H3 0.000 1.344 1.467
H4 0.000 -1.344 1.467
C5 -1.296 0.675 -0.265
C6 1.296 0.675 -0.265
C7 1.296 -0.675 -0.265
C8 -1.296 -0.675 -0.265
H9 -1.936 1.425 -0.722
H10 1.936 1.425 -0.722
H11 1.936 -1.425 -0.722
H12 -1.936 -1.425 -0.722

Atom - Atom Distances (Å)
  C1 C2 H3 H4 C5 C6 C7 C8 H9 H10 H11 H12
C11.57531.09252.33001.52361.52362.10912.10912.39042.39043.19433.1943
C21.57532.33001.09252.10912.10911.52361.52363.19433.19432.39042.3904
H31.09252.33002.68872.26502.26502.95982.95982.92342.92344.02614.0261
H42.33001.09252.68872.95982.95982.26502.26504.02614.02612.92342.9234
C51.52362.10912.26502.95982.59282.92331.35011.08623.34933.88152.2423
C61.52362.10912.26502.95982.59281.35012.92333.34931.08622.24233.8815
C72.10911.52362.95982.26502.92331.35012.59283.88152.24231.08623.3493
C82.10911.52362.95982.26501.35012.92332.59282.24233.88153.34931.0862
H92.39043.19432.92344.02611.08623.34933.88152.24233.87154.80752.8501
H102.39043.19432.92344.02613.34931.08622.24233.88153.87152.85014.8075
H113.19432.39044.02612.92343.88152.24231.08623.34934.80752.85013.8715
H123.19432.39044.02612.92342.24233.88153.34931.08622.85014.80753.8715

picture of Dewar Benzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 120.635 C1 C2 H7 85.763
C1 C2 H8 85.763 C1 C5 H8 94.237
C1 C5 H9 131.951 C1 C6 H7 94.237
C1 C6 H10 131.951 C2 C1 C3 120.635
C2 C1 C5 85.763 C2 C1 C6 85.763
C2 H7 C6 94.237 C2 H7 H11 131.951
C2 H8 C5 94.237 C2 H8 H12 131.951
C3 C1 C5 119.027 C3 C1 C6 119.027
C4 C2 H7 119.027 C4 C2 H8 119.027
C5 C1 C6 116.612 C5 H8 H12 133.670
C6 H7 H11 133.670 H7 C6 H10 133.670
H8 C5 H9 133.670
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability