Jump to
S1C2
S1C3
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -224.042241 |
| Energy at 298.15K | -224.048677 |
| HF Energy | -224.042241 |
| Nuclear repulsion energy | 125.272806 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3938 |
3562 |
37.51 |
|
|
|
| 2 |
A |
3831 |
3464 |
4.02 |
|
|
|
| 3 |
A |
1986 |
1797 |
572.57 |
|
|
|
| 4 |
A |
1820 |
1646 |
1.61 |
|
|
|
| 5 |
A |
1307 |
1182 |
7.50 |
|
|
|
| 6 |
A |
1025 |
927 |
9.18 |
|
|
|
| 7 |
A |
647 |
586 |
93.91 |
|
|
|
| 8 |
A |
514 |
465 |
3.27 |
|
|
|
| 9 |
A |
413 |
373 |
99.15 |
|
|
|
| 10 |
B |
3938 |
3562 |
22.13 |
|
|
|
| 11 |
B |
3827 |
3461 |
52.04 |
|
|
|
| 12 |
B |
1828 |
1653 |
220.37 |
|
|
|
| 13 |
B |
1538 |
1391 |
250.64 |
|
|
|
| 14 |
B |
1164 |
1053 |
29.52 |
|
|
|
| 15 |
B |
888 |
803 |
151.32 |
|
|
|
| 16 |
B |
632 |
571 |
98.08 |
|
|
|
| 17 |
B |
597 |
540 |
254.05 |
|
|
|
| 18 |
B |
483 |
437 |
96.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15188.1 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 13736.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.147 |
| O2 |
0.000 |
0.000 |
1.338 |
| N3 |
-0.118 |
1.146 |
-0.603 |
| N4 |
0.118 |
-1.146 |
-0.603 |
| H5 |
0.000 |
1.977 |
-0.074 |
| H6 |
0.325 |
1.161 |
-1.492 |
| H7 |
0.000 |
-1.977 |
-0.074 |
| H8 |
-0.325 |
-1.161 |
-1.492 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.1912 | 1.3743 | 1.3743 | 1.9896 | 2.0346 | 1.9896 | 2.0346 |
O2 | 1.1912 | | 2.2571 | 2.2571 | 2.4297 | 3.0762 | 2.4297 | 3.0762 | N3 | 1.3743 | 2.2571 | | 2.3032 | 0.9930 | 0.9930 | 3.1697 | 2.4806 | N4 | 1.3743 | 2.2571 | 2.3032 | | 3.1697 | 2.4806 | 0.9930 | 0.9930 | H5 | 1.9896 | 2.4297 | 0.9930 | 3.1697 | | 1.6684 | 3.9548 | 3.4593 | H6 | 2.0346 | 3.0762 | 0.9930 | 2.4806 | 1.6684 | | 3.4593 | 2.4112 | H7 | 1.9896 | 2.4297 | 3.1697 | 0.9930 | 3.9548 | 3.4593 | | 1.6684 | H8 | 2.0346 | 3.0762 | 2.4806 | 0.9930 | 3.4593 | 2.4112 | 1.6684 | |
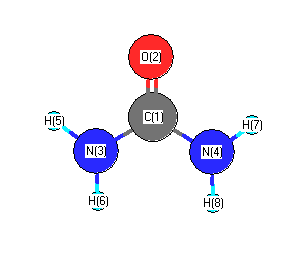 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
113.402 |
|
C1 |
N3 |
H6 |
117.607 |
| C1 |
N4 |
H7 |
113.402 |
|
C1 |
N4 |
H8 |
117.607 |
| O2 |
C1 |
N3 |
123.075 |
|
O2 |
C1 |
N4 |
123.075 |
| N3 |
C1 |
N4 |
113.851 |
|
H5 |
N3 |
H6 |
114.300 |
| H7 |
N4 |
H8 |
114.300 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.772 |
|
|
|
| 2 |
O |
-0.498 |
|
|
|
| 3 |
N |
-0.874 |
|
|
|
| 4 |
N |
-0.874 |
|
|
|
| 5 |
H |
0.378 |
|
|
|
| 6 |
H |
0.359 |
|
|
|
| 7 |
H |
0.378 |
|
|
|
| 8 |
H |
0.359 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.853 |
3.853 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.009 |
-0.007 |
-3.857 |
3.857 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.339 |
2.781 |
0.000 |
| y |
2.781 |
-17.069 |
0.000 |
| z |
0.000 |
0.000 |
-25.320 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.145 |
2.781 |
0.000 |
| y |
2.781 |
8.261 |
0.000 |
| z |
0.000 |
0.000 |
-4.116 |
|
| Polar |
|---|
| 3z2-r2 | -8.233 |
|---|
| x2-y2 | -8.270 |
|---|
| xy | 2.781 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.294 |
-0.046 |
0.000 |
| y |
-0.046 |
4.029 |
0.000 |
| z |
0.000 |
0.000 |
4.409 |
<r2> (average value of r
2) Å
2
| <r2> |
67.404 |
| (<r2>)1/2 |
8.210 |
Jump to
S1C1
S1C3
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -224.040134 |
| Energy at 298.15K | |
| HF Energy | -224.040134 |
| Nuclear repulsion energy | 125.359035 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3977 |
3597 |
44.95 |
|
|
|
| 2 |
A' |
3858 |
3490 |
11.49 |
|
|
|
| 3 |
A' |
1967 |
1779 |
617.18 |
|
|
|
| 4 |
A' |
1824 |
1650 |
10.22 |
|
|
|
| 5 |
A' |
1293 |
1170 |
6.72 |
|
|
|
| 6 |
A' |
1037 |
937 |
13.21 |
|
|
|
| 7 |
A' |
867 |
785 |
61.50 |
|
|
|
| 8 |
A' |
585 |
529 |
30.11 |
|
|
|
| 9 |
A' |
521 |
471 |
15.92 |
|
|
|
| 10 |
A' |
375 |
340 |
510.06 |
|
|
|
| 11 |
A" |
3973 |
3594 |
26.23 |
|
|
|
| 12 |
A" |
3850 |
3482 |
55.73 |
|
|
|
| 13 |
A" |
1813 |
1640 |
240.38 |
|
|
|
| 14 |
A" |
1541 |
1394 |
272.55 |
|
|
|
| 15 |
A" |
1121 |
1014 |
21.73 |
|
|
|
| 16 |
A" |
625 |
565 |
27.47 |
|
|
|
| 17 |
A" |
447 |
404 |
50.74 |
|
|
|
| 18 |
A" |
102i |
92i |
51.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14786.6 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 13373.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.011 |
0.143 |
0.000 |
| O2 |
0.031 |
1.336 |
0.000 |
| N3 |
0.031 |
-0.593 |
1.153 |
| N4 |
0.031 |
-0.593 |
-1.153 |
| H5 |
-0.141 |
-0.080 |
1.983 |
| H6 |
-0.235 |
-1.547 |
1.159 |
| H7 |
-0.141 |
-0.080 |
-1.983 |
| H8 |
-0.235 |
-1.547 |
-1.159 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.1937 | 1.3675 | 1.3675 | 2.0017 | 2.0633 | 2.0017 | 2.0633 |
O2 | 1.1937 | | 2.2472 | 2.2472 | 2.4434 | 3.1185 | 2.4434 | 3.1185 | N3 | 1.3675 | 2.2472 | | 2.3055 | 0.9911 | 0.9903 | 3.1824 | 2.5147 | N4 | 1.3675 | 2.2472 | 2.3055 | | 3.1824 | 2.5147 | 0.9911 | 0.9903 | H5 | 2.0017 | 2.4434 | 0.9911 | 3.1824 | | 1.6849 | 3.9668 | 3.4687 | H6 | 2.0633 | 3.1185 | 0.9903 | 2.5147 | 1.6849 | | 3.4687 | 2.3176 | H7 | 2.0017 | 2.4434 | 3.1824 | 0.9911 | 3.9668 | 3.4687 | | 1.6849 | H8 | 2.0633 | 3.1185 | 2.5147 | 0.9903 | 3.4687 | 2.3176 | 1.6849 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
115.207 |
|
C1 |
N3 |
H6 |
121.281 |
| C1 |
N4 |
H7 |
115.207 |
|
C1 |
N4 |
H8 |
121.281 |
| O2 |
C1 |
N3 |
122.512 |
|
O2 |
C1 |
N4 |
122.512 |
| N3 |
C1 |
N4 |
114.904 |
|
H5 |
N3 |
H6 |
116.503 |
| H7 |
N4 |
H8 |
116.503 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.807 |
|
|
|
| 2 |
O |
-0.513 |
|
|
|
| 3 |
N |
-0.887 |
|
|
|
| 4 |
N |
-0.887 |
|
|
|
| 5 |
H |
0.384 |
|
|
|
| 6 |
H |
0.356 |
|
|
|
| 7 |
H |
0.384 |
|
|
|
| 8 |
H |
0.356 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.625 |
-4.309 |
0.000 |
4.605 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.387 |
1.646 |
0.000 |
| y |
1.646 |
-24.548 |
0.000 |
| z |
0.000 |
0.000 |
-16.994 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.616 |
1.646 |
0.000 |
| y |
1.646 |
-3.357 |
0.000 |
| z |
0.000 |
0.000 |
7.973 |
|
| Polar |
|---|
| 3z2-r2 | 15.947 |
|---|
| x2-y2 | -0.839 |
|---|
| xy | 1.646 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.230 |
0.094 |
0.000 |
| y |
0.094 |
4.362 |
0.000 |
| z |
0.000 |
0.000 |
4.025 |
<r2> (average value of r
2) Å
2
| <r2> |
67.387 |
| (<r2>)1/2 |
8.209 |
Jump to
S1C1
S1C2
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -224.039707 |
| Energy at 298.15K | |
| HF Energy | -224.039707 |
| Nuclear repulsion energy | 125.475992 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
4009 |
3625 |
61.69 |
|
|
|
| 2 |
A1 |
3879 |
3508 |
7.90 |
|
|
|
| 3 |
A1 |
1954 |
1767 |
659.55 |
|
|
|
| 4 |
A1 |
1810 |
1637 |
0.20 |
|
|
|
| 5 |
A1 |
1267 |
1146 |
2.43 |
|
|
|
| 6 |
A1 |
1045 |
945 |
12.23 |
|
|
|
| 7 |
A1 |
520 |
471 |
2.82 |
|
|
|
| 8 |
A2 |
430 |
389 |
0.00 |
|
|
|
| 9 |
A2 |
431i |
389i |
0.00 |
|
|
|
| 10 |
B1 |
869 |
786 |
33.59 |
|
|
|
| 11 |
B1 |
610 |
552 |
9.30 |
|
|
|
| 12 |
B1 |
271i |
245i |
642.28 |
|
|
|
| 13 |
B2 |
4006 |
3623 |
31.39 |
|
|
|
| 14 |
B2 |
3871 |
3501 |
77.84 |
|
|
|
| 15 |
B2 |
1804 |
1631 |
288.64 |
|
|
|
| 16 |
B2 |
1543 |
1396 |
298.87 |
|
|
|
| 17 |
B2 |
1098 |
993 |
14.69 |
|
|
|
| 18 |
B2 |
619 |
559 |
19.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14315.6 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 12947.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.143 |
| O2 |
0.000 |
0.000 |
1.339 |
| N3 |
0.000 |
1.148 |
-0.588 |
| N4 |
0.000 |
-1.148 |
-0.588 |
| H5 |
0.000 |
2.003 |
-0.091 |
| H6 |
0.000 |
1.170 |
-1.576 |
| H7 |
0.000 |
-2.003 |
-0.091 |
| H8 |
0.000 |
-1.170 |
-1.576 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.1957 | 1.3612 | 1.3612 | 2.0170 | 2.0800 | 2.0170 | 2.0800 |
O2 | 1.1957 | | 2.2433 | 2.2433 | 2.4615 | 3.1415 | 2.4615 | 3.1415 | N3 | 1.3612 | 2.2433 | | 2.2959 | 0.9893 | 0.9883 | 3.1902 | 2.5199 | N4 | 1.3612 | 2.2433 | 2.2959 | | 3.1902 | 2.5199 | 0.9893 | 0.9883 | H5 | 2.0170 | 2.4615 | 0.9893 | 3.1902 | | 1.7028 | 4.0067 | 3.5038 | H6 | 2.0800 | 3.1415 | 0.9883 | 2.5199 | 1.7028 | | 3.5038 | 2.3403 | H7 | 2.0170 | 2.4615 | 3.1902 | 0.9893 | 4.0067 | 3.5038 | | 1.7028 | H8 | 2.0800 | 3.1415 | 2.5199 | 0.9883 | 3.5038 | 2.3403 | 1.7028 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.335 |
|
C1 |
N3 |
H6 |
123.797 |
| C1 |
N4 |
H7 |
117.335 |
|
C1 |
N4 |
H8 |
123.797 |
| O2 |
C1 |
N3 |
122.509 |
|
O2 |
C1 |
N4 |
122.509 |
| N3 |
C1 |
N4 |
114.982 |
|
H5 |
N3 |
H6 |
118.868 |
| H7 |
N4 |
H8 |
118.868 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.853 |
|
|
|
| 2 |
O |
-0.526 |
|
|
|
| 3 |
N |
-0.925 |
|
|
|
| 4 |
N |
-0.925 |
|
|
|
| 5 |
H |
0.394 |
|
|
|
| 6 |
H |
0.368 |
|
|
|
| 7 |
H |
0.394 |
|
|
|
| 8 |
H |
0.368 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.627 |
4.627 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.729 |
0.000 |
0.000 |
| y |
0.000 |
-16.286 |
0.000 |
| z |
0.000 |
0.000 |
-24.093 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.540 |
0.000 |
0.000 |
| y |
0.000 |
8.625 |
0.000 |
| z |
0.000 |
0.000 |
-3.086 |
|
| Polar |
|---|
| 3z2-r2 | -6.171 |
|---|
| x2-y2 | -9.443 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.177 |
0.000 |
0.000 |
| y |
0.000 |
4.005 |
0.000 |
| z |
0.000 |
0.000 |
4.292 |
<r2> (average value of r
2) Å
2
| <r2> |
67.276 |
| (<r2>)1/2 |
8.202 |
