Jump to
S1C2
S1C3
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -225.073125 |
| Energy at 298.15K | -225.079347 |
| HF Energy | -225.073125 |
| Nuclear repulsion energy | 124.074413 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3734 |
3579 |
23.38 |
|
|
|
| 2 |
A |
3621 |
3471 |
1.92 |
|
|
|
| 3 |
A |
1860 |
1783 |
421.71 |
|
|
|
| 4 |
A |
1679 |
1609 |
0.00 |
|
|
|
| 5 |
A |
1203 |
1153 |
3.47 |
|
|
|
| 6 |
A |
973 |
933 |
7.70 |
|
|
|
| 7 |
A |
602 |
577 |
90.22 |
|
|
|
| 8 |
A |
476 |
456 |
4.44 |
|
|
|
| 9 |
A |
390 |
374 |
78.73 |
|
|
|
| 10 |
B |
3733 |
3578 |
22.68 |
|
|
|
| 11 |
B |
3617 |
3467 |
35.08 |
|
|
|
| 12 |
B |
1682 |
1612 |
221.52 |
|
|
|
| 13 |
B |
1446 |
1386 |
207.42 |
|
|
|
| 14 |
B |
1070 |
1025 |
22.55 |
|
|
|
| 15 |
B |
804 |
770 |
97.94 |
|
|
|
| 16 |
B |
588 |
563 |
132.15 |
|
|
|
| 17 |
B |
555 |
532 |
176.15 |
|
|
|
| 18 |
B |
454 |
436 |
125.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14242.7 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 13651.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.145 |
| O2 |
0.000 |
0.000 |
1.356 |
| N3 |
0.000 |
1.156 |
-0.612 |
| N4 |
0.000 |
-1.156 |
-0.612 |
| H5 |
0.203 |
1.978 |
-0.068 |
| H6 |
0.463 |
1.130 |
-1.506 |
| H7 |
-0.203 |
-1.978 |
-0.068 |
| H8 |
-0.463 |
-1.130 |
-1.506 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2107 | 1.3817 | 1.3817 | 1.9997 | 2.0536 | 1.9997 | 2.0536 |
O2 | 1.2107 | | 2.2821 | 2.2821 | 2.4454 | 3.1115 | 2.4454 | 3.1115 | N3 | 1.3817 | 2.2821 | | 2.3118 | 1.0065 | 1.0072 | 3.1872 | 2.4975 | N4 | 1.3817 | 2.2821 | 2.3118 | | 3.1872 | 2.4975 | 1.0065 | 1.0072 | H5 | 1.9997 | 2.4454 | 1.0065 | 3.1872 | | 1.6901 | 3.9767 | 3.4885 | H6 | 2.0536 | 3.1115 | 1.0072 | 2.4975 | 1.6901 | | 3.4885 | 2.4414 | H7 | 1.9997 | 2.4454 | 3.1872 | 1.0065 | 3.9767 | 3.4885 | | 1.6901 | H8 | 2.0536 | 3.1115 | 2.4975 | 1.0072 | 3.4885 | 2.4414 | 1.6901 | |
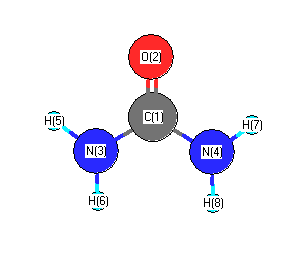 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
112.772 |
|
C1 |
N3 |
H6 |
117.689 |
| C1 |
N4 |
H7 |
112.772 |
|
C1 |
N4 |
H8 |
117.689 |
| O2 |
C1 |
N3 |
123.221 |
|
O2 |
C1 |
N4 |
123.221 |
| N3 |
C1 |
N4 |
113.558 |
|
H5 |
N3 |
H6 |
114.127 |
| H7 |
N4 |
H8 |
114.127 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.584 |
|
|
|
| 2 |
O |
-0.393 |
|
|
|
| 3 |
N |
-0.802 |
|
|
|
| 4 |
N |
-0.802 |
|
|
|
| 5 |
H |
0.362 |
|
|
|
| 6 |
H |
0.345 |
|
|
|
| 7 |
H |
0.362 |
|
|
|
| 8 |
H |
0.345 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.631 |
3.631 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.701 |
3.496 |
0.000 |
| y |
3.496 |
-17.542 |
0.000 |
| z |
0.000 |
0.000 |
-24.804 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.528 |
3.496 |
0.000 |
| y |
3.496 |
7.210 |
0.000 |
| z |
0.000 |
0.000 |
-3.683 |
|
| Polar |
|---|
| 3z2-r2 | -7.366 |
|---|
| x2-y2 | -7.159 |
|---|
| xy | 3.496 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.447 |
0.157 |
0.000 |
| y |
0.157 |
4.699 |
0.000 |
| z |
0.000 |
0.000 |
4.871 |
<r2> (average value of r
2) Å
2
| <r2> |
68.173 |
| (<r2>)1/2 |
8.257 |
Jump to
S1C1
S1C3
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -225.071110 |
| Energy at 298.15K | |
| HF Energy | -225.071110 |
| Nuclear repulsion energy | 124.123509 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3768 |
3612 |
26.22 |
|
|
|
| 2 |
A' |
3649 |
3497 |
7.65 |
|
|
|
| 3 |
A' |
1849 |
1772 |
453.22 |
|
|
|
| 4 |
A' |
1683 |
1613 |
13.74 |
|
|
|
| 5 |
A' |
1193 |
1144 |
2.70 |
|
|
|
| 6 |
A' |
981 |
940 |
8.89 |
|
|
|
| 7 |
A' |
787 |
755 |
28.77 |
|
|
|
| 8 |
A' |
560 |
536 |
36.16 |
|
|
|
| 9 |
A' |
483 |
463 |
14.16 |
|
|
|
| 10 |
A' |
368 |
353 |
441.30 |
|
|
|
| 11 |
A" |
3765 |
3609 |
29.59 |
|
|
|
| 12 |
A" |
3639 |
3488 |
35.38 |
|
|
|
| 13 |
A" |
1671 |
1601 |
245.00 |
|
|
|
| 14 |
A" |
1451 |
1390 |
213.89 |
|
|
|
| 15 |
A" |
1028 |
985 |
18.19 |
|
|
|
| 16 |
A" |
576 |
552 |
22.73 |
|
|
|
| 17 |
A" |
407 |
390 |
56.39 |
|
|
|
| 18 |
A" |
95i |
91i |
49.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13881.0 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 13304.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.008 |
0.141 |
0.000 |
| O2 |
0.034 |
1.353 |
0.000 |
| N3 |
0.034 |
-0.600 |
1.159 |
| N4 |
0.034 |
-0.600 |
-1.159 |
| H5 |
-0.146 |
-0.073 |
1.995 |
| H6 |
-0.251 |
-1.564 |
1.172 |
| H7 |
-0.146 |
-0.073 |
-1.995 |
| H8 |
-0.251 |
-1.564 |
-1.172 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2123 | 1.3761 | 1.3761 | 2.0122 | 2.0852 | 2.0122 | 2.0852 |
O2 | 1.2123 | | 2.2715 | 2.2715 | 2.4591 | 3.1566 | 2.4591 | 3.1566 | N3 | 1.3761 | 2.2715 | | 2.3176 | 1.0045 | 1.0047 | 3.2024 | 2.5381 | N4 | 1.3761 | 2.2715 | 2.3176 | | 3.2024 | 2.5381 | 1.0045 | 1.0047 | H5 | 2.0122 | 2.4591 | 1.0045 | 3.2024 | | 1.7057 | 3.9896 | 3.5016 | H6 | 2.0852 | 3.1566 | 1.0047 | 2.5381 | 1.7057 | | 3.5016 | 2.3440 | H7 | 2.0122 | 2.4591 | 3.2024 | 1.0045 | 3.9896 | 3.5016 | | 1.7057 | H8 | 2.0852 | 3.1566 | 2.5381 | 1.0047 | 3.5016 | 2.3440 | 1.7057 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
114.501 |
|
C1 |
N3 |
H6 |
121.506 |
| C1 |
N4 |
H7 |
114.501 |
|
C1 |
N4 |
H8 |
121.506 |
| O2 |
C1 |
N3 |
122.577 |
|
O2 |
C1 |
N4 |
122.577 |
| N3 |
C1 |
N4 |
114.730 |
|
H5 |
N3 |
H6 |
116.194 |
| H7 |
N4 |
H8 |
116.194 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.623 |
|
|
|
| 2 |
O |
-0.406 |
|
|
|
| 3 |
N |
-0.814 |
|
|
|
| 4 |
N |
-0.814 |
|
|
|
| 5 |
H |
0.366 |
|
|
|
| 6 |
H |
0.339 |
|
|
|
| 7 |
H |
0.366 |
|
|
|
| 8 |
H |
0.339 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.617 |
-4.063 |
0.000 |
4.373 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.411 |
1.688 |
0.000 |
| y |
1.688 |
-24.001 |
0.000 |
| z |
0.000 |
0.000 |
-16.840 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.991 |
1.688 |
0.000 |
| y |
1.688 |
-2.876 |
0.000 |
| z |
0.000 |
0.000 |
7.866 |
|
| Polar |
|---|
| 3z2-r2 | 15.733 |
|---|
| x2-y2 | -1.410 |
|---|
| xy | 1.688 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.357 |
0.114 |
0.000 |
| y |
0.114 |
4.850 |
0.000 |
| z |
0.000 |
0.000 |
4.721 |
<r2> (average value of r
2) Å
2
| <r2> |
68.213 |
| (<r2>)1/2 |
8.259 |
Jump to
S1C1
S1C2
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -225.070691 |
| Energy at 298.15K | |
| HF Energy | -225.070691 |
| Nuclear repulsion energy | 124.235451 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3804 |
3646 |
37.94 |
|
|
|
| 2 |
A1 |
3674 |
3521 |
4.61 |
|
|
|
| 3 |
A1 |
1842 |
1766 |
487.25 |
|
|
|
| 4 |
A1 |
1668 |
1599 |
1.20 |
|
|
|
| 5 |
A1 |
1167 |
1119 |
0.21 |
|
|
|
| 6 |
A1 |
989 |
948 |
7.58 |
|
|
|
| 7 |
A1 |
481 |
461 |
3.41 |
|
|
|
| 8 |
A2 |
375 |
359 |
0.00 |
|
|
|
| 9 |
A2 |
404i |
387i |
0.00 |
|
|
|
| 10 |
B1 |
791 |
758 |
11.35 |
|
|
|
| 11 |
B1 |
578 |
554 |
10.71 |
|
|
|
| 12 |
B1 |
256i |
245i |
563.05 |
|
|
|
| 13 |
B2 |
3801 |
3643 |
36.77 |
|
|
|
| 14 |
B2 |
3665 |
3513 |
52.39 |
|
|
|
| 15 |
B2 |
1662 |
1593 |
309.29 |
|
|
|
| 16 |
B2 |
1455 |
1395 |
219.10 |
|
|
|
| 17 |
B2 |
1002 |
961 |
11.03 |
|
|
|
| 18 |
B2 |
568 |
545 |
14.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13431.5 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 12874.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.142 |
| O2 |
0.000 |
0.000 |
1.356 |
| N3 |
0.000 |
1.154 |
-0.595 |
| N4 |
0.000 |
-1.154 |
-0.595 |
| H5 |
0.000 |
2.017 |
-0.085 |
| H6 |
0.000 |
1.186 |
-1.597 |
| H7 |
0.000 |
-2.017 |
-0.085 |
| H8 |
0.000 |
-1.186 |
-1.597 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2140 | 1.3697 | 1.3697 | 2.0299 | 2.1051 | 2.0299 | 2.1051 |
O2 | 1.2140 | | 2.2671 | 2.2671 | 2.4790 | 3.1824 | 2.4790 | 3.1824 | N3 | 1.3697 | 2.2671 | | 2.3088 | 1.0023 | 1.0023 | 3.2124 | 2.5461 | N4 | 1.3697 | 2.2671 | 2.3088 | | 3.2124 | 2.5461 | 1.0023 | 1.0023 | H5 | 2.0299 | 2.4790 | 1.0023 | 3.2124 | | 1.7252 | 4.0344 | 3.5424 | H6 | 2.1051 | 3.1824 | 1.0023 | 2.5461 | 1.7252 | | 3.5424 | 2.3728 | H7 | 2.0299 | 2.4790 | 3.2124 | 1.0023 | 4.0344 | 3.5424 | | 1.7252 | H8 | 2.1051 | 3.1824 | 2.5461 | 1.0023 | 3.5424 | 2.3728 | 1.7252 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
116.842 |
|
C1 |
N3 |
H6 |
124.390 |
| C1 |
N4 |
H7 |
116.842 |
|
C1 |
N4 |
H8 |
124.390 |
| O2 |
C1 |
N3 |
122.562 |
|
O2 |
C1 |
N4 |
122.562 |
| N3 |
C1 |
N4 |
114.877 |
|
H5 |
N3 |
H6 |
118.768 |
| H7 |
N4 |
H8 |
118.768 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.677 |
|
|
|
| 2 |
O |
-0.419 |
|
|
|
| 3 |
N |
-0.853 |
|
|
|
| 4 |
N |
-0.853 |
|
|
|
| 5 |
H |
0.375 |
|
|
|
| 6 |
H |
0.350 |
|
|
|
| 7 |
H |
0.375 |
|
|
|
| 8 |
H |
0.350 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.383 |
4.383 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.771 |
0.000 |
0.000 |
| y |
0.000 |
-16.106 |
0.000 |
| z |
0.000 |
0.000 |
-23.497 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.970 |
0.000 |
0.000 |
| y |
0.000 |
8.529 |
0.000 |
| z |
0.000 |
0.000 |
-2.559 |
|
| Polar |
|---|
| 3z2-r2 | -5.117 |
|---|
| x2-y2 | -9.666 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.278 |
0.000 |
0.000 |
| y |
0.000 |
4.700 |
0.000 |
| z |
0.000 |
0.000 |
4.791 |
<r2> (average value of r
2) Å
2
| <r2> |
68.126 |
| (<r2>)1/2 |
8.254 |
