Jump to
S1C2
S1C3
Energy calculated at M06-2X/6-311G*
| | hartrees |
|---|
| Energy at 0K | -225.231687 |
| Energy at 298.15K | -225.237959 |
| HF Energy | -225.231687 |
| Nuclear repulsion energy | 123.994037 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3728 |
3728 |
35.76 |
|
|
|
| 2 |
A |
3619 |
3619 |
3.49 |
|
|
|
| 3 |
A |
1875 |
1875 |
461.75 |
|
|
|
| 4 |
A |
1675 |
1675 |
0.16 |
|
|
|
| 5 |
A |
1205 |
1205 |
3.12 |
|
|
|
| 6 |
A |
974 |
974 |
9.09 |
|
|
|
| 7 |
A |
610 |
610 |
112.67 |
|
|
|
| 8 |
A |
482 |
482 |
3.07 |
|
|
|
| 9 |
A |
410 |
410 |
72.24 |
|
|
|
| 10 |
B |
3729 |
3729 |
23.78 |
|
|
|
| 11 |
B |
3615 |
3615 |
51.35 |
|
|
|
| 12 |
B |
1679 |
1679 |
230.76 |
|
|
|
| 13 |
B |
1439 |
1439 |
217.83 |
|
|
|
| 14 |
B |
1074 |
1074 |
22.44 |
|
|
|
| 15 |
B |
808 |
808 |
108.15 |
|
|
|
| 16 |
B |
594 |
594 |
112.02 |
|
|
|
| 17 |
B |
556 |
556 |
233.61 |
|
|
|
| 18 |
B |
471 |
471 |
89.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14270.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 14270.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.148 |
| O2 |
0.000 |
0.000 |
1.355 |
| N3 |
0.000 |
1.157 |
-0.613 |
| N4 |
0.000 |
-1.157 |
-0.613 |
| H5 |
0.201 |
1.982 |
-0.069 |
| H6 |
0.472 |
1.125 |
-1.504 |
| H7 |
-0.201 |
-1.982 |
-0.069 |
| H8 |
-0.472 |
-1.125 |
-1.504 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2072 | 1.3853 | 1.3853 | 2.0038 | 2.0533 | 2.0038 | 2.0533 |
O2 | 1.2072 | | 2.2834 | 2.2834 | 2.4487 | 3.1082 | 2.4487 | 3.1082 | N3 | 1.3853 | 2.2834 | | 2.3150 | 1.0081 | 1.0083 | 3.1925 | 2.4952 | N4 | 1.3853 | 2.2834 | 2.3150 | | 3.1925 | 2.4952 | 1.0081 | 1.0083 | H5 | 2.0038 | 2.4487 | 1.0081 | 3.1925 | | 1.6929 | 3.9841 | 3.4878 | H6 | 2.0533 | 3.1082 | 1.0083 | 2.4952 | 1.6929 | | 3.4878 | 2.4400 | H7 | 2.0038 | 2.4487 | 3.1925 | 1.0081 | 3.9841 | 3.4878 | | 1.6929 | H8 | 2.0533 | 3.1082 | 2.4952 | 1.0083 | 3.4878 | 2.4400 | 1.6929 | |
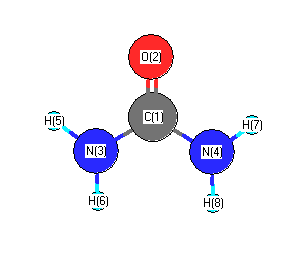 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
112.749 |
|
C1 |
N3 |
H6 |
117.280 |
| C1 |
N4 |
H7 |
112.749 |
|
C1 |
N4 |
H8 |
117.280 |
| O2 |
C1 |
N3 |
123.323 |
|
O2 |
C1 |
N4 |
123.323 |
| N3 |
C1 |
N4 |
113.353 |
|
H5 |
N3 |
H6 |
114.185 |
| H7 |
N4 |
H8 |
114.185 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.618 |
|
|
|
| 2 |
O |
-0.410 |
|
|
|
| 3 |
N |
-0.818 |
|
|
|
| 4 |
N |
-0.818 |
|
|
|
| 5 |
H |
0.365 |
|
|
|
| 6 |
H |
0.349 |
|
|
|
| 7 |
H |
0.365 |
|
|
|
| 8 |
H |
0.349 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.581 |
3.581 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.814 |
3.544 |
0.000 |
| y |
3.544 |
-17.745 |
0.000 |
| z |
0.000 |
0.000 |
-25.044 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.420 |
3.544 |
0.000 |
| y |
3.544 |
7.184 |
0.000 |
| z |
0.000 |
0.000 |
-3.765 |
|
| Polar |
|---|
| 3z2-r2 | -7.530 |
|---|
| x2-y2 | -7.069 |
|---|
| xy | 3.544 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.438 |
0.150 |
0.000 |
| y |
0.150 |
4.595 |
0.000 |
| z |
0.000 |
0.000 |
4.836 |
<r2> (average value of r
2) Å
2
| <r2> |
68.361 |
| (<r2>)1/2 |
8.268 |
Jump to
S1C1
S1C3
Energy calculated at M06-2X/6-311G*
| | hartrees |
|---|
| Energy at 0K | -225.229503 |
| Energy at 298.15K | |
| HF Energy | -225.229503 |
| Nuclear repulsion energy | 124.061144 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3762 |
3762 |
40.29 |
|
|
|
| 2 |
A' |
3644 |
3644 |
11.96 |
|
|
|
| 3 |
A' |
1860 |
1860 |
499.33 |
|
|
|
| 4 |
A' |
1680 |
1680 |
14.98 |
|
|
|
| 5 |
A' |
1194 |
1194 |
2.64 |
|
|
|
| 6 |
A' |
982 |
982 |
11.31 |
|
|
|
| 7 |
A' |
790 |
790 |
32.78 |
|
|
|
| 8 |
A' |
562 |
562 |
33.71 |
|
|
|
| 9 |
A' |
490 |
490 |
17.16 |
|
|
|
| 10 |
A' |
382 |
382 |
456.14 |
|
|
|
| 11 |
A" |
3760 |
3760 |
32.09 |
|
|
|
| 12 |
A" |
3635 |
3635 |
50.38 |
|
|
|
| 13 |
A" |
1667 |
1667 |
256.13 |
|
|
|
| 14 |
A" |
1444 |
1444 |
226.12 |
|
|
|
| 15 |
A" |
1029 |
1029 |
17.86 |
|
|
|
| 16 |
A" |
584 |
584 |
24.40 |
|
|
|
| 17 |
A" |
414 |
414 |
56.88 |
|
|
|
| 18 |
A" |
20i |
20i |
50.89 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13929.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13929.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.009 |
0.144 |
0.000 |
| O2 |
0.034 |
1.353 |
0.000 |
| N3 |
0.034 |
-0.601 |
1.160 |
| N4 |
0.034 |
-0.601 |
-1.160 |
| H5 |
-0.146 |
-0.075 |
1.998 |
| H6 |
-0.254 |
-1.564 |
1.168 |
| H7 |
-0.146 |
-0.075 |
-1.998 |
| H8 |
-0.254 |
-1.564 |
-1.168 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2092 | 1.3789 | 1.3789 | 2.0161 | 2.0862 | 2.0161 | 2.0862 |
O2 | 1.2092 | | 2.2724 | 2.2724 | 2.4624 | 3.1557 | 2.4624 | 3.1557 | N3 | 1.3789 | 2.2724 | | 2.3199 | 1.0060 | 1.0056 | 3.2068 | 2.5357 | N4 | 1.3789 | 2.2724 | 2.3199 | | 3.2068 | 2.5357 | 1.0060 | 1.0056 | H5 | 2.0161 | 2.4624 | 1.0060 | 3.2068 | | 1.7091 | 3.9965 | 3.5008 | H6 | 2.0862 | 3.1557 | 1.0056 | 2.5357 | 1.7091 | | 3.5008 | 2.3357 | H7 | 2.0161 | 2.4624 | 3.2068 | 1.0060 | 3.9965 | 3.5008 | | 1.7091 | H8 | 2.0862 | 3.1557 | 2.5357 | 1.0056 | 3.5008 | 2.3357 | 1.7091 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
114.519 |
|
C1 |
N3 |
H6 |
121.272 |
| C1 |
N4 |
H7 |
114.519 |
|
C1 |
N4 |
H8 |
121.272 |
| O2 |
C1 |
N3 |
122.676 |
|
O2 |
C1 |
N4 |
122.676 |
| N3 |
C1 |
N4 |
114.538 |
|
H5 |
N3 |
H6 |
116.335 |
| H7 |
N4 |
H8 |
116.335 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.657 |
|
|
|
| 2 |
O |
-0.424 |
|
|
|
| 3 |
N |
-0.831 |
|
|
|
| 4 |
N |
-0.831 |
|
|
|
| 5 |
H |
0.370 |
|
|
|
| 6 |
H |
0.344 |
|
|
|
| 7 |
H |
0.370 |
|
|
|
| 8 |
H |
0.344 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.645 |
-4.048 |
0.000 |
4.369 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.558 |
1.728 |
0.000 |
| y |
1.728 |
-24.196 |
0.000 |
| z |
0.000 |
0.000 |
-17.018 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.951 |
1.728 |
0.000 |
| y |
1.728 |
-2.908 |
0.000 |
| z |
0.000 |
0.000 |
7.859 |
|
| Polar |
|---|
| 3z2-r2 | 15.718 |
|---|
| x2-y2 | -1.363 |
|---|
| xy | 1.728 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.346 |
0.113 |
0.000 |
| y |
0.113 |
4.806 |
0.000 |
| z |
0.000 |
0.000 |
4.614 |
<r2> (average value of r
2) Å
2
| <r2> |
68.382 |
| (<r2>)1/2 |
8.269 |
Jump to
S1C1
S1C2
Energy calculated at M06-2X/6-311G*
| | hartrees |
|---|
| Energy at 0K | -225.229011 |
| Energy at 298.15K | |
| HF Energy | -225.229011 |
| Nuclear repulsion energy | 124.184173 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3809 |
3809 |
56.71 |
|
|
|
| 2 |
A1 |
3681 |
3681 |
7.67 |
|
|
|
| 3 |
A1 |
1850 |
1850 |
539.79 |
|
|
|
| 4 |
A1 |
1665 |
1665 |
2.64 |
|
|
|
| 5 |
A1 |
1168 |
1168 |
0.17 |
|
|
|
| 6 |
A1 |
990 |
990 |
9.96 |
|
|
|
| 7 |
A1 |
489 |
489 |
2.99 |
|
|
|
| 8 |
A2 |
371 |
371 |
0.00 |
|
|
|
| 9 |
A2 |
424i |
424i |
0.00 |
|
|
|
| 10 |
B1 |
793 |
793 |
13.80 |
|
|
|
| 11 |
B1 |
572 |
572 |
13.74 |
|
|
|
| 12 |
B1 |
278i |
278i |
577.14 |
|
|
|
| 13 |
B2 |
3806 |
3806 |
34.67 |
|
|
|
| 14 |
B2 |
3672 |
3672 |
75.09 |
|
|
|
| 15 |
B2 |
1659 |
1659 |
321.38 |
|
|
|
| 16 |
B2 |
1448 |
1448 |
235.89 |
|
|
|
| 17 |
B2 |
1003 |
1003 |
10.28 |
|
|
|
| 18 |
B2 |
576 |
576 |
15.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13424.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13424.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.145 |
| O2 |
0.000 |
0.000 |
1.356 |
| N3 |
0.000 |
1.155 |
-0.596 |
| N4 |
0.000 |
-1.155 |
-0.596 |
| H5 |
0.000 |
2.021 |
-0.087 |
| H6 |
0.000 |
1.181 |
-1.599 |
| H7 |
0.000 |
-2.021 |
-0.087 |
| H8 |
0.000 |
-1.181 |
-1.599 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2111 | 1.3721 | 1.3721 | 2.0339 | 2.1060 | 2.0339 | 2.1060 |
O2 | 1.2111 | | 2.2679 | 2.2679 | 2.4831 | 3.1820 | 2.4831 | 3.1820 | N3 | 1.3721 | 2.2679 | | 2.3100 | 1.0038 | 1.0032 | 3.2160 | 2.5424 | N4 | 1.3721 | 2.2679 | 2.3100 | | 3.2160 | 2.5424 | 1.0038 | 1.0032 | H5 | 2.0339 | 2.4831 | 1.0038 | 3.2160 | | 1.7287 | 4.0411 | 3.5405 | H6 | 2.1060 | 3.1820 | 1.0032 | 2.5424 | 1.7287 | | 3.5405 | 2.3625 | H7 | 2.0339 | 2.4831 | 3.2160 | 1.0038 | 4.0411 | 3.5405 | | 1.7287 | H8 | 2.1060 | 3.1820 | 2.5424 | 1.0032 | 3.5405 | 2.3625 | 1.7287 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
116.898 |
|
C1 |
N3 |
H6 |
124.168 |
| C1 |
N4 |
H7 |
116.898 |
|
C1 |
N4 |
H8 |
124.168 |
| O2 |
C1 |
N3 |
122.671 |
|
O2 |
C1 |
N4 |
122.671 |
| N3 |
C1 |
N4 |
114.659 |
|
H5 |
N3 |
H6 |
118.934 |
| H7 |
N4 |
H8 |
118.934 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.710 |
|
|
|
| 2 |
O |
-0.437 |
|
|
|
| 3 |
N |
-0.872 |
|
|
|
| 4 |
N |
-0.872 |
|
|
|
| 5 |
H |
0.380 |
|
|
|
| 6 |
H |
0.355 |
|
|
|
| 7 |
H |
0.380 |
|
|
|
| 8 |
H |
0.355 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.387 |
4.387 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.930 |
0.000 |
0.000 |
| y |
0.000 |
-16.263 |
0.000 |
| z |
0.000 |
0.000 |
-23.680 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.958 |
0.000 |
0.000 |
| y |
0.000 |
8.542 |
0.000 |
| z |
0.000 |
0.000 |
-2.583 |
|
| Polar |
|---|
| 3z2-r2 | -5.167 |
|---|
| x2-y2 | -9.667 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.273 |
0.000 |
0.000 |
| y |
0.000 |
4.596 |
0.000 |
| z |
0.000 |
0.000 |
4.746 |
<r2> (average value of r
2) Å
2
| <r2> |
68.279 |
| (<r2>)1/2 |
8.263 |
