return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O2 (β–Propiolactone)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-266.581504
Energy at 298.15K-266.586975
HF Energy-265.699044
Nuclear repulsion energy171.693634
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3143 2976 7.65      
2 A' 3137 2971 27.68      
3 A' 1926 1824 412.78      
4 A' 1565 1482 0.79      
5 A' 1498 1419 16.54      
6 A' 1375 1303 16.34      
7 A' 1245 1179 21.63      
8 A' 1127 1068 172.89      
9 A' 1047 992 0.74      
10 A' 979 927 97.26      
11 A' 915 867 4.26      
12 A' 759 719 1.79      
13 A' 503 476 3.88      
14 A" 3219 3049 16.29      
15 A" 3205 3035 3.42      
16 A" 1230 1165 0.08      
17 A" 1190 1127 0.83      
18 A" 1079 1022 3.07      
19 A" 816 773 2.47      
20 A" 523 495 2.56      
21 A" 172 163 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 15325.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 14515.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.41387 0.17544 0.12934

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -1.039 -0.278 0.000
C2 0.000 0.624 0.000
O3 -0.064 1.813 0.000
C4 1.087 -0.448 0.000
H5 1.710 -0.445 0.894
H6 1.710 -0.445 -0.894
C7 -0.106 -1.405 0.000
H8 -0.239 -2.006 -0.899
H9 -0.239 -2.006 0.899

Atom - Atom Distances (Å)
  O1 C2 O3 C4 H5 H6 C7 H8 H9
O11.37612.30782.13282.89622.89621.46372.10602.1060
C21.37611.19131.52642.20592.20592.03192.78952.7895
O32.30781.19132.53743.00773.00773.21903.92783.9278
C42.13281.52642.53741.09001.09001.52902.23422.2342
H52.89622.20593.00771.09001.78792.24053.07422.4973
H62.89622.20593.00771.09001.78792.24052.49733.0742
C71.46372.03193.21901.52902.24052.24051.08931.0893
H82.10602.78953.92782.23423.07422.49731.08931.7978
H92.10602.78953.92782.23422.49733.07421.08931.7978

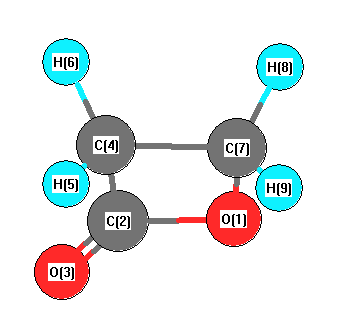
picture of β–Propiolactone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 O3 127.870 O1 C2 C4 94.442
O1 C7 C4 90.878 O1 C7 H8 110.296
O1 C7 H9 110.296 C2 O1 C7 91.312
C2 C4 H5 113.899 C2 C4 H6 113.899
C2 C4 C7 83.367 O3 C2 C4 137.688
C4 C7 H8 116.133 C4 C7 H9 116.133
H5 C4 H6 110.191 H5 C4 C7 116.628
H6 C4 C7 116.628 H8 C7 H9 111.213
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability