return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5NO (Isoxazole, 5-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-284.707415
Energy at 298.15K-284.713799
HF Energy-283.688012
Nuclear repulsion energy223.953619
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3315 3139 0.05      
2 A 3286 3112 1.87      
3 A 3198 3028 6.86      
4 A 3172 3004 6.76      
5 A 3092 2928 11.98      
6 A 1654 1567 25.50      
7 A 1529 1448 11.93      
8 A 1512 1432 11.19      
9 A 1495 1416 21.12      
10 A 1450 1373 2.41      
11 A 1384 1310 10.88      
12 A 1302 1233 14.90      
13 A 1216 1152 10.01      
14 A 1076 1019 1.74      
15 A 1076 1019 3.02      
16 A 1050 994 15.51      
17 A 1018 964 1.81      
18 A 967 916 10.81      
19 A 937 887 9.61      
20 A 849 804 5.82      
21 A 809 766 45.37      
22 A 668 633 0.55      
23 A 643 609 1.37      
24 A 637 603 0.38      
25 A 334 316 2.04      
26 A 245 232 2.68      
27 A 90 85 0.68      

Unscaled Zero Point Vibrational Energy (zpe) 19000.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 17995.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.30750 0.11920 0.08730

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.104 0.016 -0.000
H2 -2.456 -0.520 0.884
H3 -2.544 1.014 -0.001
H4 -2.456 -0.522 -0.883
C5 -0.621 0.116 -0.000
O6 0.064 -1.049 0.000
N7 1.418 -0.799 -0.000
C8 1.532 0.520 0.000
H9 2.524 0.951 0.000
C10 0.267 1.157 0.000
H11 0.044 2.212 -0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 O6 N7 C8 H9 C10 H11
C11.09231.09021.09231.48622.41483.61523.67004.72062.63093.0719
H21.09231.77281.76702.13402.72203.98374.21475.26683.31793.8078
H31.09021.77281.77282.12213.32454.35724.10535.06762.81462.8522
H41.09231.76701.77282.13402.72143.98334.21485.26713.31843.8085
C51.48622.13402.12212.13401.35102.23522.19003.25321.36802.1993
O62.41482.72203.32452.72141.35101.37752.14833.17022.21483.2613
N73.61523.98374.35723.98332.23521.37751.32382.06992.26953.3103
C83.67004.21474.10534.21482.19002.14831.32381.08151.41582.2534
H94.72065.26685.06765.26713.25323.17022.06991.08152.26562.7816
C102.63093.31792.81463.31841.36802.21482.26951.41582.26561.0791
H113.07193.80782.85223.80852.19933.26133.31032.25342.78161.0791

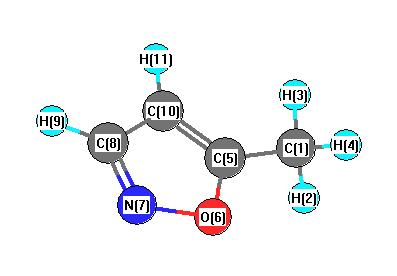
picture of Isoxazole, 5-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 O6 116.587 C1 C5 C10 134.329
H2 C1 H3 108.637 H2 C1 H4 107.972
H2 C1 C5 110.784 H3 C1 H4 108.639
H3 C1 C5 109.958 H4 C1 C5 110.780
C5 O6 N7 110.001 C5 C10 C8 103.741
C5 C10 H11 127.598 O6 C5 C10 109.084
O6 N7 C8 105.347 N7 C8 H9 118.418
N7 C8 C10 111.827 C8 C10 H11 128.662
H9 C8 C10 129.755
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability