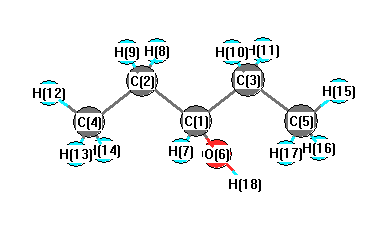
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3828 |
3685 |
10.60 |
|
|
|
| 2 |
A |
3132 |
3015 |
30.50 |
|
|
|
| 3 |
A |
3109 |
2993 |
36.30 |
|
|
|
| 4 |
A |
3103 |
2987 |
75.43 |
|
|
|
| 5 |
A |
3099 |
2984 |
49.99 |
|
|
|
| 6 |
A |
3076 |
2961 |
19.52 |
|
|
|
| 7 |
A |
3067 |
2952 |
1.97 |
|
|
|
| 8 |
A |
3038 |
2925 |
22.95 |
|
|
|
| 9 |
A |
3038 |
2924 |
66.38 |
|
|
|
| 10 |
A |
3030 |
2917 |
25.85 |
|
|
|
| 11 |
A |
3029 |
2916 |
11.88 |
|
|
|
| 12 |
A |
2955 |
2845 |
52.66 |
|
|
|
| 13 |
A |
1527 |
1470 |
9.06 |
|
|
|
| 14 |
A |
1522 |
1465 |
8.02 |
|
|
|
| 15 |
A |
1512 |
1456 |
8.90 |
|
|
|
| 16 |
A |
1509 |
1453 |
8.29 |
|
|
|
| 17 |
A |
1489 |
1434 |
5.52 |
|
|
|
| 18 |
A |
1478 |
1423 |
0.94 |
|
|
|
| 19 |
A |
1446 |
1392 |
8.30 |
|
|
|
| 20 |
A |
1419 |
1366 |
2.73 |
|
|
|
| 21 |
A |
1412 |
1360 |
8.09 |
|
|
|
| 22 |
A |
1404 |
1352 |
14.19 |
|
|
|
| 23 |
A |
1373 |
1322 |
0.29 |
|
|
|
| 24 |
A |
1342 |
1292 |
16.25 |
|
|
|
| 25 |
A |
1315 |
1266 |
1.09 |
|
|
|
| 26 |
A |
1291 |
1243 |
8.69 |
|
|
|
| 27 |
A |
1263 |
1216 |
28.34 |
|
|
|
| 28 |
A |
1176 |
1132 |
25.80 |
|
|
|
| 29 |
A |
1158 |
1114 |
3.06 |
|
|
|
| 30 |
A |
1102 |
1061 |
9.48 |
|
|
|
| 31 |
A |
1075 |
1035 |
25.10 |
|
|
|
| 32 |
A |
1046 |
1007 |
3.57 |
|
|
|
| 33 |
A |
1030 |
992 |
0.67 |
|
|
|
| 34 |
A |
984 |
947 |
54.04 |
|
|
|
| 35 |
A |
941 |
906 |
10.00 |
|
|
|
| 36 |
A |
866 |
834 |
4.81 |
|
|
|
| 37 |
A |
781 |
752 |
1.97 |
|
|
|
| 38 |
A |
769 |
740 |
0.22 |
|
|
|
| 39 |
A |
497 |
479 |
3.24 |
|
|
|
| 40 |
A |
474 |
456 |
9.46 |
|
|
|
| 41 |
A |
397 |
382 |
4.84 |
|
|
|
| 42 |
A |
310 |
298 |
85.22 |
|
|
|
| 43 |
A |
305 |
294 |
22.88 |
|
|
|
| 44 |
A |
240 |
231 |
30.65 |
|
|
|
| 45 |
A |
216 |
208 |
3.12 |
|
|
|
| 46 |
A |
186 |
179 |
0.07 |
|
|
|
| 47 |
A |
106 |
102 |
1.02 |
|
|
|
| 48 |
A |
92 |
89 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36278.5 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 34925.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.023 |
|
|
|
| 2 |
C |
-0.431 |
|
|
|
| 3 |
C |
-0.406 |
|
|
|
| 4 |
C |
-0.655 |
|
|
|
| 5 |
C |
-0.691 |
|
|
|
| 6 |
O |
-0.572 |
|
|
|
| 7 |
H |
0.187 |
|
|
|
| 8 |
H |
0.212 |
|
|
|
| 9 |
H |
0.222 |
|
|
|
| 10 |
H |
0.220 |
|
|
|
| 11 |
H |
0.212 |
|
|
|
| 12 |
H |
0.217 |
|
|
|
| 13 |
H |
0.209 |
|
|
|
| 14 |
H |
0.241 |
|
|
|
| 15 |
H |
0.232 |
|
|
|
| 16 |
H |
0.215 |
|
|
|
| 17 |
H |
0.217 |
|
|
|
| 18 |
H |
0.394 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.144 |
-0.452 |
1.038 |
1.610 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.774 |
2.153 |
0.280 |
| y |
2.153 |
-39.142 |
1.622 |
| z |
0.280 |
1.622 |
-40.272 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.067 |
2.153 |
0.280 |
| y |
2.153 |
0.881 |
1.622 |
| z |
0.280 |
1.622 |
-0.814 |
|
| Polar |
|---|
| 3z2-r2 | -1.627 |
|---|
| x2-y2 | -0.632 |
|---|
| xy | 2.153 |
|---|
| xz | 0.280 |
|---|
| yz | 1.622 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.627 |
0.128 |
0.057 |
| y |
0.128 |
8.630 |
0.144 |
| z |
0.057 |
0.144 |
8.222 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |