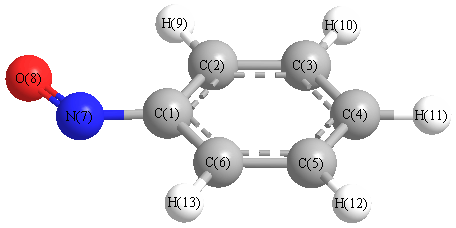
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3395 |
3070 |
3.59 |
82.73 |
0.21 |
0.35 |
| 2 |
A' |
3377 |
3054 |
14.16 |
176.28 |
0.15 |
0.26 |
| 3 |
A' |
3366 |
3044 |
26.51 |
63.91 |
0.27 |
0.43 |
| 4 |
A' |
3356 |
3035 |
14.18 |
105.39 |
0.73 |
0.84 |
| 5 |
A' |
3342 |
3023 |
0.76 |
49.93 |
0.75 |
0.86 |
| 6 |
A' |
1928 |
1744 |
195.63 |
83.58 |
0.31 |
0.47 |
| 7 |
A' |
1791 |
1620 |
4.36 |
47.80 |
0.48 |
0.64 |
| 8 |
A' |
1777 |
1607 |
45.31 |
66.62 |
0.52 |
0.68 |
| 9 |
A' |
1647 |
1490 |
0.98 |
4.49 |
0.30 |
0.46 |
| 10 |
A' |
1612 |
1458 |
17.20 |
2.05 |
0.51 |
0.67 |
| 11 |
A' |
1458 |
1319 |
6.70 |
1.04 |
0.37 |
0.54 |
| 12 |
A' |
1351 |
1222 |
12.60 |
0.16 |
0.74 |
0.85 |
| 13 |
A' |
1299 |
1175 |
25.43 |
25.38 |
0.30 |
0.46 |
| 14 |
A' |
1256 |
1136 |
60.53 |
39.97 |
0.20 |
0.33 |
| 15 |
A' |
1221 |
1105 |
22.39 |
10.89 |
0.74 |
0.85 |
| 16 |
A' |
1165 |
1054 |
0.76 |
0.90 |
0.67 |
0.80 |
| 17 |
A' |
1112 |
1006 |
2.14 |
5.67 |
0.09 |
0.17 |
| 18 |
A' |
1085 |
981 |
1.21 |
34.45 |
0.09 |
0.17 |
| 19 |
A' |
900 |
814 |
21.29 |
7.59 |
0.36 |
0.52 |
| 20 |
A' |
731 |
661 |
15.65 |
3.87 |
0.16 |
0.28 |
| 21 |
A' |
671 |
607 |
0.05 |
7.17 |
0.75 |
0.86 |
| 22 |
A' |
484 |
438 |
0.16 |
2.75 |
0.33 |
0.50 |
| 23 |
A' |
283 |
256 |
2.61 |
2.50 |
0.46 |
0.63 |
| 24 |
A" |
1126 |
1018 |
0.19 |
0.56 |
0.75 |
0.86 |
| 25 |
A" |
1111 |
1005 |
0.02 |
0.12 |
0.75 |
0.86 |
| 26 |
A" |
1069 |
967 |
5.81 |
0.63 |
0.75 |
0.86 |
| 27 |
A" |
955 |
864 |
0.00 |
0.37 |
0.75 |
0.86 |
| 28 |
A" |
853 |
771 |
72.85 |
0.54 |
0.75 |
0.86 |
| 29 |
A" |
755 |
683 |
31.94 |
0.44 |
0.75 |
0.86 |
| 30 |
A" |
523 |
473 |
4.15 |
0.45 |
0.75 |
0.86 |
| 31 |
A" |
460 |
416 |
0.04 |
0.02 |
0.75 |
0.86 |
| 32 |
A" |
274 |
248 |
0.01 |
3.56 |
0.75 |
0.86 |
| 33 |
A" |
113 |
102 |
0.36 |
0.22 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 22923.9 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 20732.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.049 |
|
|
|
| 2 |
C |
-0.158 |
|
|
|
| 3 |
C |
-0.229 |
|
|
|
| 4 |
C |
-0.188 |
|
|
|
| 5 |
C |
-0.229 |
|
|
|
| 6 |
C |
-0.172 |
|
|
|
| 7 |
N |
0.004 |
|
|
|
| 8 |
O |
-0.270 |
|
|
|
| 9 |
H |
0.258 |
|
|
|
| 10 |
H |
0.230 |
|
|
|
| 11 |
H |
0.229 |
|
|
|
| 12 |
H |
0.229 |
|
|
|
| 13 |
H |
0.244 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.138 |
-3.309 |
0.000 |
3.499 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.811 |
2.650 |
0.000 |
| y |
2.650 |
-48.492 |
0.000 |
| z |
0.000 |
0.000 |
-48.708 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.789 |
2.650 |
0.000 |
| y |
2.650 |
-3.232 |
0.000 |
| z |
0.000 |
0.000 |
-3.557 |
|
| Polar |
|---|
| 3z2-r2 | -7.115 |
|---|
| x2-y2 | 6.681 |
|---|
| xy | 2.650 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.895 |
-1.384 |
0.000 |
| y |
-1.384 |
13.346 |
0.000 |
| z |
0.000 |
0.000 |
4.562 |
<r2> (average value of r
2) Å
2
| <r2> |
246.106 |
| (<r2>)1/2 |
15.688 |