return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,4-Pentadiene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-194.745494
Energy at 298.15K-194.753025
HF Energy-193.983385
Nuclear repulsion energy155.432682
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3278 3105 13.18      
2 A 3189 3020 3.75      
3 A 3176 3008 3.76      
4 A 3070 2907 20.77      
5 A 1710 1619 0.72      
6 A 1521 1441 9.13      
7 A 1470 1392 0.00      
8 A 1331 1261 0.16      
9 A 1288 1220 0.01      
10 A 1097 1039 2.07      
11 A 1022 968 19.67      
12 A 934 885 3.09      
13 A 907 859 46.90      
14 A 687 651 12.23      
15 A 376 356 0.32      
16 A 309 293 0.32      
17 A 87 82 0.03      
18 A 3278 3105 16.71      
19 A 3188 3019 10.00      
20 A 3177 3009 21.52      
21 A 3125 2959 11.91      
22 A 1699 1609 14.67      
23 A 1473 1395 5.40      
24 A 1340 1269 0.44      
25 A 1318 1249 1.34      
26 A 1196 1133 3.05      
27 A 1021 967 22.69      
28 A 989 937 3.46      
29 A 910 862 6.25      
30 A 905 857 43.34      
31 A 608 576 14.43      
32 A 459 435 2.12      
33 A 99 93 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 25116.6 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 23787.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.62994 0.07977 0.07915

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.698
C2 0.000 1.238 -0.157
C3 0.000 -1.238 -0.157
C4 1.001 2.124 -0.209
C5 -1.001 -2.124 -0.209
H6 -0.883 -0.009 1.347
H7 0.883 0.009 1.347
H8 -0.877 1.381 -0.788
H9 0.877 -1.381 -0.788
H10 0.958 2.991 -0.859
H11 -0.958 -2.991 -0.859
H12 1.892 2.008 0.402
H13 -1.892 -2.008 0.402

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.50431.50432.51762.51761.09631.09632.20982.20983.50583.50582.77422.7742
C21.50432.47521.33873.50842.14412.13391.08972.83252.11814.39262.11763.7978
C31.50432.47523.50841.33872.13392.14412.83251.08974.39262.11813.79782.1176
C42.51761.33873.50844.69733.24412.62902.10163.55501.08455.51671.08625.0811
C52.51763.50841.33874.69732.62903.24413.55502.10165.51671.08455.08111.0862
H61.09632.14412.13393.24412.62901.76662.54753.08874.15453.71073.55792.4300
H71.09632.13392.14412.62903.24411.76663.08872.54753.71074.15452.43003.5579
H82.20981.08972.83252.10163.55502.54753.08873.27172.44284.37353.07823.7320
H92.20982.83251.08973.55502.10163.08872.54753.27174.37352.44283.73203.0782
H103.50582.11814.39261.08455.51674.15453.71072.44284.37356.28221.85215.8909
H113.50584.39262.11815.51671.08453.71074.15454.37352.44286.28225.89091.8521
H122.77422.11763.79781.08625.08113.55792.43003.07823.73201.85215.89095.5169
H132.77423.79782.11765.08111.08622.43003.55793.73203.07825.89091.85215.5169

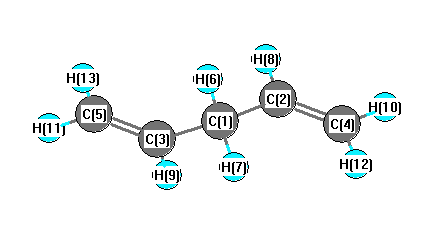
picture of 1,4-Pentadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 124.539 C1 C2 H8 115.921
C1 C3 C5 124.539 C1 C3 H9 115.921
C2 C1 C3 110.713 C2 C1 H6 110.077
C2 C1 H7 109.276 C2 C4 H10 121.528
C2 C4 H12 121.344 C3 C1 H6 109.276
C3 C1 H7 110.077 C3 C5 H11 121.528
C3 C5 H13 121.344 C4 C2 H8 119.518
C5 C3 H9 119.518 H6 C1 H7 107.360
H10 C4 H12 117.128 H11 C5 H13 117.128
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-311G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.476      
2 C -0.184      
3 C -0.184      
4 C -0.450      
5 C -0.450      
6 H 0.225      
7 H 0.225      
8 H 0.212      
9 H 0.212      
10 H 0.221      
11 H 0.221      
12 H 0.213      
13 H 0.213      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.257 2.698 0.000
y 2.698 9.711 0.000
z 0.000 0.000 5.900


<r2> (average value of r2) Å2
<r2> 156.980
(<r2>)1/2 12.529