Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -615.866234 |
| Energy at 298.15K | -615.873328 |
| HF Energy | -615.061614 |
| Nuclear repulsion energy | 205.896271 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3215 |
3045 |
6.66 |
|
|
|
| 2 |
A' |
3171 |
3003 |
23.05 |
|
|
|
| 3 |
A' |
3167 |
3000 |
6.28 |
|
|
|
| 4 |
A' |
3111 |
2947 |
18.95 |
|
|
|
| 5 |
A' |
3074 |
2911 |
23.71 |
|
|
|
| 6 |
A' |
1751 |
1659 |
0.31 |
|
|
|
| 7 |
A' |
1535 |
1454 |
14.94 |
|
|
|
| 8 |
A' |
1511 |
1431 |
4.28 |
|
|
|
| 9 |
A' |
1454 |
1377 |
6.31 |
|
|
|
| 10 |
A' |
1381 |
1308 |
25.25 |
|
|
|
| 11 |
A' |
1348 |
1277 |
3.50 |
|
|
|
| 12 |
A' |
1330 |
1260 |
0.26 |
|
|
|
| 13 |
A' |
1150 |
1089 |
0.02 |
|
|
|
| 14 |
A' |
1080 |
1023 |
9.38 |
|
|
|
| 15 |
A' |
940 |
890 |
9.09 |
|
|
|
| 16 |
A' |
755 |
715 |
17.76 |
|
|
|
| 17 |
A' |
600 |
568 |
2.36 |
|
|
|
| 18 |
A' |
342 |
324 |
0.66 |
|
|
|
| 19 |
A' |
181 |
172 |
0.62 |
|
|
|
| 20 |
A" |
3167 |
3000 |
5.62 |
|
|
|
| 21 |
A" |
3149 |
2982 |
17.14 |
|
|
|
| 22 |
A" |
1519 |
1439 |
9.64 |
|
|
|
| 23 |
A" |
1240 |
1175 |
4.54 |
|
|
|
| 24 |
A" |
1084 |
1026 |
0.13 |
|
|
|
| 25 |
A" |
983 |
931 |
45.25 |
|
|
|
| 26 |
A" |
974 |
922 |
2.73 |
|
|
|
| 27 |
A" |
714 |
676 |
0.21 |
|
|
|
| 28 |
A" |
237 |
224 |
5.87 |
|
|
|
| 29 |
A" |
195 |
184 |
1.00 |
|
|
|
| 30 |
A" |
102 |
96 |
1.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22228.8 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 21052.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.316 |
0.297 |
0.000 |
| C2 |
0.000 |
1.012 |
0.000 |
| C3 |
1.220 |
0.465 |
0.000 |
| C4 |
2.491 |
1.259 |
0.000 |
| Cl5 |
-1.193 |
-1.482 |
0.000 |
| H6 |
-1.897 |
0.571 |
0.883 |
| H7 |
-1.897 |
0.571 |
-0.883 |
| H8 |
-0.098 |
2.098 |
0.000 |
| H9 |
1.315 |
-0.618 |
0.000 |
| H10 |
2.292 |
2.333 |
0.000 |
| H11 |
3.098 |
1.023 |
0.879 |
| H12 |
3.098 |
1.023 |
-0.879 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4977 | 2.5418 | 3.9267 | 1.7831 | 1.0918 | 1.0918 | 2.1747 | 2.7861 | 4.1431 | 4.5587 | 4.5587 |
C2 | 1.4977 | | 1.3371 | 2.5030 | 2.7642 | 2.1380 | 2.1380 | 1.0913 | 2.0944 | 2.6457 | 3.2199 | 3.2199 | C3 | 2.5418 | 1.3371 | | 1.4985 | 3.1003 | 3.2411 | 3.2411 | 2.0992 | 1.0871 | 2.1539 | 2.1470 | 2.1470 | C4 | 3.9267 | 2.5030 | 1.4985 | | 4.5915 | 4.5279 | 4.5279 | 2.7215 | 2.2149 | 1.0920 | 1.0939 | 1.0939 | Cl5 | 1.7831 | 2.7642 | 3.1003 | 4.5915 | | 2.3431 | 2.3431 | 3.7441 | 2.6527 | 5.1670 | 5.0455 | 5.0455 | H6 | 1.0918 | 2.1380 | 3.2411 | 4.5279 | 2.3431 | | 1.7665 | 2.5194 | 3.5370 | 4.6291 | 5.0145 | 5.3152 | H7 | 1.0918 | 2.1380 | 3.2411 | 4.5279 | 2.3431 | 1.7665 | | 2.5194 | 3.5370 | 4.6291 | 5.3152 | 5.0145 | H8 | 2.1747 | 1.0913 | 2.0992 | 2.7215 | 3.7441 | 2.5194 | 2.5194 | | 3.0624 | 2.4017 | 3.4844 | 3.4844 | H9 | 2.7861 | 2.0944 | 1.0871 | 2.2149 | 2.6527 | 3.5370 | 3.5370 | 3.0624 | | 3.1085 | 2.5775 | 2.5775 | H10 | 4.1431 | 2.6457 | 2.1539 | 1.0920 | 5.1670 | 4.6291 | 4.6291 | 2.4017 | 3.1085 | | 1.7710 | 1.7710 | H11 | 4.5587 | 3.2199 | 2.1470 | 1.0939 | 5.0455 | 5.0145 | 5.3152 | 3.4844 | 2.5775 | 1.7710 | | 1.7581 | H12 | 4.5587 | 3.2199 | 2.1470 | 1.0939 | 5.0455 | 5.3152 | 5.0145 | 3.4844 | 2.5775 | 1.7710 | 1.7581 | |
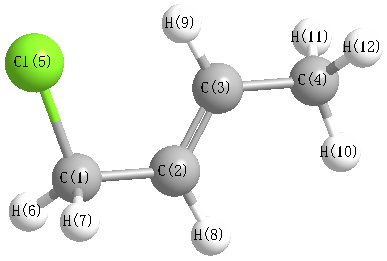 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.352 |
|
C1 |
C2 |
H8 |
113.348 |
| C2 |
C1 |
Cl5 |
114.537 |
|
C2 |
C1 |
H6 |
110.318 |
| C2 |
C1 |
H7 |
110.318 |
|
C2 |
C3 |
C4 |
123.845 |
| C2 |
C3 |
H9 |
119.169 |
|
C3 |
C2 |
H8 |
119.300 |
| C3 |
C4 |
H10 |
111.530 |
|
C3 |
C4 |
H11 |
110.863 |
| C3 |
C4 |
H12 |
110.863 |
|
C4 |
C3 |
H9 |
116.985 |
| Cl5 |
C1 |
H6 |
106.696 |
|
Cl5 |
C1 |
H7 |
106.696 |
| H6 |
C1 |
H7 |
107.993 |
|
H10 |
C4 |
H11 |
108.233 |
| H10 |
C4 |
H12 |
108.233 |
|
H11 |
C4 |
H12 |
106.951 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -615.868661 |
| Energy at 298.15K | -615.875740 |
| HF Energy | -615.064388 |
| Nuclear repulsion energy | 201.768615 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3202 |
3033 |
16.13 |
|
|
|
| 2 |
A |
3189 |
3020 |
5.34 |
|
|
|
| 3 |
A |
3182 |
3014 |
22.28 |
|
|
|
| 4 |
A |
3168 |
3000 |
2.71 |
|
|
|
| 5 |
A |
3150 |
2983 |
14.96 |
|
|
|
| 6 |
A |
3127 |
2962 |
18.65 |
|
|
|
| 7 |
A |
3075 |
2912 |
20.39 |
|
|
|
| 8 |
A |
1739 |
1647 |
7.81 |
|
|
|
| 9 |
A |
1532 |
1451 |
16.10 |
|
|
|
| 10 |
A |
1518 |
1438 |
10.39 |
|
|
|
| 11 |
A |
1514 |
1434 |
5.10 |
|
|
|
| 12 |
A |
1452 |
1376 |
4.77 |
|
|
|
| 13 |
A |
1383 |
1310 |
18.71 |
|
|
|
| 14 |
A |
1352 |
1280 |
5.78 |
|
|
|
| 15 |
A |
1331 |
1261 |
16.84 |
|
|
|
| 16 |
A |
1244 |
1178 |
2.98 |
|
|
|
| 17 |
A |
1157 |
1096 |
0.82 |
|
|
|
| 18 |
A |
1129 |
1069 |
2.79 |
|
|
|
| 19 |
A |
1083 |
1026 |
0.48 |
|
|
|
| 20 |
A |
990 |
938 |
42.59 |
|
|
|
| 21 |
A |
976 |
925 |
12.66 |
|
|
|
| 22 |
A |
907 |
859 |
2.24 |
|
|
|
| 23 |
A |
807 |
764 |
25.49 |
|
|
|
| 24 |
A |
728 |
689 |
31.85 |
|
|
|
| 25 |
A |
502 |
475 |
1.58 |
|
|
|
| 26 |
A |
363 |
344 |
2.04 |
|
|
|
| 27 |
A |
282 |
267 |
2.17 |
|
|
|
| 28 |
A |
199 |
188 |
1.18 |
|
|
|
| 29 |
A |
158 |
150 |
1.82 |
|
|
|
| 30 |
A |
86 |
81 |
0.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22262.2 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 21084.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.786 |
0.832 |
0.097 |
| C2 |
0.510 |
0.196 |
0.457 |
| C3 |
1.588 |
0.230 |
-0.338 |
| C4 |
2.911 |
-0.384 |
-0.003 |
| Cl5 |
-2.078 |
-0.406 |
-0.098 |
| H6 |
-1.137 |
1.508 |
0.877 |
| H7 |
-0.714 |
1.372 |
-0.846 |
| H8 |
0.552 |
-0.315 |
1.418 |
| H9 |
1.510 |
0.736 |
-1.301 |
| H10 |
2.886 |
-0.872 |
0.974 |
| H11 |
3.194 |
-1.130 |
-0.751 |
| H12 |
3.703 |
0.371 |
0.011 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4877 | 2.4876 | 3.8939 | 1.8001 | 1.0904 | 1.0898 | 2.2021 | 2.6899 | 4.1418 | 4.5181 | 4.5132 |
C2 | 1.4877 | | 1.3400 | 2.5134 | 2.7146 | 2.1469 | 2.1400 | 1.0890 | 2.0934 | 2.6558 | 3.2290 | 3.2286 | C3 | 2.4876 | 1.3400 | | 1.4972 | 3.7288 | 3.2459 | 2.6195 | 2.1103 | 1.0906 | 2.1490 | 2.1451 | 2.1478 | C4 | 3.8939 | 2.5134 | 1.4972 | | 4.9907 | 4.5549 | 4.1159 | 2.7554 | 2.2145 | 1.0916 | 1.0939 | 1.0939 | Cl5 | 1.8001 | 2.7146 | 3.7288 | 4.9907 | | 2.3454 | 2.3637 | 3.0369 | 3.9531 | 5.0997 | 5.3619 | 5.8339 | H6 | 1.0904 | 2.1469 | 3.2459 | 4.5549 | 2.3454 | | 1.7799 | 2.5429 | 3.5139 | 4.6751 | 5.3266 | 5.0464 | H7 | 1.0898 | 2.1400 | 2.6195 | 4.1159 | 2.3637 | 1.7799 | | 3.0944 | 2.3573 | 4.6159 | 4.6417 | 4.6089 | H8 | 2.2021 | 1.0890 | 2.1103 | 2.7554 | 3.0369 | 2.5429 | 3.0944 | | 3.0682 | 2.4405 | 3.5149 | 3.5182 | H9 | 2.6899 | 2.0934 | 1.0906 | 2.2145 | 3.9531 | 3.5139 | 2.3573 | 3.0682 | | 3.1065 | 2.5731 | 2.5810 | H10 | 4.1418 | 2.6558 | 2.1490 | 1.0916 | 5.0997 | 4.6751 | 4.6159 | 2.4405 | 3.1065 | | 1.7713 | 1.7715 | H11 | 4.5181 | 3.2290 | 2.1451 | 1.0939 | 5.3619 | 5.3266 | 4.6417 | 3.5149 | 2.5731 | 1.7713 | | 1.7589 | H12 | 4.5132 | 3.2286 | 2.1478 | 1.0939 | 5.8339 | 5.0464 | 4.6089 | 3.5182 | 2.5810 | 1.7715 | 1.7589 | |
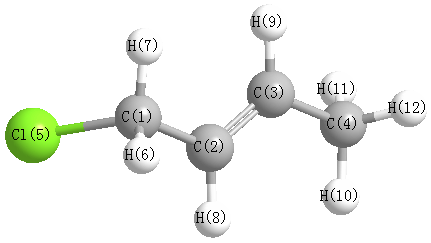 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.131 |
|
C1 |
C2 |
H8 |
116.587 |
| C2 |
C1 |
Cl5 |
110.953 |
|
C2 |
C1 |
H6 |
111.844 |
| C2 |
C1 |
H7 |
111.323 |
|
C2 |
C3 |
C4 |
124.626 |
| C2 |
C3 |
H9 |
118.556 |
|
C3 |
C2 |
H8 |
120.282 |
| C3 |
C4 |
H10 |
111.259 |
|
C3 |
C4 |
H11 |
110.801 |
| C3 |
C4 |
H12 |
111.025 |
|
C4 |
C3 |
H9 |
116.818 |
| Cl5 |
C1 |
H6 |
105.838 |
|
Cl5 |
C1 |
H7 |
107.181 |
| H6 |
C1 |
H7 |
109.454 |
|
H10 |
C4 |
H11 |
108.284 |
| H10 |
C4 |
H12 |
108.303 |
|
H11 |
C4 |
H12 |
107.015 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
