Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -616.557082 |
| Energy at 298.15K | -616.564167 |
| HF Energy | -616.295179 |
| Nuclear repulsion energy | 205.188493 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3191 |
3191 |
9.61 |
|
|
|
| 2 |
A' |
3143 |
3143 |
22.10 |
|
|
|
| 3 |
A' |
3132 |
3132 |
17.80 |
|
|
|
| 4 |
A' |
3095 |
3095 |
21.61 |
|
|
|
| 5 |
A' |
3051 |
3051 |
30.76 |
|
|
|
| 6 |
A' |
1755 |
1755 |
0.75 |
|
|
|
| 7 |
A' |
1529 |
1529 |
14.13 |
|
|
|
| 8 |
A' |
1505 |
1505 |
2.85 |
|
|
|
| 9 |
A' |
1448 |
1448 |
4.31 |
|
|
|
| 10 |
A' |
1367 |
1367 |
26.20 |
|
|
|
| 11 |
A' |
1349 |
1349 |
10.68 |
|
|
|
| 12 |
A' |
1325 |
1325 |
2.14 |
|
|
|
| 13 |
A' |
1138 |
1138 |
0.12 |
|
|
|
| 14 |
A' |
1063 |
1063 |
10.34 |
|
|
|
| 15 |
A' |
927 |
927 |
9.98 |
|
|
|
| 16 |
A' |
720 |
720 |
22.38 |
|
|
|
| 17 |
A' |
589 |
589 |
5.42 |
|
|
|
| 18 |
A' |
338 |
338 |
1.07 |
|
|
|
| 19 |
A' |
176 |
176 |
0.69 |
|
|
|
| 20 |
A" |
3141 |
3141 |
8.48 |
|
|
|
| 21 |
A" |
3105 |
3105 |
24.27 |
|
|
|
| 22 |
A" |
1515 |
1515 |
8.74 |
|
|
|
| 23 |
A" |
1228 |
1228 |
4.60 |
|
|
|
| 24 |
A" |
1085 |
1085 |
0.35 |
|
|
|
| 25 |
A" |
994 |
994 |
44.00 |
|
|
|
| 26 |
A" |
967 |
967 |
0.00 |
|
|
|
| 27 |
A" |
729 |
729 |
0.45 |
|
|
|
| 28 |
A" |
239 |
239 |
5.47 |
|
|
|
| 29 |
A" |
201 |
201 |
1.35 |
|
|
|
| 30 |
A" |
109 |
109 |
1.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22076.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 22076.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.315 |
0.298 |
0.000 |
| C2 |
0.000 |
1.015 |
0.000 |
| C3 |
1.221 |
0.483 |
0.000 |
| C4 |
2.491 |
1.279 |
0.000 |
| Cl5 |
-1.193 |
-1.501 |
0.000 |
| H6 |
-1.899 |
0.561 |
0.882 |
| H7 |
-1.899 |
0.561 |
-0.882 |
| H8 |
-0.111 |
2.100 |
0.000 |
| H9 |
1.323 |
-0.598 |
0.000 |
| H10 |
2.296 |
2.353 |
0.000 |
| H11 |
3.100 |
1.044 |
0.877 |
| H12 |
3.100 |
1.044 |
-0.877 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4978 | 2.5420 | 3.9298 | 1.8029 | 1.0901 | 1.0901 | 2.1668 | 2.7856 | 4.1550 | 4.5621 | 4.5621 |
C2 | 1.4978 | | 1.3316 | 2.5046 | 2.7851 | 2.1428 | 2.1428 | 1.0899 | 2.0864 | 2.6576 | 3.2215 | 3.2215 | C3 | 2.5420 | 1.3316 | | 1.4989 | 3.1248 | 3.2431 | 3.2431 | 2.0944 | 1.0858 | 2.1573 | 2.1483 | 2.1483 | C4 | 3.9298 | 2.5046 | 1.4989 | | 4.6154 | 4.5349 | 4.5349 | 2.7280 | 2.2106 | 1.0913 | 1.0936 | 1.0936 | Cl5 | 1.8029 | 2.7851 | 3.1248 | 4.6154 | | 2.3510 | 2.3510 | 3.7597 | 2.6736 | 5.1993 | 5.0673 | 5.0673 | H6 | 1.0901 | 2.1428 | 3.2431 | 4.5349 | 2.3510 | | 1.7632 | 2.5185 | 3.5361 | 4.6469 | 5.0223 | 5.3214 | H7 | 1.0901 | 2.1428 | 3.2431 | 4.5349 | 2.3510 | 1.7632 | | 2.5185 | 3.5361 | 4.6469 | 5.3214 | 5.0223 | H8 | 2.1668 | 1.0899 | 2.0944 | 2.7280 | 3.7597 | 2.5185 | 2.5185 | | 3.0550 | 2.4208 | 3.4918 | 3.4918 | H9 | 2.7856 | 2.0864 | 1.0858 | 2.2106 | 2.6736 | 3.5361 | 3.5361 | 3.0550 | | 3.1073 | 2.5733 | 2.5733 | H10 | 4.1550 | 2.6576 | 2.1573 | 1.0913 | 5.1993 | 4.6469 | 4.6469 | 2.4208 | 3.1073 | | 1.7688 | 1.7688 | H11 | 4.5621 | 3.2215 | 2.1483 | 1.0936 | 5.0673 | 5.0223 | 5.3214 | 3.4918 | 2.5733 | 1.7688 | | 1.7548 | H12 | 4.5621 | 3.2215 | 2.1483 | 1.0936 | 5.0673 | 5.3214 | 5.0223 | 3.4918 | 2.5733 | 1.7688 | 1.7548 | |
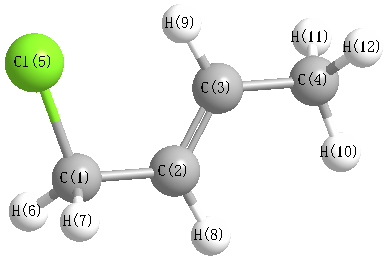 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.811 |
|
C1 |
C2 |
H8 |
112.776 |
| C2 |
C1 |
Cl5 |
114.770 |
|
C2 |
C1 |
H6 |
110.803 |
| C2 |
C1 |
H7 |
110.803 |
|
C2 |
C3 |
C4 |
124.359 |
| C2 |
C3 |
H9 |
118.973 |
|
C3 |
C2 |
H8 |
119.413 |
| C3 |
C4 |
H10 |
111.826 |
|
C3 |
C4 |
H11 |
110.954 |
| C3 |
C4 |
H12 |
110.954 |
|
C4 |
C3 |
H9 |
116.669 |
| Cl5 |
C1 |
H6 |
106.076 |
|
Cl5 |
C1 |
H7 |
106.076 |
| H6 |
C1 |
H7 |
107.944 |
|
H10 |
C4 |
H11 |
108.103 |
| H10 |
C4 |
H12 |
108.103 |
|
H11 |
C4 |
H12 |
106.700 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -616.559761 |
| Energy at 298.15K | -616.566813 |
| HF Energy | -616.297938 |
| Nuclear repulsion energy | 200.995335 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3185 |
3185 |
21.33 |
|
|
|
| 2 |
A |
3169 |
3169 |
5.76 |
|
|
|
| 3 |
A |
3154 |
3154 |
28.93 |
|
|
|
| 4 |
A |
3135 |
3135 |
5.79 |
|
|
|
| 5 |
A |
3116 |
3116 |
20.30 |
|
|
|
| 6 |
A |
3106 |
3106 |
21.83 |
|
|
|
| 7 |
A |
3052 |
3052 |
26.69 |
|
|
|
| 8 |
A |
1739 |
1739 |
12.59 |
|
|
|
| 9 |
A |
1526 |
1526 |
16.13 |
|
|
|
| 10 |
A |
1514 |
1514 |
9.45 |
|
|
|
| 11 |
A |
1511 |
1511 |
4.21 |
|
|
|
| 12 |
A |
1446 |
1446 |
3.28 |
|
|
|
| 13 |
A |
1371 |
1371 |
10.27 |
|
|
|
| 14 |
A |
1351 |
1351 |
2.28 |
|
|
|
| 15 |
A |
1318 |
1318 |
39.04 |
|
|
|
| 16 |
A |
1230 |
1230 |
3.46 |
|
|
|
| 17 |
A |
1145 |
1145 |
0.93 |
|
|
|
| 18 |
A |
1109 |
1109 |
3.17 |
|
|
|
| 19 |
A |
1083 |
1083 |
0.73 |
|
|
|
| 20 |
A |
1000 |
1000 |
40.61 |
|
|
|
| 21 |
A |
970 |
970 |
11.00 |
|
|
|
| 22 |
A |
900 |
900 |
2.88 |
|
|
|
| 23 |
A |
802 |
802 |
13.72 |
|
|
|
| 24 |
A |
686 |
686 |
64.87 |
|
|
|
| 25 |
A |
499 |
499 |
1.69 |
|
|
|
| 26 |
A |
355 |
355 |
3.93 |
|
|
|
| 27 |
A |
281 |
281 |
2.11 |
|
|
|
| 28 |
A |
206 |
206 |
1.08 |
|
|
|
| 29 |
A |
158 |
158 |
2.34 |
|
|
|
| 30 |
A |
88 |
88 |
0.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22101.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 22101.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.774 |
0.839 |
0.088 |
| C2 |
0.521 |
0.207 |
0.454 |
| C3 |
1.594 |
0.219 |
-0.339 |
| C4 |
2.921 |
-0.387 |
0.000 |
| Cl5 |
-2.095 |
-0.408 |
-0.092 |
| H6 |
-1.135 |
1.519 |
0.857 |
| H7 |
-0.716 |
1.363 |
-0.863 |
| H8 |
0.562 |
-0.282 |
1.425 |
| H9 |
1.518 |
0.702 |
-1.312 |
| H10 |
2.909 |
-0.854 |
0.986 |
| H11 |
3.200 |
-1.148 |
-0.734 |
| H12 |
3.713 |
0.367 |
-0.011 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4868 | 2.4852 | 3.8940 | 1.8256 | 1.0881 | 1.0874 | 2.1976 | 2.6893 | 4.1517 | 4.5187 | 4.5128 |
C2 | 1.4868 | | 1.3351 | 2.5138 | 2.7420 | 2.1504 | 2.1455 | 1.0872 | 2.0882 | 2.6666 | 3.2290 | 3.2298 | C3 | 2.4852 | 1.3351 | | 1.4974 | 3.7504 | 3.2508 | 2.6311 | 2.1040 | 1.0891 | 2.1529 | 2.1456 | 2.1489 | C4 | 3.8940 | 2.5138 | 1.4974 | | 5.0167 | 4.5622 | 4.1277 | 2.7572 | 2.2084 | 1.0908 | 1.0936 | 1.0937 | Cl5 | 1.8256 | 2.7420 | 3.7504 | 5.0167 | | 2.3534 | 2.3732 | 3.0624 | 3.9718 | 5.1380 | 5.3849 | 5.8599 | H6 | 1.0881 | 2.1504 | 3.2508 | 4.5622 | 2.3534 | | 1.7772 | 2.5387 | 3.5226 | 4.6900 | 5.3324 | 5.0574 | H7 | 1.0874 | 2.1455 | 2.6311 | 4.1277 | 2.3732 | 1.7772 | | 3.0943 | 2.3729 | 4.6343 | 4.6542 | 4.6192 | H8 | 2.1976 | 1.0872 | 2.1040 | 2.7572 | 3.0624 | 2.5387 | 3.0943 | | 3.0613 | 2.4546 | 3.5167 | 3.5223 | H9 | 2.6893 | 2.0882 | 1.0891 | 2.2084 | 3.9718 | 3.5226 | 2.3729 | 3.0613 | | 3.1045 | 2.5667 | 2.5736 | H10 | 4.1517 | 2.6666 | 2.1529 | 1.0908 | 5.1380 | 4.6900 | 4.6343 | 2.4546 | 3.1045 | | 1.7691 | 1.7693 | H11 | 4.5187 | 3.2290 | 2.1456 | 1.0936 | 5.3849 | 5.3324 | 4.6542 | 3.5167 | 2.5667 | 1.7691 | | 1.7553 | H12 | 4.5128 | 3.2298 | 2.1489 | 1.0937 | 5.8599 | 5.0574 | 4.6192 | 3.5223 | 2.5736 | 1.7693 | 1.7553 | |
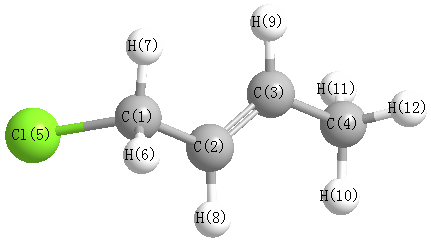 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.364 |
|
C1 |
C2 |
H8 |
116.388 |
| C2 |
C1 |
Cl5 |
111.337 |
|
C2 |
C1 |
H6 |
112.335 |
| C2 |
C1 |
H7 |
111.979 |
|
C2 |
C3 |
C4 |
125.024 |
| C2 |
C3 |
H9 |
118.599 |
|
C3 |
C2 |
H8 |
120.248 |
| C3 |
C4 |
H10 |
111.613 |
|
C3 |
C4 |
H11 |
110.841 |
| C3 |
C4 |
H12 |
111.108 |
|
C4 |
C3 |
H9 |
116.376 |
| Cl5 |
C1 |
H6 |
104.900 |
|
Cl5 |
C1 |
H7 |
106.338 |
| H6 |
C1 |
H7 |
109.552 |
|
H10 |
C4 |
H11 |
108.165 |
| H10 |
C4 |
H12 |
108.183 |
|
H11 |
C4 |
H12 |
106.743 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
