return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (hex-1-ene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-235.151704
Energy at 298.15K-235.164141
HF Energy-234.216465
Nuclear repulsion energy228.374197
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3276 3103 16.59      
2 A 3182 3014 4.92      
3 A 3170 3002 26.05      
4 A 3165 2998 34.94      
5 A 3164 2997 46.94      
6 A 3134 2968 45.07      
7 A 3116 2952 0.23      
8 A 3104 2940 1.26      
9 A 3076 2913 52.88      
10 A 3074 2911 44.39      
11 A 3065 2903 9.40      
12 A 3060 2898 3.74      
13 A 1708 1617 7.76      
14 A 1548 1466 11.66      
15 A 1541 1460 10.00      
16 A 1536 1455 2.79      
17 A 1526 1445 0.94      
18 A 1521 1441 1.75      
19 A 1475 1397 1.97      
20 A 1449 1372 6.02      
21 A 1422 1347 0.30      
22 A 1371 1299 0.36      
23 A 1363 1290 0.57      
24 A 1345 1274 0.24      
25 A 1333 1263 0.79      
26 A 1283 1215 0.06      
27 A 1276 1208 0.12      
28 A 1221 1157 0.32      
29 A 1148 1087 3.89      
30 A 1105 1047 0.18      
31 A 1071 1014 4.52      
32 A 1059 1003 1.93      
33 A 1020 966 20.72      
34 A 964 913 1.17      
35 A 944 894 1.34      
36 A 925 876 1.95      
37 A 903 855 44.72      
38 A 810 767 0.92      
39 A 734 695 3.47      
40 A 652 617 12.79      
41 A 463 439 0.89      
42 A 367 347 0.38      
43 A 363 344 0.34      
44 A 252 239 0.00      
45 A 180 171 0.02      
46 A 129 122 0.05      
47 A 91 87 0.01      
48 A 74 70 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 36878.1 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 34927.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.43613 0.04208 0.04131

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 3.043 -0.248 -0.445
C2 2.031 -0.144 0.425
C3 0.771 0.629 0.176
C4 -0.463 -0.277 0.131
C5 -1.762 0.494 -0.085
C6 -2.986 -0.416 -0.136
H7 3.016 0.258 -1.406
H8 3.926 -0.838 -0.223
H9 2.098 -0.669 1.378
H10 0.634 1.377 0.968
H11 0.857 1.178 -0.768
H12 -0.333 -1.012 -0.672
H13 -0.531 -0.848 1.066
H14 -1.881 1.229 0.720
H15 -1.688 1.066 -1.017
H16 -3.905 0.153 -0.294
H17 -2.898 -1.142 -0.949
H18 -3.095 -0.975 0.798

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.33892.51413.55384.87596.03921.08621.08462.09683.23172.63033.46923.92685.27194.94406.96136.02896.3052
C21.33891.49902.51503.88015.05522.11742.11951.09062.13542.13282.74712.73334.15674.16855.98655.21305.2066
C32.51411.49901.53172.54993.91152.77183.50242.21141.09781.09612.15222.16052.77292.76814.72324.22604.2313
C43.55382.51501.53171.52622.54023.84134.43912.87542.15452.16161.09651.09742.15092.15053.49442.80022.8032
C54.87593.88012.54991.52621.52594.96325.84384.28882.76232.79232.15792.15471.09631.09632.17962.17092.1711
C66.03925.05523.91152.54021.52596.17156.92515.31034.18784.20842.77092.76722.15852.15801.09281.09331.0934
H71.08622.11742.77183.84134.96326.17151.85173.07483.54432.43203.65644.46285.42654.78917.01036.09426.6124
H81.08462.11953.50244.43915.84386.92511.85172.43614.14263.71224.28654.63976.23595.98147.89366.86917.0965
H92.09681.09062.21142.87544.28885.31033.07482.43612.54893.09173.19882.65344.45754.80466.28545.53155.2346
H103.23172.13541.09782.15452.76234.18783.54434.14262.54891.76163.05532.51372.53193.07104.86764.74284.4124
H112.63032.13281.09612.16162.79234.20842.43203.71223.09171.76162.49523.06583.11722.56034.89424.41784.7656
H123.46922.74712.15221.09652.15792.77093.65644.28653.19883.05532.49521.75723.05912.50503.77592.58273.1290
H133.92682.73332.16051.09742.15472.76724.46284.63972.65342.51373.06581.75722.50153.05683.77313.12242.5816
H145.27194.15672.77292.15091.09632.15855.42656.23594.45752.53193.11723.05912.50151.75552.50613.07242.5172
H154.94404.16852.76812.15051.09632.15804.78915.98144.80463.07102.56032.50503.05681.75552.50372.51833.0721
H166.96135.98654.72323.49442.17961.09287.01037.89366.28544.86764.89423.77593.77312.50612.50371.76631.7662
H176.02895.21304.22602.80022.17091.09336.09426.86915.53154.74284.41782.58273.12243.07242.51831.76631.7658
H186.30525.20664.23132.80322.17111.09346.61247.09655.23464.41244.76563.12902.58162.51723.07211.76621.7658

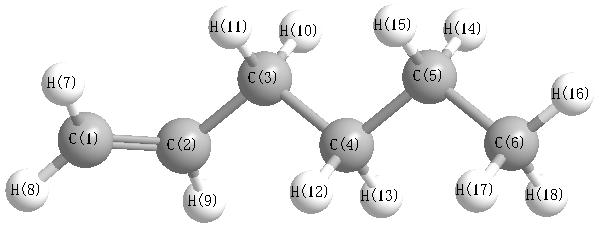
picture of hex-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.625 C1 C2 H9 118.965
C2 C1 H7 121.289 C2 C1 H8 121.630
C2 C3 C4 112.159 C2 C3 H10 109.665
C2 C3 H11 109.561 C3 C2 H9 116.395
C3 C4 C5 112.998 C3 C4 H12 108.815
C3 C4 H13 109.408 C4 C3 H10 108.919
C4 C3 H11 109.571 C4 C5 C6 112.672
C4 C5 H14 109.109 C4 C5 H15 109.075
C5 C4 H12 109.636 C5 C4 H13 109.334
C5 C6 H16 111.610 C5 C6 H17 110.880
C5 C6 H18 110.893 C6 C5 H14 109.722
C6 C5 H15 109.680 H7 C1 H8 117.081
H10 C3 H11 106.819 H12 C4 H13 106.436
H14 C5 H15 106.383 H16 C6 H17 107.796
H16 C6 H18 107.785 H17 C6 H18 107.712
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability