return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (hex-1-ene)

using model chemistry: B2PLYP=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP=FULL/6-311G*
 hartrees
Energy at 0K-235.623499
Energy at 298.15K-235.635907
HF Energy-235.317941
Nuclear repulsion energy227.969367
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3242 3242 24.42      
2 A 3157 3157 6.28      
3 A 3145 3145 34.03      
4 A 3125 3125 47.55      
5 A 3122 3122 74.32      
6 A 3095 3095 51.65      
7 A 3075 3075 2.60      
8 A 3063 3063 7.86      
9 A 3053 3053 41.95      
10 A 3046 3046 70.56      
11 A 3037 3037 3.37      
12 A 3031 3031 11.04      
13 A 1713 1713 11.26      
14 A 1543 1543 9.08      
15 A 1535 1535 8.78      
16 A 1530 1530 2.25      
17 A 1518 1518 0.63      
18 A 1512 1512 2.04      
19 A 1475 1475 1.40      
20 A 1443 1443 3.91      
21 A 1419 1419 0.62      
22 A 1374 1374 0.40      
23 A 1358 1358 0.56      
24 A 1341 1341 0.18      
25 A 1335 1335 0.66      
26 A 1290 1290 1.15      
27 A 1270 1270 0.07      
28 A 1215 1215 0.28      
29 A 1141 1141 2.81      
30 A 1081 1081 0.19      
31 A 1065 1065 6.17      
32 A 1040 1040 0.44      
33 A 1029 1029 16.59      
34 A 956 956 1.43      
35 A 940 940 0.98      
36 A 925 925 44.38      
37 A 914 914 2.00      
38 A 808 808 1.00      
39 A 739 739 3.01      
40 A 652 652 12.19      
41 A 462 462 1.04      
42 A 361 361 0.45      
43 A 360 360 0.24      
44 A 248 248 0.00      
45 A 178 178 0.03      
46 A 127 127 0.05      
47 A 89 89 0.02      
48 A 75 75 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 36625.0 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 36625.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULL/6-311G*
ABC
0.44927 0.04157 0.04082

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 3.074 -0.183 -0.443
C2 2.036 -0.199 0.394
C3 0.773 0.593 0.222
C4 -0.471 -0.296 0.096
C5 -1.769 0.501 -0.041
C6 -3.005 -0.389 -0.163
H7 3.076 0.438 -1.333
H8 3.957 -0.787 -0.269
H9 2.081 -0.840 1.273
H10 0.644 1.261 1.082
H11 0.859 1.235 -0.661
H12 -0.351 -0.957 -0.770
H13 -0.539 -0.952 0.972
H14 -1.879 1.164 0.824
H15 -1.700 1.154 -0.918
H16 -3.917 0.203 -0.264
H17 -2.937 -1.042 -1.037
H18 -3.121 -1.028 0.716

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.33322.51743.58804.90776.08921.08551.08372.08923.21252.63883.52673.95605.28674.98047.00426.10086.3584
C21.33321.50102.52743.89395.07612.11422.11471.08922.13242.13442.76222.74514.16794.18506.00355.24345.2331
C32.51741.50101.53502.55783.92322.78293.50412.20681.09731.09492.15642.16172.77882.78054.73184.24554.2466
C43.58802.52741.53501.52912.54893.89464.47022.86302.15472.16531.09551.09652.15462.15443.50042.81402.8173
C54.90773.89392.55781.52911.52805.01505.87344.28372.76832.79852.15982.15641.09571.09562.18002.17612.1761
C66.08925.07613.92322.54891.52806.24806.97435.30464.19424.22142.78092.77292.15792.15751.09211.09291.0930
H71.08552.11422.78293.89465.01506.24801.84673.06913.52542.44963.74324.50775.45254.84737.07866.19966.6894
H81.08372.11473.50414.47025.87346.97431.84672.42894.12273.71984.34054.66706.24926.01577.93616.94117.1496
H92.08921.08922.20682.86304.28375.30463.06912.42892.55313.08893.17872.63974.46064.80366.27945.52805.2346
H103.21252.13241.09732.15472.76834.19423.52544.12272.55311.75703.05612.51182.53743.08314.87184.75594.4207
H112.63882.13441.09492.16532.79854.22142.44963.71983.08891.75702.50603.06693.11562.57334.90274.44244.7806
H123.52672.76222.15641.09552.15982.78093.74324.34053.17873.05612.50601.75233.06092.50933.78382.60073.1434
H133.95602.74512.16171.09652.15642.77294.50774.66702.63972.51183.06691.75232.50843.05873.77813.12992.5952
H145.28674.16792.77882.15461.09572.15795.45256.24924.46062.53743.11563.06092.50841.75112.50243.07412.5212
H154.98044.18502.78052.15441.09562.15754.84736.01574.80363.08312.57332.50933.05871.75112.49972.52333.0738
H167.00426.00354.73183.50042.18001.09217.07867.93616.27944.87184.90273.78383.77812.50242.49971.76361.7636
H176.10085.24344.24552.81402.17611.09296.19966.94115.52804.75594.44242.60073.12993.07412.52331.76361.7631
H186.35845.23314.24662.81732.17611.09306.68947.14965.23464.42074.78063.14342.59522.52123.07381.76361.7631

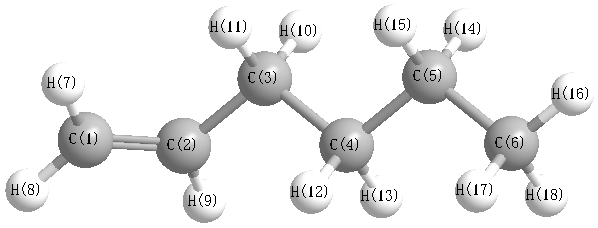
picture of hex-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 125.207 C1 C2 H9 118.841
C2 C1 H7 121.549 C2 C1 H8 121.739
C2 C3 C4 112.706 C2 C3 H10 109.328
C2 C3 H11 109.627 C3 C2 H9 115.947
C3 C4 C5 113.180 C3 C4 H12 108.973
C3 C4 H13 109.329 C4 C3 H10 108.738
C4 C3 H11 109.703 C4 C5 C6 112.970
C4 C5 H14 109.228 C4 C5 H15 109.217
C5 C4 H12 109.648 C5 C4 H13 109.324
C5 C6 H16 111.529 C5 C6 H17 111.165
C5 C6 H18 111.167 C6 C5 H14 109.559
C6 C5 H15 109.541 H7 C1 H8 116.712
H10 C3 H11 106.546 H12 C4 H13 106.144
H14 C5 H15 106.092 H16 C6 H17 107.630
H16 C6 H18 107.626 H17 C6 H18 107.532
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability