return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,5-Hexadiene)

using model chemistry: B2PLYP=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1Ag
Energy calculated at B2PLYP=FULL/6-311G*
 hartrees
Energy at 0K-234.401352
Energy at 298.15K-234.411116
HF Energy-234.099787
Nuclear repulsion energy212.309898
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3243 3243 10.71      
2 A 3158 3158 0.10      
3 A 3147 3147 50.26      
4 A 3103 3103 27.86      
5 A 3036 3036 8.75      
6 A 1711 1711 6.21      
7 A 1512 1512 3.18      
8 A 1472 1472 3.05      
9 A 1386 1386 0.47      
10 A 1338 1338 1.04      
11 A 1265 1265 0.14      
12 A 1105 1105 0.03      
13 A 1046 1046 1.26      
14 A 1029 1029 10.78      
15 A 963 963 0.00      
16 A 927 927 19.41      
17 A 792 792 0.35      
18 A 635 635 7.69      
19 A 422 422 0.01      
20 A 352 352 0.12      
21 A 97 97 0.02      
22 A 76 76 0.02      
23 B 3243 3243 35.95      
24 B 3158 3158 12.64      
25 B 3146 3146 13.86      
26 B 3084 3084 7.03      
27 B 3040 3040 46.49      
28 B 1715 1715 14.76      
29 B 1526 1526 9.79      
30 B 1474 1474 0.29      
31 B 1348 1348 0.70      
32 B 1331 1331 0.03      
33 B 1292 1292 1.46      
34 B 1196 1196 2.11      
35 B 1031 1031 29.83      
36 B 976 976 4.38      
37 B 944 944 0.03      
38 B 928 928 71.31      
39 B 665 665 22.89      
40 B 441 441 1.53      
41 B 222 222 0.25      
42 B 115 115 0.14      

Unscaled Zero Point Vibrational Energy (zpe) 31345.5 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 31345.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULL/6-311G*
ABC
0.42094 0.04670 0.04534

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULL/6-311G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.423 0.644 -0.305
C2 0.423 -0.644 -0.305
C3 0.423 1.884 -0.341
C4 -0.423 -1.884 -0.341
C5 0.451 2.818 0.609
C6 -0.451 -2.818 0.609
H7 -1.088 0.630 -1.177
H8 -1.066 0.655 0.580
H9 1.088 -0.630 -1.177
H10 1.066 -0.655 0.580
H11 1.065 1.999 -1.213
H12 -1.065 -1.999 -1.213
H13 1.091 3.690 0.532
H14 -0.169 2.745 1.496
H15 -1.091 -3.690 0.532
H16 0.169 -2.745 1.496

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.54161.50072.52822.51513.58141.09701.09372.16042.16532.20762.86793.50232.77844.46443.8834
C21.54162.52821.50073.58142.51512.16042.16531.09701.09372.86792.20764.46443.88343.50232.7784
C31.50072.52823.86091.33314.87592.13402.13842.73102.77621.08914.24842.11442.11385.84084.9860
C42.52821.50073.86094.87591.33312.73102.77622.13402.13844.24841.08915.84084.98602.11442.1138
C52.51513.58141.33314.87595.70843.21712.64213.93513.52772.09015.36901.08361.08546.68865.6403
C63.58142.51514.87591.33315.70843.93513.52773.21712.64215.36902.09016.68865.64031.08361.0854
H71.09702.16042.13402.73103.21713.93511.75712.51453.06232.55182.62924.12713.53044.64544.4850
H81.09372.16532.13842.77622.64213.52771.75713.06232.50243.09203.20323.72302.45134.34533.7318
H92.16041.09702.73102.13403.93513.21712.51453.06231.75712.62922.55184.64544.48504.12713.5304
H102.16531.09372.77622.13843.52772.64213.06232.50241.75713.20323.09204.34533.73183.72302.4513
H112.20762.86791.08914.24842.09015.36902.55183.09202.62923.20324.53002.43023.06946.32895.5359
H122.86792.20764.24841.08915.36902.09012.62923.20322.55183.09204.53006.32895.53592.43023.0694
H133.50234.46442.11445.84081.08366.68864.12713.72304.64544.34532.43026.32891.84697.69506.5710
H142.77843.88342.11384.98601.08545.64033.53042.45134.48503.73183.06945.53591.84696.57105.4997
H154.46443.50235.84082.11446.68861.08364.64544.34534.12713.72306.32892.43027.69506.57101.8469
H163.88342.77844.98602.11385.64031.08544.48503.73183.53042.45135.53593.06946.57105.49971.8469

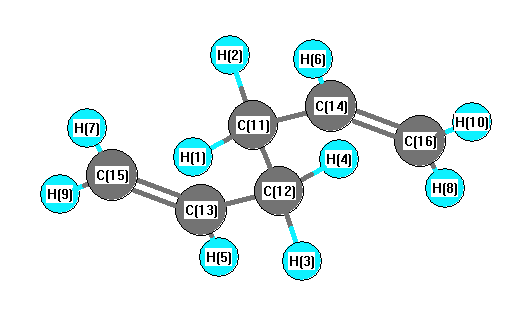
picture of 1,5-Hexadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 112.400 C1 C2 H9 108.760
C1 C2 H10 109.325 C1 C3 C5 125.018
C1 C3 H11 116.041 C2 C1 C3 112.400
C2 C1 H7 108.760 C2 C1 H8 109.325
C2 C4 C6 125.018 C2 C4 H12 116.041
C3 C1 H7 109.490 C3 C1 H8 110.028
C3 C5 H13 121.730 C3 C5 H14 121.521
C4 C2 H9 109.490 C4 C2 H10 110.028
C4 C6 H15 121.730 C4 C6 H16 121.521
C5 C3 H11 118.935 C6 C4 H12 118.935
H7 C1 H8 106.665 H9 C2 H10 106.665
H13 C5 H14 116.748 H15 C6 H16 116.748
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability