return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H8O2S ((Methylsulfonyl)ethane)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-666.677244
Energy at 298.15K-666.687566
HF Energy-665.503177
Nuclear repulsion energy352.796887
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3232 3061 0.78      
2 A 3217 3046 0.77      
3 A 3203 3034 5.70      
4 A 3184 3016 11.55      
5 A 3172 3004 2.96      
6 A 3111 2946 3.15      
7 A 3106 2942 2.41      
8 A 3098 2934 12.56      
9 A 1543 1461 10.74      
10 A 1541 1459 14.34      
11 A 1499 1419 17.74      
12 A 1496 1417 3.76      
13 A 1487 1409 3.99      
14 A 1457 1380 3.77      
15 A 1403 1329 31.03      
16 A 1373 1301 154.27      
17 A 1344 1273 77.00      
18 A 1299 1230 9.22      
19 A 1166 1105 166.22      
20 A 1112 1053 1.94      
21 A 1099 1041 12.25      
22 A 1024 970 6.18      
23 A 1003 950 24.30      
24 A 992 940 10.43      
25 A 822 778 64.19      
26 A 756 716 20.24      
27 A 684 648 13.78      
28 A 504 477 24.01      
29 A 459 434 35.14      
30 A 411 389 7.66      
31 A 325 308 1.19      
32 A 295 280 1.61      
33 A 243 230 2.30      
34 A 219 208 0.80      
35 A 195 185 0.41      
36 A 89 84 2.70      

Unscaled Zero Point Vibrational Energy (zpe) 25580.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 24227.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.13969 0.07979 0.07726

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.254 0.161 0.039
H2 2.253 -0.046 1.109
H3 3.199 -0.191 -0.380
H4 2.211 1.242 -0.105
C5 1.094 -0.549 -0.639
H6 1.156 -1.633 -0.515
H7 1.022 -0.339 -1.709
C8 -0.744 1.569 -0.319
H9 -0.720 1.704 -1.399
H10 -1.736 1.815 0.060
H11 0.002 2.183 0.182
S12 -0.499 -0.153 0.078
O13 -0.374 -0.257 1.527
O14 -1.505 -0.916 -0.649

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 H11 S12 O13 O14
C11.08991.09161.09131.51992.17522.19613.33173.64644.31993.03032.77083.04873.9703
H21.08991.76981.77022.15732.52213.08913.69214.26564.52583.30122.94042.66824.2393
H31.09161.76981.76142.15132.50452.55494.31824.47075.34544.02123.72604.05034.7671
H41.09131.77021.76142.17733.09002.54682.98103.23763.99232.41853.05303.40454.3314
C51.51992.15732.15132.17731.09361.09252.82222.99033.75313.05431.79072.63302.6244
H62.17522.52212.50453.09001.09361.76613.72923.92964.53804.04792.29862.89992.7598
H72.19613.08912.55492.54681.09251.76612.94802.70253.92123.31282.35383.52542.8001
C83.33173.69214.31822.98102.82223.72922.94801.08911.09011.08811.78412.62302.6196
H93.64644.26564.47073.23762.99033.92962.70251.08911.78151.80292.38333.54002.8358
H104.31994.52585.34543.99233.75314.53803.92121.09011.78151.78092.32512.88202.8310
H113.03033.30124.02122.41853.05434.04793.31281.08811.80291.78092.39112.81163.5445
S122.77082.94043.72603.05301.79072.29862.35381.78412.38332.32512.39111.45841.4575
O133.04872.66824.05033.40452.63302.89993.52542.62303.54002.88202.81161.45842.5402
O143.97034.23934.76714.33142.62442.75982.80012.61962.83582.83103.54451.45752.5402

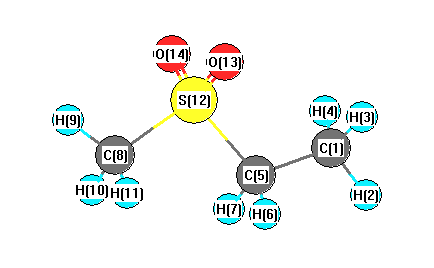
picture of (Methylsulfonyl)ethane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 111.632 C1 C5 H7 113.411
C1 C5 S12 113.383 H2 C1 H3 108.444
H2 C1 H4 108.498 H2 C1 C5 110.414
H3 C1 H4 107.597 H3 C1 C5 109.840
H4 C1 C5 111.940 C5 S12 C8 104.275
C5 S12 O13 107.829 C5 S12 O14 107.351
H6 C5 H7 107.779 H6 C5 S12 103.003
H7 C5 S12 106.944 C8 S12 O13 107.567
C8 S12 O14 107.401 H9 C8 H10 109.663
H9 C8 H11 111.799 H9 C8 S12 109.716
H10 C8 H11 109.682 H10 C8 S12 105.427
H11 C8 S12 110.356 O13 S12 O14 121.195
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability