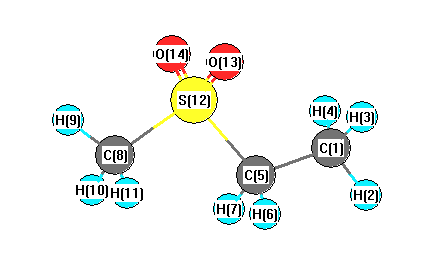
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3204 |
3058 |
2.18 |
|
|
|
| 2 |
A |
3196 |
3050 |
1.02 |
|
|
|
| 3 |
A |
3170 |
3025 |
10.13 |
|
|
|
| 4 |
A |
3149 |
3006 |
6.41 |
|
|
|
| 5 |
A |
3139 |
2995 |
14.11 |
|
|
|
| 6 |
A |
3094 |
2953 |
4.41 |
|
|
|
| 7 |
A |
3091 |
2950 |
2.52 |
|
|
|
| 8 |
A |
3074 |
2934 |
17.79 |
|
|
|
| 9 |
A |
1523 |
1454 |
10.33 |
|
|
|
| 10 |
A |
1520 |
1451 |
13.30 |
|
|
|
| 11 |
A |
1478 |
1410 |
15.22 |
|
|
|
| 12 |
A |
1475 |
1408 |
4.00 |
|
|
|
| 13 |
A |
1467 |
1400 |
4.02 |
|
|
|
| 14 |
A |
1433 |
1368 |
3.20 |
|
|
|
| 15 |
A |
1372 |
1310 |
29.41 |
|
|
|
| 16 |
A |
1350 |
1288 |
146.99 |
|
|
|
| 17 |
A |
1319 |
1259 |
80.61 |
|
|
|
| 18 |
A |
1282 |
1224 |
10.50 |
|
|
|
| 19 |
A |
1148 |
1095 |
167.30 |
|
|
|
| 20 |
A |
1099 |
1049 |
1.88 |
|
|
|
| 21 |
A |
1078 |
1029 |
12.33 |
|
|
|
| 22 |
A |
1007 |
961 |
3.62 |
|
|
|
| 23 |
A |
984 |
940 |
28.33 |
|
|
|
| 24 |
A |
976 |
932 |
12.57 |
|
|
|
| 25 |
A |
805 |
769 |
57.20 |
|
|
|
| 26 |
A |
734 |
701 |
24.73 |
|
|
|
| 27 |
A |
657 |
627 |
12.54 |
|
|
|
| 28 |
A |
496 |
473 |
22.95 |
|
|
|
| 29 |
A |
447 |
426 |
33.89 |
|
|
|
| 30 |
A |
401 |
382 |
8.05 |
|
|
|
| 31 |
A |
315 |
301 |
1.02 |
|
|
|
| 32 |
A |
286 |
273 |
1.64 |
|
|
|
| 33 |
A |
231 |
221 |
2.11 |
|
|
|
| 34 |
A |
209 |
200 |
1.12 |
|
|
|
| 35 |
A |
197 |
188 |
0.14 |
|
|
|
| 36 |
A |
90 |
86 |
2.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25248.9 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 24097.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.675 |
|
|
|
| 2 |
H |
0.270 |
|
|
|
| 3 |
H |
0.244 |
|
|
|
| 4 |
H |
0.231 |
|
|
|
| 5 |
C |
-0.649 |
|
|
|
| 6 |
H |
0.300 |
|
|
|
| 7 |
H |
0.272 |
|
|
|
| 8 |
C |
-0.848 |
|
|
|
| 9 |
H |
0.272 |
|
|
|
| 10 |
H |
0.300 |
|
|
|
| 11 |
H |
0.276 |
|
|
|
| 12 |
S |
1.018 |
|
|
|
| 13 |
O |
-0.507 |
|
|
|
| 14 |
O |
-0.503 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.235 |
3.007 |
-2.337 |
4.997 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.490 |
-5.547 |
-1.381 |
| y |
-5.547 |
-42.081 |
-0.600 |
| z |
-1.381 |
-0.600 |
-48.629 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.865 |
-5.547 |
-1.381 |
| y |
-5.547 |
4.479 |
-0.600 |
| z |
-1.381 |
-0.600 |
-5.343 |
|
| Polar |
|---|
| 3z2-r2 | -10.687 |
|---|
| x2-y2 | -2.410 |
|---|
| xy | -5.547 |
|---|
| xz | -1.381 |
|---|
| yz | -0.600 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
191.062 |
| (<r2>)1/2 |
13.823 |