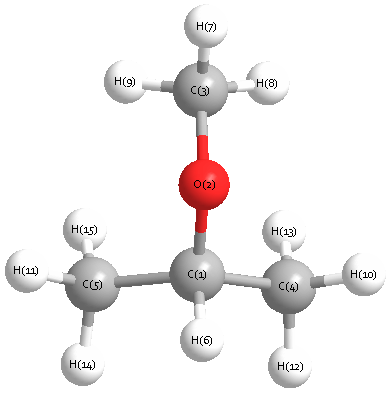
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3126 |
3009 |
49.85 |
|
|
|
| 2 |
A |
3123 |
3006 |
16.08 |
|
|
|
| 3 |
A |
3121 |
3004 |
19.21 |
|
|
|
| 4 |
A |
3115 |
2999 |
92.89 |
|
|
|
| 5 |
A |
3106 |
2990 |
0.02 |
|
|
|
| 6 |
A |
3058 |
2944 |
13.49 |
|
|
|
| 7 |
A |
3050 |
2936 |
92.00 |
|
|
|
| 8 |
A |
3044 |
2930 |
13.85 |
|
|
|
| 9 |
A |
3037 |
2923 |
1.88 |
|
|
|
| 10 |
A |
2996 |
2885 |
53.61 |
|
|
|
| 11 |
A |
1528 |
1471 |
16.65 |
|
|
|
| 12 |
A |
1523 |
1466 |
1.77 |
|
|
|
| 13 |
A |
1514 |
1457 |
2.70 |
|
|
|
| 14 |
A |
1509 |
1453 |
9.97 |
|
|
|
| 15 |
A |
1496 |
1440 |
0.86 |
|
|
|
| 16 |
A |
1493 |
1438 |
2.43 |
|
|
|
| 17 |
A |
1481 |
1426 |
0.53 |
|
|
|
| 18 |
A |
1422 |
1369 |
23.55 |
|
|
|
| 19 |
A |
1406 |
1354 |
0.28 |
|
|
|
| 20 |
A |
1402 |
1350 |
26.75 |
|
|
|
| 21 |
A |
1365 |
1314 |
3.05 |
|
|
|
| 22 |
A |
1237 |
1191 |
21.46 |
|
|
|
| 23 |
A |
1205 |
1160 |
134.67 |
|
|
|
| 24 |
A |
1183 |
1139 |
7.88 |
|
|
|
| 25 |
A |
1182 |
1138 |
1.78 |
|
|
|
| 26 |
A |
1146 |
1103 |
17.68 |
|
|
|
| 27 |
A |
1077 |
1037 |
34.27 |
|
|
|
| 28 |
A |
936 |
901 |
0.10 |
|
|
|
| 29 |
A |
923 |
888 |
16.14 |
|
|
|
| 30 |
A |
919 |
885 |
0.28 |
|
|
|
| 31 |
A |
791 |
762 |
8.98 |
|
|
|
| 32 |
A |
549 |
529 |
2.07 |
|
|
|
| 33 |
A |
417 |
401 |
9.50 |
|
|
|
| 34 |
A |
364 |
351 |
1.02 |
|
|
|
| 35 |
A |
305 |
293 |
0.74 |
|
|
|
| 36 |
A |
254 |
245 |
1.64 |
|
|
|
| 37 |
A |
248 |
238 |
0.03 |
|
|
|
| 38 |
A |
176 |
170 |
1.80 |
|
|
|
| 39 |
A |
44 |
43 |
3.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29934.2 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 28817.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.461 |
|
|
|
| 2 |
H |
0.201 |
|
|
|
| 3 |
H |
0.223 |
|
|
|
| 4 |
H |
0.201 |
|
|
|
| 5 |
O |
-0.337 |
|
|
|
| 6 |
C |
-0.669 |
|
|
|
| 7 |
H |
0.222 |
|
|
|
| 8 |
H |
0.234 |
|
|
|
| 9 |
H |
0.216 |
|
|
|
| 10 |
C |
-0.669 |
|
|
|
| 11 |
H |
0.222 |
|
|
|
| 12 |
H |
0.216 |
|
|
|
| 13 |
H |
0.235 |
|
|
|
| 14 |
C |
-0.044 |
|
|
|
| 15 |
H |
0.209 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.588 |
0.002 |
1.149 |
1.291 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.369 |
0.001 |
2.264 |
| y |
0.001 |
-33.473 |
-0.003 |
| z |
2.264 |
-0.003 |
-34.181 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.458 |
0.001 |
2.264 |
| y |
0.001 |
-0.698 |
-0.003 |
| z |
2.264 |
-0.003 |
-1.759 |
|
| Polar |
|---|
| 3z2-r2 | -3.518 |
|---|
| x2-y2 | 2.104 |
|---|
| xy | 0.001 |
|---|
| xz | 2.264 |
|---|
| yz | -0.003 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
133.931 |
| (<r2>)1/2 |
11.573 |