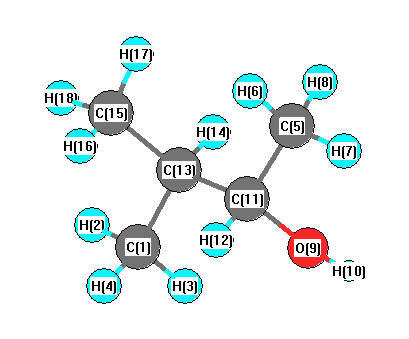
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3818 |
3675 |
8.04 |
|
|
|
| 2 |
A |
3144 |
3026 |
18.42 |
|
|
|
| 3 |
A |
3126 |
3010 |
29.09 |
|
|
|
| 4 |
A |
3115 |
2999 |
41.68 |
|
|
|
| 5 |
A |
3101 |
2986 |
96.26 |
|
|
|
| 6 |
A |
3097 |
2982 |
19.70 |
|
|
|
| 7 |
A |
3093 |
2978 |
8.92 |
|
|
|
| 8 |
A |
3036 |
2923 |
38.52 |
|
|
|
| 9 |
A |
3033 |
2920 |
45.36 |
|
|
|
| 10 |
A |
3030 |
2917 |
21.07 |
|
|
|
| 11 |
A |
3023 |
2910 |
9.22 |
|
|
|
| 12 |
A |
2955 |
2845 |
51.11 |
|
|
|
| 13 |
A |
1535 |
1478 |
7.91 |
|
|
|
| 14 |
A |
1521 |
1464 |
6.04 |
|
|
|
| 15 |
A |
1514 |
1457 |
16.99 |
|
|
|
| 16 |
A |
1507 |
1450 |
6.52 |
|
|
|
| 17 |
A |
1502 |
1446 |
2.75 |
|
|
|
| 18 |
A |
1500 |
1444 |
4.79 |
|
|
|
| 19 |
A |
1437 |
1383 |
8.94 |
|
|
|
| 20 |
A |
1423 |
1370 |
7.85 |
|
|
|
| 21 |
A |
1415 |
1362 |
17.81 |
|
|
|
| 22 |
A |
1408 |
1355 |
6.33 |
|
|
|
| 23 |
A |
1378 |
1326 |
5.76 |
|
|
|
| 24 |
A |
1349 |
1299 |
0.59 |
|
|
|
| 25 |
A |
1344 |
1294 |
20.06 |
|
|
|
| 26 |
A |
1277 |
1229 |
27.46 |
|
|
|
| 27 |
A |
1199 |
1155 |
6.02 |
|
|
|
| 28 |
A |
1184 |
1140 |
14.42 |
|
|
|
| 29 |
A |
1156 |
1113 |
5.09 |
|
|
|
| 30 |
A |
1109 |
1067 |
87.70 |
|
|
|
| 31 |
A |
1086 |
1046 |
7.38 |
|
|
|
| 32 |
A |
994 |
957 |
5.02 |
|
|
|
| 33 |
A |
973 |
937 |
4.23 |
|
|
|
| 34 |
A |
957 |
921 |
12.91 |
|
|
|
| 35 |
A |
935 |
900 |
1.26 |
|
|
|
| 36 |
A |
902 |
869 |
7.91 |
|
|
|
| 37 |
A |
791 |
761 |
2.38 |
|
|
|
| 38 |
A |
527 |
507 |
4.46 |
|
|
|
| 39 |
A |
463 |
446 |
9.44 |
|
|
|
| 40 |
A |
413 |
398 |
1.37 |
|
|
|
| 41 |
A |
370 |
356 |
4.93 |
|
|
|
| 42 |
A |
366 |
353 |
5.17 |
|
|
|
| 43 |
A |
293 |
283 |
56.18 |
|
|
|
| 44 |
A |
284 |
273 |
65.75 |
|
|
|
| 45 |
A |
244 |
234 |
1.33 |
|
|
|
| 46 |
A |
226 |
218 |
0.68 |
|
|
|
| 47 |
A |
211 |
203 |
0.46 |
|
|
|
| 48 |
A |
82 |
79 |
1.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36220.8 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 34869.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.644 |
|
|
|
| 2 |
H |
0.215 |
|
|
|
| 3 |
H |
0.249 |
|
|
|
| 4 |
H |
0.206 |
|
|
|
| 5 |
C |
-0.664 |
|
|
|
| 6 |
H |
0.227 |
|
|
|
| 7 |
H |
0.206 |
|
|
|
| 8 |
H |
0.228 |
|
|
|
| 9 |
O |
-0.575 |
|
|
|
| 10 |
H |
0.387 |
|
|
|
| 11 |
C |
0.018 |
|
|
|
| 12 |
H |
0.186 |
|
|
|
| 13 |
C |
-0.265 |
|
|
|
| 14 |
H |
0.216 |
|
|
|
| 15 |
C |
-0.659 |
|
|
|
| 16 |
H |
0.214 |
|
|
|
| 17 |
H |
0.228 |
|
|
|
| 18 |
H |
0.227 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.912 |
0.902 |
0.895 |
1.564 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.354 |
-1.099 |
2.196 |
| y |
-1.099 |
-41.983 |
-1.369 |
| z |
2.196 |
-1.369 |
-39.899 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.587 |
-1.099 |
2.196 |
| y |
-1.099 |
-3.856 |
-1.369 |
| z |
2.196 |
-1.369 |
-0.731 |
|
| Polar |
|---|
| 3z2-r2 | -1.462 |
|---|
| x2-y2 | 5.629 |
|---|
| xy | -1.099 |
|---|
| xz | 2.196 |
|---|
| yz | -1.369 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
195.079 |
| (<r2>)1/2 |
13.967 |