return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (2-Butanol, 3-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-272.246756
Energy at 298.15K-272.260346
HF Energy-271.236483
Nuclear repulsion energy265.176532
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3838 3635 13.84      
2 A 3199 3030 12.31      
3 A 3186 3017 18.25      
4 A 3172 3005 30.42      
5 A 3157 2990 78.65      
6 A 3155 2988 8.26      
7 A 3151 2984 8.05      
8 A 3078 2915 40.68      
9 A 3074 2911 11.48      
10 A 3071 2908 21.57      
11 A 3070 2908 13.16      
12 A 3010 2851 45.13      
13 A 1561 1479 7.87      
14 A 1546 1464 8.33      
15 A 1541 1459 18.02      
16 A 1531 1450 7.73      
17 A 1529 1448 1.79      
18 A 1524 1443 4.74      
19 A 1462 1384 10.01      
20 A 1450 1373 9.34      
21 A 1442 1366 18.39      
22 A 1435 1359 10.27      
23 A 1401 1327 7.58      
24 A 1373 1301 1.35      
25 A 1360 1288 17.35      
26 A 1296 1228 24.97      
27 A 1221 1157 5.17      
28 A 1210 1146 14.91      
29 A 1180 1117 2.32      
30 A 1134 1074 86.03      
31 A 1117 1058 6.62      
32 A 1008 955 4.54      
33 A 994 941 4.62      
34 A 980 928 10.46      
35 A 949 899 1.03      
36 A 919 870 7.97      
37 A 809 766 2.33      
38 A 538 509 4.73      
39 A 474 449 9.85      
40 A 420 398 1.20      
41 A 379 359 3.42      
42 A 378 358 8.58      
43 A 300 284 25.70      
44 A 293 277 96.14      
45 A 240 227 1.91      
46 A 231 219 2.34      
47 A 211 200 0.55      
48 A 86 82 2.03      

Unscaled Zero Point Vibrational Energy (zpe) 36840.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 34891.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.14669 0.10079 0.06508

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.566 -1.143 0.077
H2 -2.539 -1.087 -0.419
H3 -1.094 -2.088 -0.186
H4 -1.742 -1.136 1.159
C5 1.639 1.055 -0.043
H6 1.307 2.008 0.373
H7 2.639 0.857 0.356
H8 1.721 1.147 -1.129
O9 1.232 -1.326 -0.147
H10 2.103 -1.431 0.245
C11 0.695 -0.086 0.310
H12 0.566 -0.120 1.405
C13 -0.686 0.039 -0.324
H14 -0.531 0.005 -1.411
C15 -1.358 1.361 0.042
H16 -1.407 1.483 1.129
H17 -0.835 2.224 -0.374
H18 -2.383 1.383 -0.338

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 O9 H10 C11 H12 C13 H14 C15 H16 H17 H18
C11.09351.08931.09553.88814.27414.66484.18362.81333.68462.50702.71151.52692.14602.51212.83273.47442.6870
H21.09351.77351.76814.71004.99955.58494.86223.78854.70213.46313.72822.17002.49212.75613.20623.72372.4764
H31.08931.77351.77054.16804.78104.78614.39122.44853.29262.73113.02642.17052.49053.46663.81854.32423.7064
H41.09551.76811.77054.20414.44924.87954.73673.25383.96332.78622.53312.16633.06142.76162.64003.80232.9994
C53.88814.71004.16804.20411.09141.09491.09312.41772.54561.52192.15202.55312.77133.01393.29202.75604.0459
H64.27414.99954.78104.44921.09141.76061.78033.37503.53242.18202.47892.88723.25112.76252.86622.27853.8091
H74.66485.58494.78614.87951.09491.76061.76982.64532.35272.16092.52103.49133.72754.04114.16703.80395.0969
H84.18364.86224.39124.73671.09311.78031.76982.70492.94632.15443.06002.76952.54013.30133.87302.87474.1861
O92.81333.78852.44853.25382.41773.37502.64532.70490.96131.42672.07552.36092.54473.73674.06024.11394.5215
H103.68464.70213.29263.96332.54563.53242.35272.94630.96131.94872.32973.20413.42684.45174.64734.73035.3279
C112.50703.46312.73112.78621.52192.18202.16092.15441.42671.94871.10361.52502.11462.52582.74812.85343.4715
H122.71153.72823.02642.53312.15202.47892.52103.06002.07552.32971.10362.14083.02462.78462.55723.25913.7408
C131.52692.17002.17052.16632.55312.88723.49132.76952.36093.20411.52502.14081.09851.52732.17192.19062.1648
H142.14602.49212.49053.06142.77133.25113.72752.54012.54473.42682.11463.02461.09852.15243.06672.46802.5459
C152.51212.75613.46662.76163.01392.76254.04113.30133.73674.45172.52582.78461.52732.15241.09541.09161.0929
H162.83273.20623.81852.64003.29202.86624.16703.87304.06024.64732.74812.55722.17193.06671.09541.77081.7646
H173.47443.72374.32423.80232.75602.27853.80392.87474.11394.73032.85343.25912.19062.46801.09161.77081.7617
H182.68702.47643.70642.99944.04593.80915.09694.18614.52155.32793.47153.74082.16482.54591.09291.76461.7617

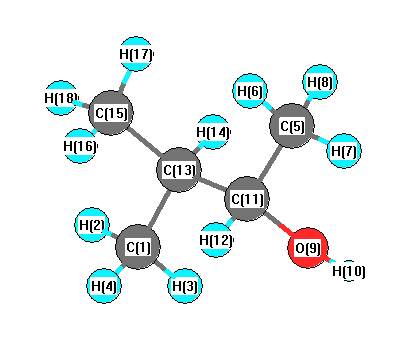
picture of 2-Butanol, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C13 C11 110.462 C1 C13 H14 108.553
C1 C13 C15 110.674 H2 C1 H3 108.672
H2 C1 H4 107.749 H2 C1 C13 110.723
H3 C1 H4 108.264 H3 C1 C13 111.016
H4 C1 C13 110.317 C5 C11 O9 110.112
C5 C11 H12 109.058 C5 C11 C13 113.841
H6 C5 H7 107.276 H6 C5 H8 109.167
H6 C5 C11 112.168 H7 C5 H8 107.978
H7 C5 C11 110.268 H8 C5 C11 109.863
O9 C11 H12 109.557 O9 C11 C13 106.182
H10 O9 C11 107.787 C11 C13 H14 106.284
C11 C13 C15 111.692 H12 C11 C13 107.992
C13 C15 H16 110.740 C13 C15 H17 112.476
C13 C15 H18 110.321 H14 C13 C15 109.021
H16 C15 H17 108.130 H16 C15 H18 107.490
H17 C15 H18 107.495
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability