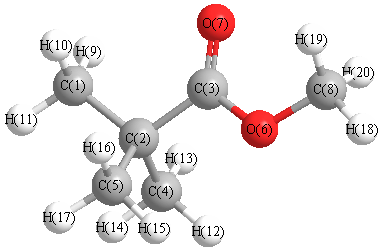
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3174 |
3056 |
19.11 |
|
|
|
| 2 |
A |
3139 |
3022 |
26.30 |
|
|
|
| 3 |
A |
3135 |
3018 |
42.02 |
|
|
|
| 4 |
A |
3134 |
3017 |
26.75 |
|
|
|
| 5 |
A |
3131 |
3014 |
0.09 |
|
|
|
| 6 |
A |
3123 |
3006 |
39.56 |
|
|
|
| 7 |
A |
3108 |
2992 |
29.36 |
|
|
|
| 8 |
A |
3106 |
2990 |
13.57 |
|
|
|
| 9 |
A |
3062 |
2948 |
35.51 |
|
|
|
| 10 |
A |
3050 |
2937 |
27.88 |
|
|
|
| 11 |
A |
3040 |
2927 |
13.90 |
|
|
|
| 12 |
A |
3038 |
2925 |
34.30 |
|
|
|
| 13 |
A |
1817 |
1749 |
225.31 |
|
|
|
| 14 |
A |
1537 |
1479 |
34.07 |
|
|
|
| 15 |
A |
1517 |
1460 |
16.12 |
|
|
|
| 16 |
A |
1512 |
1456 |
9.24 |
|
|
|
| 17 |
A |
1512 |
1456 |
4.62 |
|
|
|
| 18 |
A |
1502 |
1446 |
0.21 |
|
|
|
| 19 |
A |
1497 |
1441 |
9.57 |
|
|
|
| 20 |
A |
1497 |
1441 |
0.14 |
|
|
|
| 21 |
A |
1488 |
1432 |
0.03 |
|
|
|
| 22 |
A |
1480 |
1425 |
9.40 |
|
|
|
| 23 |
A |
1439 |
1385 |
12.72 |
|
|
|
| 24 |
A |
1410 |
1358 |
8.10 |
|
|
|
| 25 |
A |
1403 |
1351 |
9.04 |
|
|
|
| 26 |
A |
1317 |
1267 |
77.94 |
|
|
|
| 27 |
A |
1263 |
1216 |
5.74 |
|
|
|
| 28 |
A |
1239 |
1193 |
2.13 |
|
|
|
| 29 |
A |
1223 |
1177 |
117.33 |
|
|
|
| 30 |
A |
1191 |
1147 |
225.67 |
|
|
|
| 31 |
A |
1184 |
1140 |
0.57 |
|
|
|
| 32 |
A |
1065 |
1025 |
11.29 |
|
|
|
| 33 |
A |
1054 |
1014 |
0.41 |
|
|
|
| 34 |
A |
1022 |
984 |
5.12 |
|
|
|
| 35 |
A |
971 |
934 |
0.04 |
|
|
|
| 36 |
A |
957 |
921 |
2.96 |
|
|
|
| 37 |
A |
951 |
916 |
1.66 |
|
|
|
| 38 |
A |
885 |
852 |
13.80 |
|
|
|
| 39 |
A |
807 |
777 |
1.71 |
|
|
|
| 40 |
A |
786 |
756 |
5.03 |
|
|
|
| 41 |
A |
592 |
570 |
2.08 |
|
|
|
| 42 |
A |
497 |
478 |
2.83 |
|
|
|
| 43 |
A |
379 |
365 |
0.99 |
|
|
|
| 44 |
A |
363 |
349 |
10.79 |
|
|
|
| 45 |
A |
340 |
328 |
0.40 |
|
|
|
| 46 |
A |
321 |
309 |
7.23 |
|
|
|
| 47 |
A |
291 |
280 |
2.26 |
|
|
|
| 48 |
A |
268 |
258 |
0.07 |
|
|
|
| 49 |
A |
259 |
249 |
2.01 |
|
|
|
| 50 |
A |
214 |
206 |
0.25 |
|
|
|
| 51 |
A |
199 |
192 |
1.77 |
|
|
|
| 52 |
A |
134 |
129 |
1.30 |
|
|
|
| 53 |
A |
119 |
114 |
1.10 |
|
|
|
| 54 |
A |
35 |
34 |
0.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38385.9 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 36954.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.610 |
|
|
|
| 2 |
C |
-0.261 |
|
|
|
| 3 |
C |
0.510 |
|
|
|
| 4 |
C |
-0.627 |
|
|
|
| 5 |
C |
-0.627 |
|
|
|
| 6 |
O |
-0.312 |
|
|
|
| 7 |
O |
-0.351 |
|
|
|
| 8 |
C |
-0.481 |
|
|
|
| 9 |
H |
0.232 |
|
|
|
| 10 |
H |
0.232 |
|
|
|
| 11 |
H |
0.217 |
|
|
|
| 12 |
H |
0.240 |
|
|
|
| 13 |
H |
0.228 |
|
|
|
| 14 |
H |
0.225 |
|
|
|
| 15 |
H |
0.240 |
|
|
|
| 16 |
H |
0.228 |
|
|
|
| 17 |
H |
0.225 |
|
|
|
| 18 |
H |
0.227 |
|
|
|
| 19 |
H |
0.233 |
|
|
|
| 20 |
H |
0.233 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.212 |
-1.744 |
0.000 |
1.757 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.587 |
1.756 |
0.001 |
| y |
1.756 |
-55.057 |
0.000 |
| z |
0.001 |
0.000 |
-48.972 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.428 |
1.756 |
0.001 |
| y |
1.756 |
-8.777 |
0.000 |
| z |
0.001 |
0.000 |
0.350 |
|
| Polar |
|---|
| 3z2-r2 | 0.700 |
|---|
| x2-y2 | 11.470 |
|---|
| xy | 1.756 |
|---|
| xz | 0.001 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.017 |
0.158 |
-0.000 |
| y |
0.158 |
10.638 |
0.000 |
| z |
-0.000 |
0.000 |
9.592 |
<r2> (average value of r
2) Å
2
| <r2> |
296.291 |
| (<r2>)1/2 |
17.213 |