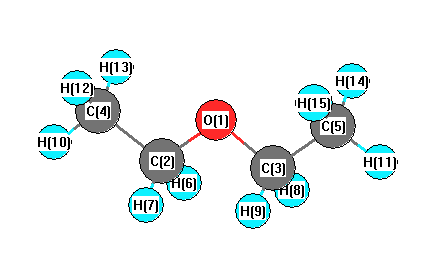
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3124 |
3007 |
44.52 |
|
|
|
| 2 |
A1 |
3049 |
2935 |
9.27 |
|
|
|
| 3 |
A1 |
2972 |
2861 |
125.08 |
|
|
|
| 4 |
A1 |
1551 |
1493 |
2.96 |
|
|
|
| 5 |
A1 |
1515 |
1459 |
4.31 |
|
|
|
| 6 |
A1 |
1463 |
1408 |
5.28 |
|
|
|
| 7 |
A1 |
1408 |
1355 |
1.97 |
|
|
|
| 8 |
A1 |
1191 |
1147 |
22.49 |
|
|
|
| 9 |
A1 |
1070 |
1030 |
13.64 |
|
|
|
| 10 |
A1 |
868 |
836 |
7.05 |
|
|
|
| 11 |
A1 |
442 |
426 |
0.03 |
|
|
|
| 12 |
A1 |
194 |
186 |
0.52 |
|
|
|
| 13 |
A2 |
3127 |
3010 |
0.00 |
|
|
|
| 14 |
A2 |
2991 |
2879 |
0.00 |
|
|
|
| 15 |
A2 |
1495 |
1440 |
0.00 |
|
|
|
| 16 |
A2 |
1306 |
1257 |
0.00 |
|
|
|
| 17 |
A2 |
1177 |
1133 |
0.00 |
|
|
|
| 18 |
A2 |
820 |
790 |
0.00 |
|
|
|
| 19 |
A2 |
254 |
244 |
0.00 |
|
|
|
| 20 |
A2 |
103 |
100 |
0.00 |
|
|
|
| 21 |
B1 |
3127 |
3010 |
70.95 |
|
|
|
| 22 |
B1 |
2989 |
2877 |
143.10 |
|
|
|
| 23 |
B1 |
1496 |
1440 |
16.18 |
|
|
|
| 24 |
B1 |
1314 |
1265 |
4.48 |
|
|
|
| 25 |
B1 |
1207 |
1162 |
13.14 |
|
|
|
| 26 |
B1 |
834 |
803 |
0.90 |
|
|
|
| 27 |
B1 |
260 |
251 |
1.61 |
|
|
|
| 28 |
B1 |
108 |
104 |
4.20 |
|
|
|
| 29 |
B2 |
3124 |
3007 |
22.37 |
|
|
|
| 30 |
B2 |
3048 |
2934 |
34.62 |
|
|
|
| 31 |
B2 |
2959 |
2849 |
9.79 |
|
|
|
| 32 |
B2 |
1531 |
1474 |
2.66 |
|
|
|
| 33 |
B2 |
1512 |
1455 |
3.61 |
|
|
|
| 34 |
B2 |
1422 |
1369 |
51.57 |
|
|
|
| 35 |
B2 |
1389 |
1337 |
26.76 |
|
|
|
| 36 |
B2 |
1177 |
1133 |
219.21 |
|
|
|
| 37 |
B2 |
1107 |
1066 |
4.40 |
|
|
|
| 38 |
B2 |
955 |
920 |
4.87 |
|
|
|
| 39 |
B2 |
434 |
418 |
3.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30055.0 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 28934.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.334 |
|
|
|
| 2 |
C |
-0.211 |
|
|
|
| 3 |
C |
-0.211 |
|
|
|
| 4 |
C |
-0.667 |
|
|
|
| 5 |
C |
-0.667 |
|
|
|
| 6 |
H |
0.188 |
|
|
|
| 7 |
H |
0.188 |
|
|
|
| 8 |
H |
0.188 |
|
|
|
| 9 |
H |
0.188 |
|
|
|
| 10 |
H |
0.215 |
|
|
|
| 11 |
H |
0.215 |
|
|
|
| 12 |
H |
0.227 |
|
|
|
| 13 |
H |
0.227 |
|
|
|
| 14 |
H |
0.227 |
|
|
|
| 15 |
H |
0.227 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.101 |
1.101 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
176.065 |
| (<r2>)1/2 |
13.269 |