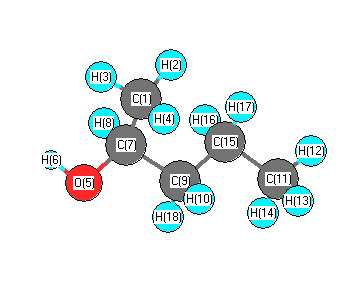
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3812 |
3669 |
5.75 |
|
|
|
| 2 |
A |
3119 |
3003 |
34.88 |
|
|
|
| 3 |
A |
3107 |
2991 |
48.14 |
|
|
|
| 4 |
A |
3104 |
2988 |
78.82 |
|
|
|
| 5 |
A |
3096 |
2981 |
34.82 |
|
|
|
| 6 |
A |
3080 |
2965 |
27.21 |
|
|
|
| 7 |
A |
3051 |
2937 |
3.99 |
|
|
|
| 8 |
A |
3036 |
2923 |
58.40 |
|
|
|
| 9 |
A |
3032 |
2919 |
22.04 |
|
|
|
| 10 |
A |
3030 |
2917 |
32.64 |
|
|
|
| 11 |
A |
3021 |
2908 |
12.13 |
|
|
|
| 12 |
A |
2962 |
2852 |
56.37 |
|
|
|
| 13 |
A |
1522 |
1465 |
10.48 |
|
|
|
| 14 |
A |
1514 |
1458 |
6.93 |
|
|
|
| 15 |
A |
1512 |
1456 |
9.16 |
|
|
|
| 16 |
A |
1511 |
1454 |
3.69 |
|
|
|
| 17 |
A |
1505 |
1449 |
1.70 |
|
|
|
| 18 |
A |
1492 |
1437 |
0.85 |
|
|
|
| 19 |
A |
1438 |
1385 |
17.03 |
|
|
|
| 20 |
A |
1423 |
1370 |
8.07 |
|
|
|
| 21 |
A |
1418 |
1365 |
23.61 |
|
|
|
| 22 |
A |
1398 |
1346 |
7.58 |
|
|
|
| 23 |
A |
1381 |
1329 |
2.62 |
|
|
|
| 24 |
A |
1339 |
1289 |
1.57 |
|
|
|
| 25 |
A |
1320 |
1270 |
25.05 |
|
|
|
| 26 |
A |
1294 |
1246 |
5.02 |
|
|
|
| 27 |
A |
1251 |
1204 |
17.36 |
|
|
|
| 28 |
A |
1189 |
1145 |
14.53 |
|
|
|
| 29 |
A |
1153 |
1110 |
31.79 |
|
|
|
| 30 |
A |
1111 |
1069 |
46.90 |
|
|
|
| 31 |
A |
1098 |
1057 |
5.81 |
|
|
|
| 32 |
A |
1052 |
1012 |
4.14 |
|
|
|
| 33 |
A |
1012 |
974 |
4.26 |
|
|
|
| 34 |
A |
970 |
934 |
10.67 |
|
|
|
| 35 |
A |
921 |
886 |
22.75 |
|
|
|
| 36 |
A |
880 |
847 |
0.66 |
|
|
|
| 37 |
A |
843 |
811 |
0.14 |
|
|
|
| 38 |
A |
747 |
719 |
3.67 |
|
|
|
| 39 |
A |
491 |
472 |
9.69 |
|
|
|
| 40 |
A |
450 |
433 |
1.80 |
|
|
|
| 41 |
A |
413 |
398 |
10.63 |
|
|
|
| 42 |
A |
321 |
309 |
12.87 |
|
|
|
| 43 |
A |
313 |
302 |
116.17 |
|
|
|
| 44 |
A |
265 |
255 |
1.70 |
|
|
|
| 45 |
A |
247 |
238 |
0.80 |
|
|
|
| 46 |
A |
182 |
175 |
1.79 |
|
|
|
| 47 |
A |
114 |
110 |
0.01 |
|
|
|
| 48 |
A |
82 |
79 |
0.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36309.3 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 34955.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.666 |
|
|
|
| 2 |
H |
0.224 |
|
|
|
| 3 |
H |
0.207 |
|
|
|
| 4 |
H |
0.227 |
|
|
|
| 5 |
O |
-0.573 |
|
|
|
| 6 |
H |
0.385 |
|
|
|
| 7 |
C |
0.007 |
|
|
|
| 8 |
H |
0.188 |
|
|
|
| 9 |
C |
-0.441 |
|
|
|
| 10 |
H |
0.218 |
|
|
|
| 11 |
C |
-0.647 |
|
|
|
| 12 |
H |
0.220 |
|
|
|
| 13 |
H |
0.219 |
|
|
|
| 14 |
H |
0.220 |
|
|
|
| 15 |
C |
-0.447 |
|
|
|
| 16 |
H |
0.212 |
|
|
|
| 17 |
H |
0.219 |
|
|
|
| 18 |
H |
0.230 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.462 |
1.330 |
-0.817 |
1.628 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.795 |
-1.614 |
1.946 |
| y |
-1.614 |
-42.630 |
0.861 |
| z |
1.946 |
0.861 |
-39.848 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.444 |
-1.614 |
1.946 |
| y |
-1.614 |
-4.809 |
0.861 |
| z |
1.946 |
0.861 |
-0.635 |
|
| Polar |
|---|
| 3z2-r2 | -1.270 |
|---|
| x2-y2 | 6.835 |
|---|
| xy | -1.614 |
|---|
| xz | 1.946 |
|---|
| yz | 0.861 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
239.188 |
| (<r2>)1/2 |
15.466 |