return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CONH2 (Acetamide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-208.733602
Energy at 298.15K-208.739625
HF Energy-208.023637
Nuclear repulsion energy121.576619
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3779 3579 29.84      
2 A 3643 3450 32.55      
3 A 3220 3049 5.22      
4 A 3177 3008 10.66      
5 A 3092 2929 5.76      
6 A 1823 1726 282.21      
7 A 1693 1603 90.95      
8 A 1526 1445 11.66      
9 A 1515 1435 12.29      
10 A 1441 1365 100.15      
11 A 1373 1300 91.93      
12 A 1166 1105 0.42      
13 A 1077 1020 4.26      
14 A 1001 948 8.87      
15 A 870 824 2.64      
16 A 659 624 29.80      
17 A 565 535 22.68      
18 A 517 490 27.72      
19 A 429 406 22.65      
20 A 390 369 230.83      
21 A 71 67 0.89      

Unscaled Zero Point Vibrational Energy (zpe) 16512.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15638.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.35835 0.31101 0.17215

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.368 -0.298 -0.001
C2 0.079 0.153 -0.007
N3 0.994 -0.871 -0.054
O4 0.412 1.324 0.009
H5 -1.993 0.534 -0.317
H6 -1.533 -1.154 -0.658
H7 -1.659 -0.584 1.014
H8 1.950 -0.615 0.129
H9 0.718 -1.806 0.186

Atom - Atom Distances (Å)
  C1 C2 N3 O4 H5 H6 H7 H8 H9
C11.51552.43132.40851.08791.09151.09343.33622.5805
C21.51551.37411.21802.12892.17452.14592.02772.0697
N32.43131.37412.27193.31142.61402.87451.00661.0046
O42.40851.21802.27192.55193.21992.98982.47853.1504
H51.08792.12893.31142.55191.78221.77014.13143.6162
H61.09152.17452.61403.21991.78221.77133.61192.4909
H71.09342.14592.87452.98981.77011.77133.71652.7980
H83.33622.02771.00662.47854.13143.61193.71651.7153
H92.58052.06971.00463.15043.61622.49092.79801.7153

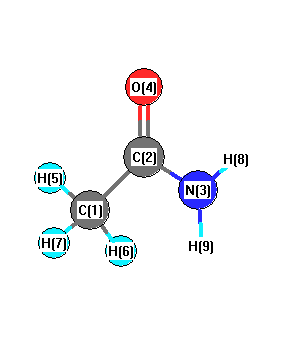
picture of Acetamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 N3 114.488 C1 C2 O4 123.183
C2 C1 H5 108.607 C2 C1 H6 112.020
C2 C1 H7 109.619 C2 N3 H8 115.947
C2 N3 H9 120.142 N3 C2 O4 122.321
H5 C1 H6 109.720 H5 C1 H7 108.483
H6 C1 H7 108.329 H8 N3 H9 117.052
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability