return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (2,4-Hexadiene, (Z,Z)-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-233.954899
Energy at 298.15K-233.964120
HF Energy-233.032631
Nuclear repulsion energy213.064434
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3217 3047 57.75      
2 A' 3210 3040 0.00      
3 A' 3196 3027 0.00      
4 A' 3189 3021 21.65      
5 A' 3176 3008 9.81      
6 A' 3171 3003 0.00      
7 A' 3075 2913 47.76      
8 A' 3075 2912 0.03      
9 A' 1728 1637 0.00      
10 A' 1670 1581 8.96      
11 A' 1532 1451 0.00      
12 A' 1528 1447 25.97      
13 A' 1490 1411 0.00      
14 A' 1457 1380 15.14      
15 A' 1445 1369 0.00      
16 A' 1391 1318 2.82      
17 A' 1324 1254 0.00      
18 A' 1289 1221 0.71      
19 A' 1192 1129 0.00      
20 A' 1110 1052 0.39      
21 A' 1027 973 0.00      
22 A' 948 898 26.11      
23 A' 937 888 0.00      
24 A' 641 607 0.00      
25 A' 523 495 14.62      
26 A' 278 264 0.00      
27 A' 192 182 0.19      
28 A" 3140 2974 1.29      
29 A" 3140 2974 33.51      
30 A" 1521 1441 0.10      
31 A" 1521 1441 18.35      
32 A" 1071 1015 0.00      
33 A" 1070 1013 1.80      
34 A" 959 908 0.25      
35 A" 927 878 0.00      
36 A" 778 737 0.00      
37 A" 683 647 80.35      
38 A" 452 428 0.00      
39 A" 297 282 0.11      
40 A" 86 81 0.00      
41 A" 79 75 0.71      
42 A" 43 41 0.18      

Unscaled Zero Point Vibrational Energy (zpe) 31389.9 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 29729.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.38699 0.05215 0.04675

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.924 -2.599 0.000
C2 1.093 -1.522 0.000
H3 -0.966 -1.228 0.000
C4 0.000 -0.728 0.000
H5 0.966 1.228 0.000
C6 0.000 0.728 0.000
H7 -0.924 2.599 0.000
C8 -1.093 1.522 0.000
H9 -2.656 0.016 0.000
H10 -3.048 1.497 0.878
H11 -3.048 1.497 -0.878
C12 -2.531 1.098 0.000
H13 2.656 -0.016 0.000
H14 3.048 -1.497 0.878
H15 3.048 -1.497 -0.878
C16 2.531 -1.098 0.000

Atom - Atom Distances (Å)
  H1 C2 H3 C4 H5 C6 H7 C8 H9 H10 H11 C12 H13 H14 H15 C16
H11.08962.33442.08613.82653.45265.51584.58784.43265.77225.77225.05983.10942.54942.54942.1985
C21.08962.07951.35092.75282.50174.58783.74784.05185.19935.19934.47192.17042.14372.14371.4991
H32.33442.07951.08733.12392.18123.82652.75282.09823.53973.53972.80353.81854.11764.11763.4989
C42.08611.35091.08732.18121.45633.45262.50172.75793.87463.87463.12112.74933.26393.26392.5577
H53.82652.75283.12392.18121.08732.33442.07953.81854.11764.11763.49892.09823.53973.53972.8035
C63.45262.50172.18121.45631.08732.08611.35092.74933.26393.26392.55772.75793.87463.87463.1211
H75.51584.58783.82653.45262.33442.08611.08963.10942.54942.54942.19854.43265.77225.77225.0598
C84.58783.74782.75282.50172.07951.35091.08962.17042.14372.14371.49914.05175.19935.19934.4719
H94.43264.05182.09822.75793.81852.74933.10942.17041.76571.76571.08945.31125.96595.96595.3047
H105.77225.19933.53973.87464.11763.26392.54942.14371.76571.75701.09455.96596.79147.01506.2153
H115.77225.19933.53973.87464.11763.26392.54942.14371.76571.75701.09455.96597.01506.79146.2153
C125.05984.47192.80353.12113.49892.55772.19851.49911.08941.09451.09455.30476.21536.21535.5177
H133.10942.17043.81852.74932.09822.75794.43264.05175.31125.96595.96595.30471.76571.76571.0894
H142.54942.14374.11763.26393.53973.87465.77225.19935.96596.79147.01506.21531.76571.75701.0945
H152.54942.14374.11763.26393.53973.87465.77225.19935.96597.01506.79146.21531.76571.75701.0945
C162.19851.49913.49892.55772.80353.12115.05984.47195.30476.21536.21535.51771.08941.09451.0945

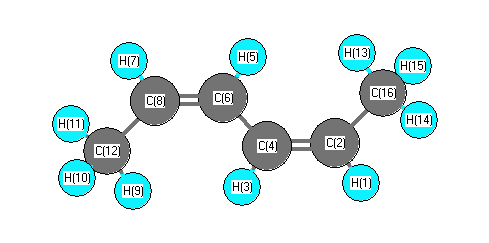
picture of 2,4-Hexadiene, (Z,Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 117.076 H1 C2 C16 115.352
C2 C4 H3 116.646 C2 C4 C6 126.006
C2 C16 H13 113.000 C2 C16 H14 110.520
C2 C16 H15 110.520 H3 C4 C6 117.348
C4 C2 C16 127.572 C4 C6 H5 117.348
C4 C6 C8 126.006 H5 C6 C8 116.646
C6 C8 H7 117.076 C6 C8 C12 127.572
H7 C8 C12 115.352 C8 C12 H9 113.000
C8 C12 H10 110.519 C8 C12 H11 110.519
H9 C12 H10 107.897 H9 C12 H11 107.897
H10 C12 H11 106.763 H13 C16 H14 107.897
H13 C16 H15 107.897 H14 C16 H15 106.762
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability