Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -190.236972 |
| Energy at 298.15K | |
| HF Energy | -189.578880 |
| Nuclear repulsion energy | 88.046569 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3121 |
2956 |
29.49 |
195.15 |
0.13 |
0.24 |
| 2 |
A1 |
2334 |
2210 |
714.23 |
35.20 |
0.35 |
0.52 |
| 3 |
A1 |
1781 |
1687 |
12.34 |
11.33 |
0.40 |
0.57 |
| 4 |
A1 |
1507 |
1427 |
2.55 |
18.23 |
0.52 |
0.69 |
| 5 |
A1 |
913 |
865 |
2.32 |
36.98 |
0.26 |
0.42 |
| 6 |
B1 |
1007 |
954 |
27.30 |
0.13 |
0.75 |
0.86 |
| 7 |
B1 |
650 |
616 |
21.27 |
0.65 |
0.75 |
0.86 |
| 8 |
B1 |
218 |
207 |
1.03 |
2.62 |
0.75 |
0.86 |
| 9 |
B2 |
3202 |
3032 |
10.70 |
123.17 |
0.75 |
0.86 |
| 10 |
B2 |
1083 |
1026 |
1.13 |
0.18 |
0.75 |
0.86 |
| 11 |
B2 |
475 |
450 |
12.42 |
1.46 |
0.75 |
0.86 |
| 12 |
B2 |
168i |
159i |
21.35 |
0.25 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8061.3 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 7634.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.882 |
| C2 |
0.000 |
0.000 |
-0.558 |
| C3 |
0.000 |
0.000 |
0.723 |
| O4 |
0.000 |
0.000 |
1.903 |
| H5 |
0.000 |
0.924 |
-2.461 |
| H6 |
0.000 |
-0.924 |
-2.461 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3242 | 2.6053 | 3.7855 | 1.0904 | 1.0904 |
C2 | 1.3242 | | 1.2810 | 2.4613 | 2.1153 | 2.1153 | C3 | 2.6053 | 1.2810 | | 1.1802 | 3.3152 | 3.3152 | O4 | 3.7855 | 2.4613 | 1.1802 | | 4.4607 | 4.4607 | H5 | 1.0904 | 2.1153 | 3.3152 | 4.4607 | | 1.8487 | H6 | 1.0904 | 2.1153 | 3.3152 | 4.4607 | 1.8487 | |
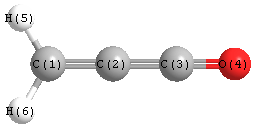 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
122.035 |
| C2 |
C1 |
H6 |
122.035 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
115.930 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -190.239649 |
| Energy at 298.15K | |
| HF Energy | -189.577127 |
| Nuclear repulsion energy | 88.392603 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3227 |
3056 |
8.94 |
104.25 |
0.67 |
0.80 |
| 2 |
A' |
3134 |
2968 |
26.91 |
137.28 |
0.14 |
0.25 |
| 3 |
A' |
2219 |
2102 |
684.51 |
48.20 |
0.41 |
0.59 |
| 4 |
A' |
1724 |
1632 |
6.50 |
3.39 |
0.60 |
0.75 |
| 5 |
A' |
1513 |
1433 |
1.30 |
16.19 |
0.56 |
0.71 |
| 6 |
A' |
1102 |
1043 |
21.61 |
6.25 |
0.07 |
0.14 |
| 7 |
A' |
947 |
897 |
2.00 |
33.84 |
0.18 |
0.31 |
| 8 |
A' |
530 |
502 |
10.81 |
5.08 |
0.59 |
0.74 |
| 9 |
A' |
195 |
185 |
20.82 |
5.42 |
0.73 |
0.84 |
| 10 |
A" |
1012 |
958 |
30.71 |
0.23 |
0.75 |
0.86 |
| 11 |
A" |
722 |
684 |
9.19 |
1.28 |
0.75 |
0.86 |
| 12 |
A" |
288 |
272 |
3.43 |
4.45 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8305.4 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 7866.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.124 |
-1.798 |
0.000 |
| C2 |
-0.408 |
-0.570 |
0.000 |
| C3 |
0.000 |
0.682 |
0.000 |
| O4 |
0.128 |
1.847 |
0.000 |
| H5 |
1.198 |
-1.986 |
0.000 |
| H6 |
-0.513 |
-2.680 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3386 | 2.4832 | 3.6450 | 1.0904 | 1.0878 |
C2 | 1.3386 | | 1.3165 | 2.4752 | 2.1411 | 2.1127 | C3 | 2.4832 | 1.3165 | | 1.1719 | 2.9243 | 3.4007 | O4 | 3.6450 | 2.4752 | 1.1719 | | 3.9792 | 4.5718 | H5 | 1.0904 | 2.1411 | 2.9243 | 3.9792 | | 1.8466 | H6 | 1.0878 | 2.1127 | 3.4007 | 4.5718 | 1.8466 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
138.536 |
|
C2 |
C1 |
H5 |
123.317 |
| C2 |
C1 |
H6 |
120.741 |
|
C2 |
C3 |
O4 |
168.197 |
| H5 |
C1 |
H6 |
115.942 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
