Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -190.545439 |
| Energy at 298.15K | |
| HF Energy | -190.325951 |
| Nuclear repulsion energy | 88.332159 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3098 |
3098 |
36.91 |
206.10 |
0.13 |
0.24 |
| 2 |
A1 |
2318 |
2318 |
837.24 |
31.37 |
0.37 |
0.54 |
| 3 |
A1 |
1783 |
1783 |
22.39 |
2.94 |
0.49 |
0.65 |
| 4 |
A1 |
1504 |
1504 |
4.07 |
16.90 |
0.53 |
0.70 |
| 5 |
A1 |
921 |
921 |
5.77 |
41.81 |
0.27 |
0.42 |
| 6 |
B1 |
1014 |
1014 |
24.51 |
0.38 |
0.75 |
0.86 |
| 7 |
B1 |
652 |
652 |
23.67 |
1.09 |
0.75 |
0.86 |
| 8 |
B1 |
219 |
219 |
0.77 |
3.59 |
0.75 |
0.86 |
| 9 |
B2 |
3169 |
3169 |
15.00 |
137.66 |
0.75 |
0.86 |
| 10 |
B2 |
1083 |
1083 |
1.38 |
0.21 |
0.75 |
0.86 |
| 11 |
B2 |
472 |
472 |
13.52 |
2.01 |
0.75 |
0.86 |
| 12 |
B2 |
139i |
139i |
19.10 |
0.09 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8046.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 8046.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.875 |
| C2 |
0.000 |
0.000 |
-0.556 |
| C3 |
0.000 |
0.000 |
0.720 |
| O4 |
0.000 |
0.000 |
1.897 |
| H5 |
0.000 |
0.922 |
-2.455 |
| H6 |
0.000 |
-0.922 |
-2.455 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3190 | 2.5946 | 3.7719 | 1.0894 | 1.0894 |
C2 | 1.3190 | | 1.2756 | 2.4529 | 2.1113 | 2.1113 | C3 | 2.5946 | 1.2756 | | 1.1773 | 3.3061 | 3.3061 | O4 | 3.7719 | 2.4529 | 1.1773 | | 4.4488 | 4.4488 | H5 | 1.0894 | 2.1113 | 3.3061 | 4.4488 | | 1.8438 | H6 | 1.0894 | 2.1113 | 3.3061 | 4.4488 | 1.8438 | |
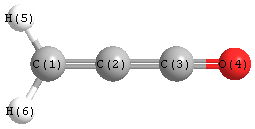 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
122.190 |
| C2 |
C1 |
H6 |
122.190 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
115.619 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -190.546620 |
| Energy at 298.15K | |
| HF Energy | -190.325632 |
| Nuclear repulsion energy | 88.566537 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3189 |
3189 |
13.57 |
121.35 |
0.68 |
0.81 |
| 2 |
A' |
3108 |
3108 |
34.86 |
156.84 |
0.15 |
0.26 |
| 3 |
A' |
2218 |
2218 |
833.03 |
39.00 |
0.40 |
0.57 |
| 4 |
A' |
1749 |
1749 |
7.67 |
1.46 |
0.72 |
0.84 |
| 5 |
A' |
1510 |
1510 |
1.25 |
17.15 |
0.57 |
0.72 |
| 6 |
A' |
1091 |
1091 |
16.94 |
4.61 |
0.12 |
0.22 |
| 7 |
A' |
943 |
943 |
2.81 |
35.52 |
0.19 |
0.32 |
| 8 |
A' |
511 |
511 |
13.30 |
5.17 |
0.57 |
0.73 |
| 9 |
A' |
161 |
161 |
22.28 |
6.46 |
0.74 |
0.85 |
| 10 |
A" |
1020 |
1020 |
26.48 |
0.57 |
0.75 |
0.86 |
| 11 |
A" |
709 |
709 |
14.81 |
1.83 |
0.75 |
0.86 |
| 12 |
A" |
276 |
276 |
1.96 |
5.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8241.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 8241.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.023 |
-1.505 |
0.000 |
| C2 |
0.000 |
-0.657 |
0.000 |
| C3 |
-0.353 |
0.595 |
0.000 |
| O4 |
-0.869 |
1.646 |
0.000 |
| H5 |
2.064 |
-1.182 |
0.000 |
| H6 |
0.865 |
-2.581 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3288 | 2.5109 | 3.6755 | 1.0895 | 1.0871 |
C2 | 1.3288 | | 1.3011 | 2.4617 | 2.1296 | 2.1090 | C3 | 2.5109 | 1.3011 | | 1.1707 | 3.0001 | 3.4013 | O4 | 3.6755 | 2.4617 | 1.1707 | | 4.0742 | 4.5685 | H5 | 1.0895 | 2.1296 | 3.0001 | 4.0742 | | 1.8417 | H6 | 1.0871 | 2.1090 | 3.4013 | 4.5685 | 1.8417 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
145.390 |
|
C2 |
C1 |
H5 |
123.128 |
| C2 |
C1 |
H6 |
121.281 |
|
C2 |
C3 |
O4 |
169.630 |
| H5 |
C1 |
H6 |
115.590 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
