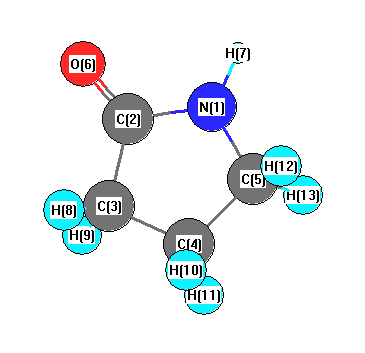
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3895 |
3522 |
34.97 |
|
|
|
| 2 |
A |
3278 |
2965 |
34.78 |
|
|
|
| 3 |
A |
3261 |
2949 |
47.38 |
|
|
|
| 4 |
A |
3243 |
2933 |
60.22 |
|
|
|
| 5 |
A |
3211 |
2904 |
26.62 |
|
|
|
| 6 |
A |
3205 |
2899 |
21.47 |
|
|
|
| 7 |
A |
3169 |
2866 |
80.65 |
|
|
|
| 8 |
A |
1981 |
1792 |
634.55 |
|
|
|
| 9 |
A |
1678 |
1518 |
3.28 |
|
|
|
| 10 |
A |
1638 |
1481 |
11.30 |
|
|
|
| 11 |
A |
1605 |
1452 |
2.64 |
|
|
|
| 12 |
A |
1601 |
1448 |
51.81 |
|
|
|
| 13 |
A |
1510 |
1366 |
13.12 |
|
|
|
| 14 |
A |
1475 |
1334 |
25.16 |
|
|
|
| 15 |
A |
1446 |
1308 |
34.64 |
|
|
|
| 16 |
A |
1386 |
1254 |
95.37 |
|
|
|
| 17 |
A |
1360 |
1230 |
27.18 |
|
|
|
| 18 |
A |
1333 |
1205 |
0.41 |
|
|
|
| 19 |
A |
1298 |
1174 |
7.09 |
|
|
|
| 20 |
A |
1201 |
1086 |
0.33 |
|
|
|
| 21 |
A |
1164 |
1052 |
22.01 |
|
|
|
| 22 |
A |
1072 |
970 |
12.23 |
|
|
|
| 23 |
A |
988 |
893 |
2.23 |
|
|
|
| 24 |
A |
983 |
889 |
4.69 |
|
|
|
| 25 |
A |
955 |
864 |
4.90 |
|
|
|
| 26 |
A |
886 |
801 |
6.08 |
|
|
|
| 27 |
A |
748 |
677 |
14.46 |
|
|
|
| 28 |
A |
697 |
631 |
29.94 |
|
|
|
| 29 |
A |
614 |
556 |
52.93 |
|
|
|
| 30 |
A |
530 |
480 |
78.92 |
|
|
|
| 31 |
A |
508 |
460 |
17.45 |
|
|
|
| 32 |
A |
208 |
189 |
6.03 |
|
|
|
| 33 |
A |
147 |
133 |
1.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26137.2 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 23638.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.701 |
-0.837 |
|
-0.816 |
| 2 |
C |
0.621 |
0.482 |
|
0.482 |
| 3 |
C |
-0.551 |
-0.722 |
|
-0.704 |
| 4 |
C |
-0.451 |
0.354 |
|
0.350 |
| 5 |
C |
-0.191 |
0.028 |
|
0.037 |
| 6 |
O |
-0.480 |
0.815 |
|
0.746 |
| 7 |
H |
0.373 |
-0.095 |
|
-0.066 |
| 8 |
H |
0.241 |
-0.069 |
|
-0.127 |
| 9 |
H |
0.251 |
0.016 |
|
0.041 |
| 10 |
H |
0.227 |
0.016 |
|
0.038 |
| 11 |
H |
0.228 |
-0.013 |
|
-0.041 |
| 12 |
H |
0.210 |
-0.027 |
|
-0.010 |
| 13 |
H |
0.222 |
0.050 |
|
0.071 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.222 |
-0.531 |
0.432 |
4.277 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-1.802 |
0.439 |
0.811 |
2.024 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.617 |
-0.233 |
0.178 |
| y |
-0.233 |
-31.642 |
-0.283 |
| z |
0.178 |
-0.283 |
-35.634 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.979 |
-0.233 |
0.178 |
| y |
-0.233 |
8.483 |
-0.283 |
| z |
0.178 |
-0.283 |
2.496 |
|
| Polar |
|---|
| 3z2-r2 | 4.991 |
|---|
| x2-y2 | -12.975 |
|---|
| xy | -0.233 |
|---|
| xz | 0.178 |
|---|
| yz | -0.283 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.618 |
-0.147 |
0.380 |
| y |
-0.147 |
6.782 |
0.054 |
| z |
0.380 |
0.054 |
5.636 |
<r2> (average value of r
2) Å
2
| <r2> |
142.206 |
| (<r2>)1/2 |
11.925 |