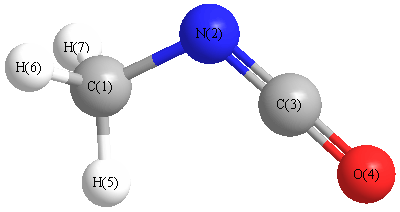
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3151 |
3024 |
19.74 |
85.20 |
0.66 |
0.79 |
| 2 |
A' |
3053 |
2929 |
51.07 |
167.24 |
0.02 |
0.03 |
| 3 |
A' |
2437 |
2339 |
1053.90 |
2.31 |
0.03 |
0.06 |
| 4 |
A' |
1546 |
1483 |
2.59 |
11.83 |
0.14 |
0.25 |
| 5 |
A' |
1505 |
1444 |
12.52 |
20.73 |
0.62 |
0.76 |
| 6 |
A' |
1475 |
1415 |
30.00 |
21.86 |
0.45 |
0.62 |
| 7 |
A' |
1166 |
1119 |
14.88 |
2.42 |
0.61 |
0.76 |
| 8 |
A' |
904 |
867 |
31.70 |
7.13 |
0.21 |
0.35 |
| 9 |
A' |
648 |
621 |
30.63 |
0.41 |
0.66 |
0.80 |
| 10 |
A' |
166 |
160 |
17.60 |
1.81 |
0.68 |
0.81 |
| 11 |
A" |
3120 |
2994 |
26.61 |
79.46 |
0.75 |
0.86 |
| 12 |
A" |
1524 |
1462 |
7.82 |
16.08 |
0.75 |
0.86 |
| 13 |
A" |
1141 |
1095 |
0.01 |
2.02 |
0.75 |
0.86 |
| 14 |
A" |
606 |
582 |
27.69 |
0.68 |
0.75 |
0.86 |
| 15 |
A" |
52 |
50 |
2.66 |
1.07 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11245.8 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 10791.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.551 |
|
|
|
| 2 |
N |
-0.390 |
|
|
|
| 3 |
C |
0.503 |
|
|
|
| 4 |
O |
-0.294 |
|
|
|
| 5 |
H |
0.249 |
|
|
|
| 6 |
H |
0.242 |
|
|
|
| 7 |
H |
0.242 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.500 |
1.567 |
0.000 |
2.950 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.267 |
-0.497 |
0.000 |
| y |
-0.497 |
-24.641 |
0.000 |
| z |
0.000 |
0.000 |
-22.405 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.256 |
-0.497 |
0.000 |
| y |
-0.497 |
-2.305 |
0.000 |
| z |
0.000 |
0.000 |
1.048 |
|
| Polar |
|---|
| 3z2-r2 | 2.097 |
|---|
| x2-y2 | 2.374 |
|---|
| xy | -0.497 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.442 |
1.848 |
0.000 |
| y |
1.848 |
5.444 |
0.000 |
| z |
0.000 |
0.000 |
2.917 |
<r2> (average value of r
2) Å
2
| <r2> |
83.175 |
| (<r2>)1/2 |
9.120 |