return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (Calicene)

using model chemistry: B2PLYP=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP=FULL/6-311G*
 hartrees
Energy at 0K-308.082370
Energy at 298.15K-308.087006
HF Energy-307.686885
Nuclear repulsion energy299.716314
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3311 3311 7.25      
2 A1 3245 3245 18.23      
3 A1 3222 3222 11.33      
4 A1 1819 1819 554.56      
5 A1 1571 1571 93.54      
6 A1 1497 1497 55.91      
7 A1 1416 1416 107.10      
8 A1 1219 1219 29.84      
9 A1 1087 1087 11.41      
10 A1 1029 1029 10.44      
11 A1 952 952 13.30      
12 A1 889 889 38.51      
13 A1 488 488 0.15      
14 A2 898 898 0.00      
15 A2 855 855 0.00      
16 A2 669 669 0.00      
17 A2 579 579 0.00      
18 A2 211 211 0.00      
19 B1 853 853 0.19      
20 B1 727 727 121.58      
21 B1 688 688 31.55      
22 B1 608 608 6.36      
23 B1 418 418 10.02      
24 B1 128 128 6.53      
25 B2 3271 3271 2.26      
26 B2 3237 3237 34.55      
27 B2 3213 3213 1.57      
28 B2 1580 1580 0.33      
29 B2 1351 1351 0.16      
30 B2 1236 1236 1.81      
31 B2 1110 1110 21.39      
32 B2 1102 1102 3.05      
33 B2 940 940 7.16      
34 B2 838 838 1.48      
35 B2 494 494 1.12      
36 B2 147 147 1.84      

Unscaled Zero Point Vibrational Energy (zpe) 23448.9 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 23448.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULL/6-311G*
ABC
0.22593 0.04745 0.03922

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 -0.046
C2 0.000 0.000 1.305
C3 0.000 0.664 2.571
C4 0.000 -0.664 2.571
C5 0.000 1.171 -0.911
C6 0.000 -1.171 -0.911
C7 0.000 0.727 -2.203
C8 0.000 -0.727 -2.203
H9 0.000 1.581 3.138
H10 0.000 -1.581 3.138
H11 0.000 2.197 -0.569
H12 0.000 -2.197 -0.569
H13 0.000 1.348 -3.090
H14 0.000 -1.348 -3.090

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.35062.69992.69991.45591.45592.27662.27663.55443.55442.25862.25863.32933.3293
C21.35061.42991.42992.50632.50633.58253.58252.42052.42052.88772.88774.59684.5968
C32.69991.42991.32783.51903.93614.77474.97291.07792.31533.49454.24815.70226.0080
C42.69991.42991.32783.93613.51904.97294.77472.31531.07794.24813.49456.00805.7022
C51.45592.50633.51903.93612.34171.36592.29624.06954.89551.08193.38542.18593.3306
C61.45592.50633.93613.51902.34172.29621.36594.89554.06953.38541.08193.33062.1859
C72.27663.58254.77474.97291.36592.29621.45495.40855.81832.19793.35021.08252.2573
C82.27663.58254.97294.77472.29621.36591.45495.81835.40853.35022.19792.25731.0825
H93.55442.42051.07792.31534.06954.89555.40855.81833.16203.75745.29286.23186.8821
H103.55442.42052.31531.07794.89554.06955.81835.40853.16205.29283.75746.88216.2318
H112.25862.88773.49454.24811.08193.38542.19793.35023.75745.29284.39442.66004.3504
H122.25862.88774.24813.49453.38541.08193.35022.19795.29283.75744.39444.35042.6600
H133.32934.59685.70226.00802.18593.33061.08252.25736.23186.88212.66004.35042.6967
H143.32934.59686.00805.70223.33062.18592.25731.08256.88216.23184.35042.66002.6967

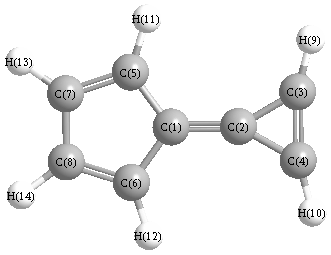
picture of Calicene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 152.334 C1 C2 C4 152.334
C1 C5 C7 107.524 C1 C5 H11 125.094
C1 C6 C8 107.524 C1 C6 H12 125.094
C2 C1 C5 126.466 C2 C1 C6 126.466
C2 C3 C4 62.334 C2 C3 H9 149.369
C2 C4 C3 62.334 C2 C4 H10 149.369
C3 C2 C4 55.332 C3 C4 H10 148.298
C4 C3 H9 148.298 C5 C1 C6 107.068
C5 C7 C8 108.942 C5 C7 H13 126.057
C6 C8 C7 108.942 C6 C8 H14 126.057
C7 C5 H11 127.383 C7 C8 H14 125.001
C8 C6 H12 127.383 C8 C7 H13 125.001
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability