return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SSH (Hydrogen methyl disulfide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-835.912915
Energy at 298.15K 
HF Energy-835.262986
Nuclear repulsion energy149.014643
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3203 3034 5.63 58.34 0.75 0.86
2 A 3190 3022 5.34 87.93 0.73 0.85
3 A 3090 2927 18.31 134.83 0.02 0.03
4 A 2649 2509 11.14 150.71 0.31 0.47
5 A 1515 1435 12.54 10.93 0.75 0.86
6 A 1497 1418 10.84 12.75 0.75 0.86
7 A 1424 1349 0.95 2.11 0.73 0.84
8 A 1029 974 7.96 2.97 0.72 0.84
9 A 1024 970 13.73 3.17 0.57 0.73
10 A 922 873 6.98 17.25 0.62 0.76
11 A 748 709 1.61 11.97 0.36 0.53
12 A 512 485 0.76 18.97 0.30 0.46
13 A 317 301 24.50 9.62 0.75 0.86
14 A 250 236 0.24 3.08 0.66 0.80
15 A 200 190 0.86 0.02 0.68 0.81

Unscaled Zero Point Vibrational Energy (zpe) 10785.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 10214.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.54855 0.14781 0.12201

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.613 0.702 -0.005
S2 -0.491 -0.713 0.015
S3 1.346 0.240 -0.088
H4 1.564 0.437 1.227
H5 -1.443 1.320 -0.885
H6 -2.627 0.297 -0.042
H7 -1.506 1.306 0.895

Atom - Atom Distances (Å)
  C1 S2 S3 H4 H5 H6 H7
C11.80612.99683.41811.08881.09181.0895
S21.80612.07212.64832.41822.36332.4256
S32.99682.07211.34683.09593.97373.2001
H43.41812.64831.34683.77954.38103.2082
H51.08882.41823.09593.77951.77651.7814
H61.09182.36333.97374.38101.77651.7758
H71.08952.42563.20013.20821.78141.7758

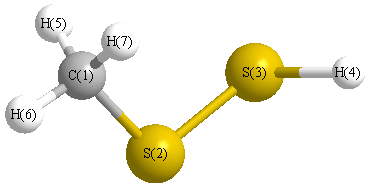
picture of Hydrogen methyl disulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 S3 100.979 S2 C1 H5 110.858
S2 C1 H6 106.673 S2 C1 H7 111.383
S2 S3 H4 99.346 H5 C1 H6 109.111
H5 C1 H7 109.732 H6 C1 H7 109.000
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability