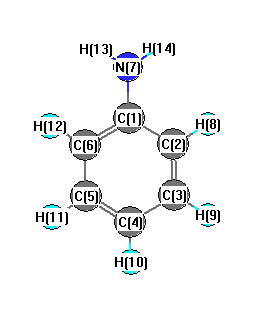
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3808 |
3444 |
10.24 |
|
|
|
| 2 |
A' |
3365 |
3043 |
24.96 |
|
|
|
| 3 |
A' |
3339 |
3020 |
11.57 |
|
|
|
| 4 |
A' |
3322 |
3004 |
20.95 |
|
|
|
| 5 |
A' |
1859 |
1681 |
106.64 |
|
|
|
| 6 |
A' |
1796 |
1624 |
69.48 |
|
|
|
| 7 |
A' |
1662 |
1503 |
68.75 |
|
|
|
| 8 |
A' |
1384 |
1252 |
56.86 |
|
|
|
| 9 |
A' |
1291 |
1168 |
9.58 |
|
|
|
| 10 |
A' |
1121 |
1014 |
2.69 |
|
|
|
| 11 |
A' |
1091 |
987 |
0.48 |
|
|
|
| 12 |
A' |
1078 |
975 |
1.33 |
|
|
|
| 13 |
A' |
982 |
888 |
22.24 |
|
|
|
| 14 |
A' |
887 |
802 |
13.89 |
|
|
|
| 15 |
A' |
843 |
763 |
126.18 |
|
|
|
| 16 |
A' |
767 |
694 |
20.88 |
|
|
|
| 17 |
A' |
726 |
657 |
281.53 |
|
|
|
| 18 |
A' |
573 |
518 |
1.85 |
|
|
|
| 19 |
A' |
553 |
501 |
55.26 |
|
|
|
| 20 |
A' |
244 |
220 |
6.54 |
|
|
|
| 21 |
A" |
3899 |
3526 |
7.92 |
|
|
|
| 22 |
A" |
3348 |
3028 |
65.94 |
|
|
|
| 23 |
A" |
3324 |
3006 |
5.45 |
|
|
|
| 24 |
A" |
1780 |
1609 |
11.42 |
|
|
|
| 25 |
A" |
1630 |
1474 |
4.36 |
|
|
|
| 26 |
A" |
1491 |
1348 |
0.02 |
|
|
|
| 27 |
A" |
1354 |
1224 |
5.55 |
|
|
|
| 28 |
A" |
1251 |
1131 |
1.68 |
|
|
|
| 29 |
A" |
1181 |
1068 |
20.37 |
|
|
|
| 30 |
A" |
1144 |
1035 |
0.44 |
|
|
|
| 31 |
A" |
1075 |
972 |
0.04 |
|
|
|
| 32 |
A" |
917 |
829 |
0.02 |
|
|
|
| 33 |
A" |
680 |
615 |
0.31 |
|
|
|
| 34 |
A" |
455 |
412 |
0.35 |
|
|
|
| 35 |
A" |
409 |
370 |
0.51 |
|
|
|
| 36 |
A" |
224 |
203 |
27.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27425.1 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 24803.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.353 |
|
|
0.512 |
| 2 |
C |
-0.267 |
|
|
-0.381 |
| 3 |
C |
-0.197 |
|
|
-0.090 |
| 4 |
C |
-0.254 |
|
|
-0.206 |
| 5 |
C |
-0.197 |
|
|
-0.090 |
| 6 |
C |
-0.267 |
|
|
-0.381 |
| 7 |
N |
-0.931 |
|
|
-0.920 |
| 8 |
H |
0.215 |
|
|
0.193 |
| 9 |
H |
0.214 |
|
|
0.145 |
| 10 |
H |
0.211 |
|
|
0.140 |
| 11 |
H |
0.214 |
|
|
0.145 |
| 12 |
H |
0.215 |
|
|
0.193 |
| 13 |
H |
0.345 |
|
|
0.370 |
| 14 |
H |
0.345 |
|
|
0.370 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.228 |
0.931 |
0.000 |
1.541 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.335 |
-3.865 |
0.000 |
| y |
-3.865 |
-35.455 |
0.000 |
| z |
0.000 |
0.000 |
-36.204 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.505 |
-3.865 |
0.000 |
| y |
-3.865 |
6.314 |
0.000 |
| z |
0.000 |
0.000 |
5.191 |
|
| Polar |
|---|
| 3z2-r2 | 10.383 |
|---|
| x2-y2 | -11.879 |
|---|
| xy | -3.865 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.658 |
-0.109 |
0.000 |
| y |
-0.109 |
12.707 |
0.000 |
| z |
0.000 |
0.000 |
11.136 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |