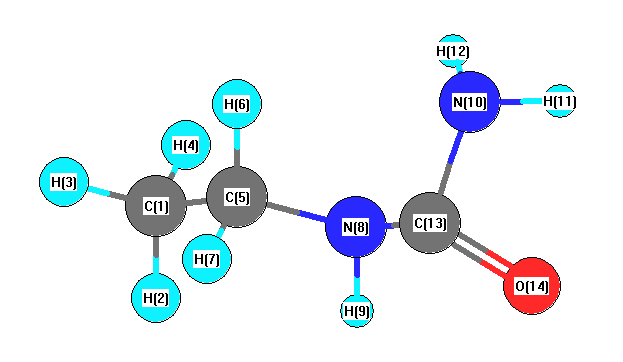
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3954 |
3576 |
28.86 |
|
|
|
| 2 |
A |
3877 |
3507 |
27.62 |
|
|
|
| 3 |
A |
3842 |
3475 |
29.15 |
|
|
|
| 4 |
A |
3258 |
2946 |
52.40 |
|
|
|
| 5 |
A |
3247 |
2937 |
71.98 |
|
|
|
| 6 |
A |
3200 |
2894 |
32.29 |
|
|
|
| 7 |
A |
3185 |
2880 |
18.96 |
|
|
|
| 8 |
A |
3178 |
2874 |
60.56 |
|
|
|
| 9 |
A |
1959 |
1771 |
682.64 |
|
|
|
| 10 |
A |
1821 |
1647 |
175.46 |
|
|
|
| 11 |
A |
1674 |
1514 |
3.74 |
|
|
|
| 12 |
A |
1643 |
1486 |
5.45 |
|
|
|
| 13 |
A |
1629 |
1474 |
8.67 |
|
|
|
| 14 |
A |
1625 |
1470 |
15.42 |
|
|
|
| 15 |
A |
1556 |
1408 |
42.57 |
|
|
|
| 16 |
A |
1550 |
1402 |
292.31 |
|
|
|
| 17 |
A |
1500 |
1357 |
37.64 |
|
|
|
| 18 |
A |
1427 |
1290 |
4.65 |
|
|
|
| 19 |
A |
1303 |
1179 |
17.44 |
|
|
|
| 20 |
A |
1271 |
1150 |
3.30 |
|
|
|
| 21 |
A |
1238 |
1119 |
54.96 |
|
|
|
| 22 |
A |
1156 |
1045 |
13.96 |
|
|
|
| 23 |
A |
1058 |
957 |
3.35 |
|
|
|
| 24 |
A |
968 |
875 |
3.24 |
|
|
|
| 25 |
A |
892 |
807 |
32.37 |
|
|
|
| 26 |
A |
878 |
794 |
66.36 |
|
|
|
| 27 |
A |
651 |
589 |
64.38 |
|
|
|
| 28 |
A |
615 |
556 |
21.81 |
|
|
|
| 29 |
A |
590 |
534 |
143.49 |
|
|
|
| 30 |
A |
534 |
483 |
49.59 |
|
|
|
| 31 |
A |
424 |
384 |
140.78 |
|
|
|
| 32 |
A |
387 |
350 |
0.82 |
|
|
|
| 33 |
A |
294 |
265 |
2.34 |
|
|
|
| 34 |
A |
215 |
195 |
3.21 |
|
|
|
| 35 |
A |
114 |
103 |
7.80 |
|
|
|
| 36 |
A |
64 |
58 |
0.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28388.2 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 25674.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.651 |
-0.365 |
|
-0.410 |
| 2 |
H |
0.216 |
0.100 |
|
0.116 |
| 3 |
H |
0.222 |
0.069 |
|
0.086 |
| 4 |
H |
0.223 |
0.096 |
|
0.108 |
| 5 |
C |
-0.208 |
0.598 |
|
0.540 |
| 6 |
H |
0.196 |
-0.066 |
|
-0.044 |
| 7 |
H |
0.217 |
-0.064 |
|
-0.046 |
| 8 |
N |
-0.712 |
-0.932 |
|
-0.878 |
| 9 |
H |
0.374 |
0.385 |
|
0.381 |
| 10 |
N |
-0.889 |
-1.044 |
|
-1.028 |
| 11 |
H |
0.378 |
0.418 |
|
0.423 |
| 12 |
H |
0.363 |
0.402 |
|
0.398 |
| 13 |
C |
0.786 |
1.125 |
|
1.053 |
| 14 |
O |
-0.517 |
-0.722 |
|
-0.698 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.207 |
2.903 |
-0.626 |
4.371 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-3.228 |
2.876 |
-0.685 |
4.377 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.946 |
7.278 |
-1.834 |
| y |
7.278 |
-35.450 |
-1.903 |
| z |
-1.834 |
-1.903 |
-38.141 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.150 |
7.278 |
-1.834 |
| y |
7.278 |
3.093 |
-1.903 |
| z |
-1.834 |
-1.903 |
-0.943 |
|
| Polar |
|---|
| 3z2-r2 | -1.886 |
|---|
| x2-y2 | -3.496 |
|---|
| xy | 7.278 |
|---|
| xz | -1.834 |
|---|
| yz | -1.903 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.170 |
-0.229 |
0.110 |
| y |
-0.229 |
6.955 |
-0.067 |
| z |
0.110 |
-0.067 |
5.083 |
<r2> (average value of r
2) Å
2
| <r2> |
200.477 |
| (<r2>)1/2 |
14.159 |